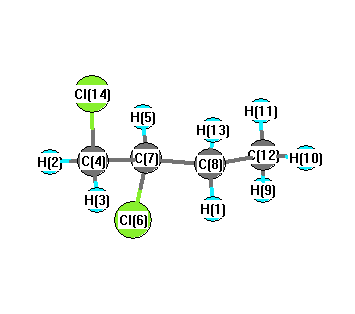
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3113 |
3062 |
3.48 |
|
|
|
| 2 |
A |
3078 |
3028 |
16.46 |
|
|
|
| 3 |
A |
3065 |
3015 |
29.40 |
|
|
|
| 4 |
A |
3046 |
2996 |
13.79 |
|
|
|
| 5 |
A |
3039 |
2988 |
2.02 |
|
|
|
| 6 |
A |
3022 |
2972 |
1.80 |
|
|
|
| 7 |
A |
2992 |
2942 |
24.25 |
|
|
|
| 8 |
A |
2982 |
2933 |
8.91 |
|
|
|
| 9 |
A |
1489 |
1464 |
8.06 |
|
|
|
| 10 |
A |
1480 |
1455 |
7.48 |
|
|
|
| 11 |
A |
1459 |
1435 |
4.07 |
|
|
|
| 12 |
A |
1453 |
1429 |
4.83 |
|
|
|
| 13 |
A |
1392 |
1369 |
4.00 |
|
|
|
| 14 |
A |
1370 |
1347 |
0.46 |
|
|
|
| 15 |
A |
1309 |
1288 |
5.48 |
|
|
|
| 16 |
A |
1298 |
1276 |
2.79 |
|
|
|
| 17 |
A |
1271 |
1250 |
10.03 |
|
|
|
| 18 |
A |
1221 |
1201 |
9.57 |
|
|
|
| 19 |
A |
1180 |
1160 |
12.60 |
|
|
|
| 20 |
A |
1109 |
1091 |
1.82 |
|
|
|
| 21 |
A |
1078 |
1060 |
1.40 |
|
|
|
| 22 |
A |
1047 |
1030 |
3.83 |
|
|
|
| 23 |
A |
1024 |
1007 |
0.64 |
|
|
|
| 24 |
A |
933 |
917 |
11.91 |
|
|
|
| 25 |
A |
817 |
803 |
6.40 |
|
|
|
| 26 |
A |
799 |
786 |
12.53 |
|
|
|
| 27 |
A |
716 |
704 |
52.30 |
|
|
|
| 28 |
A |
640 |
630 |
45.23 |
|
|
|
| 29 |
A |
452 |
445 |
1.91 |
|
|
|
| 30 |
A |
376 |
370 |
1.96 |
|
|
|
| 31 |
A |
287 |
282 |
0.10 |
|
|
|
| 32 |
A |
234 |
230 |
0.58 |
|
|
|
| 33 |
A |
213 |
210 |
3.13 |
|
|
|
| 34 |
A |
184 |
181 |
3.92 |
|
|
|
| 35 |
A |
115 |
113 |
3.41 |
|
|
|
| 36 |
A |
95 |
94 |
1.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24689.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 24281.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.170 |
|
|
|
| 2 |
H |
0.228 |
|
|
|
| 3 |
H |
0.220 |
|
|
|
| 4 |
C |
-0.400 |
|
|
|
| 5 |
H |
0.214 |
|
|
|
| 6 |
Cl |
-0.066 |
|
|
|
| 7 |
C |
-0.237 |
|
|
|
| 8 |
C |
-0.279 |
|
|
|
| 9 |
H |
0.184 |
|
|
|
| 10 |
H |
0.167 |
|
|
|
| 11 |
H |
0.163 |
|
|
|
| 12 |
C |
-0.490 |
|
|
|
| 13 |
H |
0.189 |
|
|
|
| 14 |
Cl |
-0.062 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.339 |
0.654 |
0.224 |
0.770 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-57.202 |
3.399 |
-0.265 |
| y |
3.399 |
-50.612 |
-0.185 |
| z |
-0.265 |
-0.185 |
-50.262 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.765 |
3.399 |
-0.265 |
| y |
3.399 |
3.120 |
-0.185 |
| z |
-0.265 |
-0.185 |
3.646 |
|
| Polar |
|---|
| 3z2-r2 | 7.291 |
|---|
| x2-y2 | -6.590 |
|---|
| xy | 3.399 |
|---|
| xz | -0.265 |
|---|
| yz | -0.185 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.676 |
-1.101 |
0.218 |
| y |
-1.101 |
9.772 |
-0.142 |
| z |
0.218 |
-0.142 |
7.277 |
<r2> (average value of r
2) Å
2
| <r2> |
326.378 |
| (<r2>)1/2 |
18.066 |