return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6S (Thiophene, 3-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-591.933185
Energy at 298.15K-591.939350
HF Energy-591.675900
Nuclear repulsion energy269.501398
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3296 3129 1.05      
2 A' 3285 3118 1.06      
3 A' 3240 3076 5.88      
4 A' 3162 3001 16.61      
5 A' 3075 2919 31.80      
6 A' 1617 1535 2.19      
7 A' 1546 1467 8.19      
8 A' 1479 1404 6.96      
9 A' 1463 1388 0.44      
10 A' 1438 1365 3.33      
11 A' 1282 1216 4.40      
12 A' 1207 1145 2.05      
13 A' 1129 1072 3.80      
14 A' 1035 982 7.90      
15 A' 959 910 2.80      
16 A' 879 834 15.32      
17 A' 847 804 6.89      
18 A' 671 637 0.53      
19 A' 549 521 0.59      
20 A' 321 304 0.77      
21 A" 3135 2976 20.61      
22 A" 1534 1456 5.65      
23 A" 1085 1030 2.57      
24 A" 874 830 0.03      
25 A" 771 731 73.61      
26 A" 679 644 1.94      
27 A" 596 566 7.21      
28 A" 457 434 0.20      
29 A" 234 222 2.57      
30 A" 111 105 0.08      

Unscaled Zero Point Vibrational Energy (zpe) 20976.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 19910.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.23171 0.08134 0.06089

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.219 0.791 0.000
C2 -1.258 0.287 0.000
C3 0.000 0.986 0.000
C4 1.054 0.112 0.000
H5 -1.895 -1.823 0.000
C6 -1.117 -1.072 0.000
S7 0.549 -1.547 0.000
H8 1.164 2.799 0.000
H9 -0.364 2.920 0.882
H10 -0.364 2.920 -0.882
C11 0.118 2.486 0.000
H12 2.109 0.352 0.000

Atom - Atom Distances (Å)
  H1 C2 C3 C4 H5 C6 S7 H8 H9 H10 C11 H12
H11.08492.22703.34222.63412.16423.62323.93422.95832.95832.88634.3498
C21.08491.43902.31852.20391.36592.57473.49012.91752.91752.59413.3676
C32.22701.43901.36883.38792.34092.59162.15542.15712.15711.50522.2021
C43.34222.31851.36883.52702.47271.73442.68943.26693.26692.55201.0820
H52.63412.20393.38793.52701.08142.45945.54305.06165.06164.75594.5564
C62.16421.36592.34092.47271.08141.73254.49334.15704.15703.76603.5261
S73.62322.57472.59161.73442.45941.73254.38984.64414.64414.05632.4576
H83.93423.49012.15542.68945.54304.49334.38981.76881.76881.09272.6232
H92.95832.91752.15713.26695.06164.15704.64411.76881.76461.09483.6726
H102.95832.91752.15713.26695.06164.15704.64411.76881.76461.09483.6726
C112.88632.59411.50522.55204.75593.76604.05631.09271.09481.09482.9189
H124.34983.36762.20211.08204.55643.52612.45762.62323.67263.67262.9189

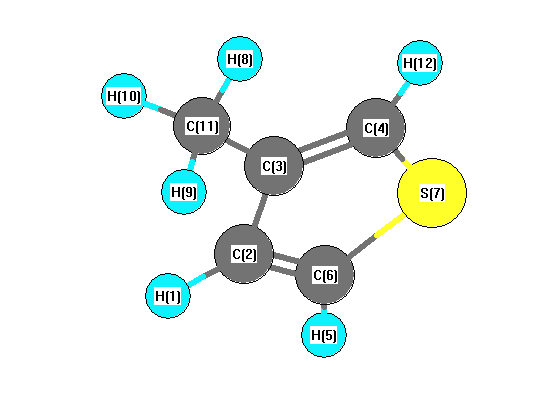
picture of Thiophene, 3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C3 123.255 H1 C2 C6 123.625
C2 C3 C4 111.302 C2 C3 C11 123.533
C2 C6 H5 128.076 C2 C6 S7 111.856
C3 C2 C6 113.120 C3 C4 S7 112.726
C3 C4 H12 127.543 C3 C11 H8 111.138
C3 C11 H9 111.151 C3 C11 H10 111.151
C4 C3 C11 125.166 C4 S7 C6 90.996
H5 C6 S7 120.068 S7 C4 H12 119.731
H8 C11 H9 107.917 H8 C11 H10 107.917
H9 C11 H10 107.400
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.165      
2 C -0.144      
3 C 0.134      
4 C -0.419      
5 H 0.198      
6 C -0.378      
7 S 0.266      
8 H 0.165      
9 H 0.169      
10 H 0.169      
11 C -0.516      
12 H 0.192      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.144 -0.434 0.000
y -0.434 12.191 0.000
z 0.000 0.000 4.940


<r2> (average value of r2) Å2
<r2> 188.025
(<r2>)1/2 13.712