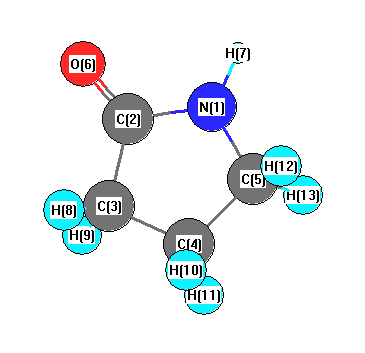
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3542 |
3484 |
18.70 |
|
|
|
| 2 |
A |
3075 |
3024 |
17.43 |
|
|
|
| 3 |
A |
3064 |
3013 |
16.57 |
|
|
|
| 4 |
A |
3020 |
2970 |
49.04 |
|
|
|
| 5 |
A |
3007 |
2958 |
10.63 |
|
|
|
| 6 |
A |
2997 |
2947 |
13.89 |
|
|
|
| 7 |
A |
2934 |
2886 |
69.10 |
|
|
|
| 8 |
A |
1789 |
1759 |
366.43 |
|
|
|
| 9 |
A |
1510 |
1485 |
3.28 |
|
|
|
| 10 |
A |
1475 |
1451 |
6.28 |
|
|
|
| 11 |
A |
1448 |
1425 |
3.51 |
|
|
|
| 12 |
A |
1411 |
1388 |
33.22 |
|
|
|
| 13 |
A |
1335 |
1313 |
9.29 |
|
|
|
| 14 |
A |
1308 |
1286 |
12.36 |
|
|
|
| 15 |
A |
1268 |
1248 |
22.77 |
|
|
|
| 16 |
A |
1234 |
1213 |
62.60 |
|
|
|
| 17 |
A |
1214 |
1194 |
17.53 |
|
|
|
| 18 |
A |
1181 |
1161 |
2.07 |
|
|
|
| 19 |
A |
1157 |
1138 |
3.66 |
|
|
|
| 20 |
A |
1070 |
1052 |
6.99 |
|
|
|
| 21 |
A |
1060 |
1043 |
8.03 |
|
|
|
| 22 |
A |
991 |
975 |
11.13 |
|
|
|
| 23 |
A |
911 |
896 |
0.24 |
|
|
|
| 24 |
A |
891 |
876 |
1.11 |
|
|
|
| 25 |
A |
873 |
859 |
3.39 |
|
|
|
| 26 |
A |
796 |
783 |
6.30 |
|
|
|
| 27 |
A |
677 |
666 |
7.57 |
|
|
|
| 28 |
A |
616 |
606 |
21.16 |
|
|
|
| 29 |
A |
551 |
542 |
42.32 |
|
|
|
| 30 |
A |
481 |
473 |
63.20 |
|
|
|
| 31 |
A |
450 |
443 |
8.83 |
|
|
|
| 32 |
A |
189 |
186 |
4.54 |
|
|
|
| 33 |
A |
141 |
139 |
1.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23832.8 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23439.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.552 |
|
|
|
| 2 |
C |
0.551 |
|
|
|
| 3 |
C |
-0.402 |
|
|
|
| 4 |
C |
-0.324 |
|
|
|
| 5 |
C |
-0.155 |
|
|
|
| 6 |
O |
-0.476 |
|
|
|
| 7 |
H |
0.325 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.187 |
|
|
|
| 10 |
H |
0.164 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
H |
0.160 |
|
|
|
| 13 |
H |
0.166 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.686 |
-0.716 |
0.351 |
3.771 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.729 |
-0.273 |
0.199 |
| y |
-0.273 |
-31.037 |
-0.202 |
| z |
0.199 |
-0.202 |
-35.200 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.611 |
-0.273 |
0.199 |
| y |
-0.273 |
7.928 |
-0.202 |
| z |
0.199 |
-0.202 |
1.683 |
|
| Polar |
|---|
| 3z2-r2 | 3.365 |
|---|
| x2-y2 | -11.692 |
|---|
| xy | -0.273 |
|---|
| xz | 0.199 |
|---|
| yz | -0.202 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.805 |
0.458 |
-0.106 |
| y |
0.458 |
7.635 |
0.055 |
| z |
-0.106 |
0.055 |
5.558 |
<r2> (average value of r
2) Å
2
| <r2> |
147.395 |
| (<r2>)1/2 |
12.141 |