return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (E)-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2h 1Ag
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-156.676700
Energy at 298.15K-156.684412
HF Energy-156.109056
Nuclear repulsion energy115.430519
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3154 3003 0.00      
2 A 3137 2987 0.00      
3 A 3054 2908 0.00      
4 A 1783 1698 0.00      
5 A 1544 1470 0.00      
6 A 1469 1399 0.00      
7 A 1361 1296 0.00      
8 A 1199 1142 0.00      
9 A 901 858 0.00      
10 A 508 483 0.00      
11 A 3114 2965 55.13      
12 A 1531 1458 11.06      
13 A 1101 1049 0.75      
14 A 1005 957 29.36      
15 A 247 235 3.00      
16 A 167 159 0.71      
17 A 3114 2966 0.00      
18 A 1531 1458 0.00      
19 A 1090 1038 0.00      
20 A 751 715 0.00      
21 A 214 203 0.00      
22 A 3159 3008 75.65      
23 A 3135 2985 18.51      
24 A 3053 2908 60.21      
25 A 1549 1475 13.56      
26 A 1468 1398 4.04      
27 A 1368 1303 3.00      
28 A 1112 1059 0.84      
29 A 1015 967 10.12      
30 A 286 273 0.74      

Unscaled Zero Point Vibrational Energy (zpe) 24059.0 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 22911.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
1.14863 0.12360 0.11644

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.670 0.000
C2 0.000 -0.670 0.000
C3 -1.235 1.528 0.000
C4 1.235 -1.528 0.000
H5 0.957 1.199 0.000
H6 -0.957 -1.199 0.000
H7 -2.143 0.913 0.000
H8 2.143 -0.913 0.000
H9 -1.266 2.181 0.884
H10 -1.266 2.181 -0.884
H11 1.266 -2.181 0.884
H12 1.266 -2.181 -0.884

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.33971.50442.52161.09362.09942.15692.66422.16002.16003.24183.2418
C21.33972.52161.50442.09941.09362.66422.15693.24183.24182.16002.1600
C31.50442.52163.93052.21732.74111.09684.16821.09881.09884.56014.5601
C42.52161.50443.93052.74112.21734.16821.09684.56014.56011.09881.0988
H51.09362.09942.21732.74113.06793.11362.42162.58602.58603.50663.5066
H62.09941.09362.74112.21733.06792.42163.11363.50663.50662.58602.5860
H72.15692.66421.09684.16823.11362.42164.65901.77721.77724.68744.6874
H82.66422.15694.16821.09682.42163.11364.65904.68744.68741.77721.7772
H92.16003.24181.09884.56012.58603.50661.77724.68741.76715.04305.3437
H102.16003.24181.09884.56012.58603.50661.77724.68741.76715.34375.0430
H113.24182.16004.56011.09883.50662.58604.68741.77725.04305.34371.7671
H123.24182.16004.56011.09883.50662.58604.68741.77725.34375.04301.7671

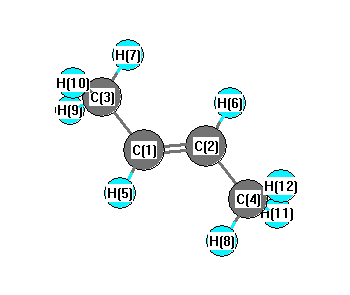
picture of 2-Butene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 124.797 C1 C2 H6 118.913
C1 C3 H7 111.062 C1 C3 H9 111.193
C1 C3 H10 111.193 C2 C1 C3 124.797
C2 C1 H5 118.913 C2 C4 H8 111.062
C2 C4 H11 111.193 C2 C4 H12 111.193
C3 C1 H5 116.290 C4 C2 H6 116.290
H7 C3 H9 108.085 H7 C3 H10 108.085
H8 C4 H11 108.085 H8 C4 H12 108.085
H9 C3 H10 107.056 H11 C4 H12 107.056
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability