Jump to
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -278.176848 |
| Energy at 298.15K | |
| HF Energy | -278.176848 |
| Nuclear repulsion energy | 125.701720 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3069 |
2936 |
0.00 |
|
|
|
| 2 |
Ag |
1547 |
1480 |
0.00 |
|
|
|
| 3 |
Ag |
1481 |
1417 |
0.00 |
|
|
|
| 4 |
Ag |
1105 |
1057 |
0.00 |
|
|
|
| 5 |
Ag |
1096 |
1049 |
0.00 |
|
|
|
| 6 |
Ag |
462 |
442 |
0.00 |
|
|
|
| 7 |
Au |
3139 |
3003 |
76.11 |
|
|
|
| 8 |
Au |
1246 |
1192 |
2.54 |
|
|
|
| 9 |
Au |
822 |
787 |
0.02 |
|
|
|
| 10 |
Au |
135 |
130 |
11.24 |
|
|
|
| 11 |
Bg |
3114 |
2979 |
0.00 |
|
|
|
| 12 |
Bg |
1313 |
1256 |
0.00 |
|
|
|
| 13 |
Bg |
1194 |
1142 |
0.00 |
|
|
|
| 14 |
Bu |
3075 |
2942 |
70.33 |
|
|
|
| 15 |
Bu |
1558 |
1491 |
2.17 |
|
|
|
| 16 |
Bu |
1380 |
1320 |
17.14 |
|
|
|
| 17 |
Bu |
1118 |
1069 |
185.42 |
|
|
|
| 18 |
Bu |
274 |
262 |
17.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13563.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12976.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.423 |
0.630 |
0.000 |
| C2 |
-0.423 |
-0.630 |
0.000 |
| F3 |
-0.423 |
1.724 |
0.000 |
| F4 |
0.423 |
-1.724 |
0.000 |
| H5 |
1.059 |
0.665 |
0.892 |
| H6 |
1.059 |
0.665 |
-0.892 |
| H7 |
-1.059 |
-0.665 |
0.892 |
| H8 |
-1.059 |
-0.665 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5173 | 1.3830 | 2.3537 | 1.0961 | 1.0961 | 2.1607 | 2.1607 |
C2 | 1.5173 | | 2.3537 | 1.3830 | 2.1607 | 2.1607 | 1.0961 | 1.0961 | F3 | 1.3830 | 2.3537 | | 3.5500 | 2.0283 | 2.0283 | 2.6279 | 2.6279 | F4 | 2.3537 | 1.3830 | 3.5500 | | 2.6279 | 2.6279 | 2.0283 | 2.0283 | H5 | 1.0961 | 2.1607 | 2.0283 | 2.6279 | | 1.7836 | 2.5013 | 3.0721 | H6 | 1.0961 | 2.1607 | 2.0283 | 2.6279 | 1.7836 | | 3.0721 | 2.5013 | H7 | 2.1607 | 1.0961 | 2.6279 | 2.0283 | 2.5013 | 3.0721 | | 1.7836 | H8 | 2.1607 | 1.0961 | 2.6279 | 2.0283 | 3.0721 | 2.5013 | 1.7836 | |
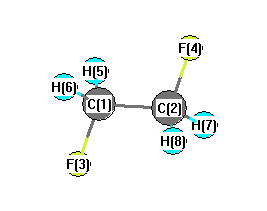 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.401 |
|
C1 |
C2 |
H7 |
110.500 |
| C1 |
C2 |
H8 |
110.500 |
|
C2 |
C1 |
F3 |
108.401 |
| C2 |
C1 |
H5 |
110.500 |
|
C2 |
C1 |
H6 |
110.500 |
| F3 |
C1 |
H5 |
109.259 |
|
F3 |
C1 |
H6 |
109.259 |
| F4 |
C2 |
H7 |
109.259 |
|
F4 |
C2 |
H8 |
109.259 |
| H5 |
C1 |
H6 |
108.899 |
|
H7 |
C2 |
H8 |
108.899 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.029 |
|
|
|
| 2 |
C |
-0.029 |
|
|
|
| 3 |
F |
-0.309 |
|
|
|
| 4 |
F |
-0.309 |
|
|
|
| 5 |
H |
0.169 |
|
|
|
| 6 |
H |
0.169 |
|
|
|
| 7 |
H |
0.169 |
|
|
|
| 8 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.522 |
2.517 |
0.000 |
| y |
2.517 |
-27.973 |
0.000 |
| z |
0.000 |
0.000 |
-21.206 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.068 |
2.517 |
0.000 |
| y |
2.517 |
-6.609 |
0.000 |
| z |
0.000 |
0.000 |
3.541 |
|
| Polar |
|---|
| 3z2-r2 | 7.083 |
|---|
| x2-y2 | 6.451 |
|---|
| xy | 2.517 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.523 |
0.032 |
0.000 |
| y |
0.032 |
3.459 |
0.000 |
| z |
0.000 |
0.000 |
3.392 |
<r2> (average value of r
2) Å
2
| <r2> |
87.776 |
| (<r2>)1/2 |
9.369 |
Jump to
S1C1
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -278.177458 |
| Energy at 298.15K | |
| HF Energy | -278.177458 |
| Nuclear repulsion energy | 127.829515 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3106 |
2971 |
22.51 |
|
|
|
| 2 |
A |
3061 |
2929 |
42.80 |
|
|
|
| 3 |
A |
1527 |
1461 |
0.51 |
|
|
|
| 4 |
A |
1465 |
1401 |
19.96 |
|
|
|
| 5 |
A |
1317 |
1260 |
1.43 |
|
|
|
| 6 |
A |
1155 |
1105 |
36.51 |
|
|
|
| 7 |
A |
1139 |
1090 |
48.16 |
|
|
|
| 8 |
A |
891 |
852 |
20.38 |
|
|
|
| 9 |
A |
324 |
310 |
0.39 |
|
|
|
| 10 |
A |
149 |
143 |
3.01 |
|
|
|
| 11 |
B |
3120 |
2985 |
58.00 |
|
|
|
| 12 |
B |
3049 |
2917 |
16.54 |
|
|
|
| 13 |
B |
1524 |
1458 |
6.95 |
|
|
|
| 14 |
B |
1432 |
1370 |
14.04 |
|
|
|
| 15 |
B |
1273 |
1218 |
4.31 |
|
|
|
| 16 |
B |
1121 |
1073 |
43.94 |
|
|
|
| 17 |
B |
923 |
883 |
33.55 |
|
|
|
| 18 |
B |
495 |
474 |
14.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13535.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 12949.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.263 |
0.706 |
0.511 |
| C2 |
-0.263 |
-0.706 |
0.511 |
| F3 |
-0.263 |
1.420 |
-0.549 |
| F4 |
0.263 |
-1.420 |
-0.549 |
| H5 |
-0.020 |
1.204 |
1.448 |
| H6 |
1.356 |
0.702 |
0.425 |
| H7 |
0.020 |
-1.204 |
1.448 |
| H8 |
-1.356 |
-0.702 |
0.425 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5064 | 1.3812 | 2.3749 | 1.0984 | 1.0967 | 2.1411 | 2.1467 |
C2 | 1.5064 | | 2.3749 | 1.3812 | 2.1411 | 2.1467 | 1.0984 | 1.0967 | F3 | 1.3812 | 2.3749 | | 2.8873 | 2.0232 | 2.0204 | 3.3090 | 2.5774 | F4 | 2.3749 | 1.3812 | 2.8873 | | 3.3090 | 2.5774 | 2.0232 | 2.0204 | H5 | 1.0984 | 2.1411 | 2.0232 | 3.3090 | | 1.7870 | 2.4078 | 2.5421 | H6 | 1.0967 | 2.1467 | 2.0204 | 2.5774 | 1.7870 | | 2.5421 | 3.0534 | H7 | 2.1411 | 1.0984 | 3.3090 | 2.0232 | 2.4078 | 2.5421 | | 1.7870 | H8 | 2.1467 | 1.0967 | 2.5774 | 2.0204 | 2.5421 | 3.0534 | 1.7870 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.593 |
|
C1 |
C2 |
H7 |
109.571 |
| C1 |
C2 |
H8 |
110.116 |
|
C2 |
C1 |
F3 |
110.593 |
| C2 |
C1 |
H5 |
109.571 |
|
C2 |
C1 |
H6 |
110.116 |
| F3 |
C1 |
H5 |
108.825 |
|
F3 |
C1 |
H6 |
108.712 |
| F4 |
C2 |
H7 |
108.825 |
|
F4 |
C2 |
H8 |
108.712 |
| H5 |
C1 |
H6 |
108.990 |
|
H7 |
C2 |
H8 |
108.990 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.019 |
|
|
|
| 2 |
C |
-0.019 |
|
|
|
| 3 |
F |
-0.305 |
|
|
|
| 4 |
F |
-0.305 |
|
|
|
| 5 |
H |
0.153 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
H |
0.153 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.453 |
2.453 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.291 |
1.392 |
0.000 |
| y |
1.392 |
-25.354 |
0.000 |
| z |
0.000 |
0.000 |
-21.774 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.273 |
1.392 |
0.000 |
| y |
1.392 |
-3.822 |
0.000 |
| z |
0.000 |
0.000 |
1.549 |
|
| Polar |
|---|
| 3z2-r2 | 3.098 |
|---|
| x2-y2 | 4.063 |
|---|
| xy | 1.392 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.451 |
0.038 |
0.000 |
| y |
0.038 |
3.563 |
0.000 |
| z |
0.000 |
0.000 |
3.499 |
<r2> (average value of r
2) Å
2
| <r2> |
79.232 |
| (<r2>)1/2 |
8.901 |
