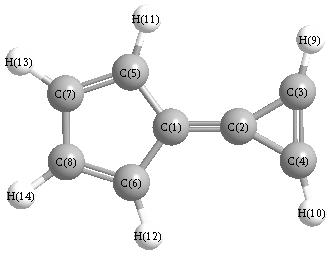
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3325 |
3181 |
6.05 |
|
|
|
| 2 |
A1 |
3257 |
3116 |
14.13 |
|
|
|
| 3 |
A1 |
3234 |
3094 |
11.63 |
|
|
|
| 4 |
A1 |
1841 |
1762 |
511.57 |
|
|
|
| 5 |
A1 |
1607 |
1537 |
122.01 |
|
|
|
| 6 |
A1 |
1520 |
1454 |
80.68 |
|
|
|
| 7 |
A1 |
1430 |
1368 |
96.85 |
|
|
|
| 8 |
A1 |
1235 |
1181 |
23.63 |
|
|
|
| 9 |
A1 |
1091 |
1043 |
10.08 |
|
|
|
| 10 |
A1 |
1038 |
993 |
9.16 |
|
|
|
| 11 |
A1 |
956 |
915 |
12.93 |
|
|
|
| 12 |
A1 |
890 |
852 |
37.24 |
|
|
|
| 13 |
A1 |
491 |
470 |
0.05 |
|
|
|
| 14 |
A2 |
919 |
879 |
0.00 |
|
|
|
| 15 |
A2 |
907 |
867 |
0.00 |
|
|
|
| 16 |
A2 |
699 |
669 |
0.00 |
|
|
|
| 17 |
A2 |
592 |
566 |
0.00 |
|
|
|
| 18 |
A2 |
211 |
202 |
0.00 |
|
|
|
| 19 |
B1 |
886 |
848 |
0.00 |
|
|
|
| 20 |
B1 |
762 |
729 |
63.46 |
|
|
|
| 21 |
B1 |
725 |
694 |
63.46 |
|
|
|
| 22 |
B1 |
633 |
605 |
3.29 |
|
|
|
| 23 |
B1 |
437 |
418 |
8.31 |
|
|
|
| 24 |
B1 |
137 |
131 |
5.50 |
|
|
|
| 25 |
B2 |
3286 |
3143 |
2.59 |
|
|
|
| 26 |
B2 |
3250 |
3109 |
28.98 |
|
|
|
| 27 |
B2 |
3223 |
3083 |
2.90 |
|
|
|
| 28 |
B2 |
1608 |
1538 |
0.34 |
|
|
|
| 29 |
B2 |
1359 |
1301 |
0.34 |
|
|
|
| 30 |
B2 |
1240 |
1186 |
1.64 |
|
|
|
| 31 |
B2 |
1123 |
1074 |
16.89 |
|
|
|
| 32 |
B2 |
1108 |
1060 |
6.58 |
|
|
|
| 33 |
B2 |
949 |
908 |
6.65 |
|
|
|
| 34 |
B2 |
830 |
794 |
1.75 |
|
|
|
| 35 |
B2 |
496 |
475 |
0.95 |
|
|
|
| 36 |
B2 |
147 |
141 |
1.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23719.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22692.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.048 |
|
|
|
| 2 |
C |
0.104 |
|
|
|
| 3 |
C |
-0.144 |
|
|
|
| 4 |
C |
-0.144 |
|
|
|
| 5 |
C |
-0.178 |
|
|
|
| 6 |
C |
-0.178 |
|
|
|
| 7 |
C |
-0.194 |
|
|
|
| 8 |
C |
-0.194 |
|
|
|
| 9 |
H |
0.209 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.137 |
|
|
|
| 12 |
H |
0.137 |
|
|
|
| 13 |
H |
0.142 |
|
|
|
| 14 |
H |
0.142 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
4.537 |
4.537 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.463 |
0.000 |
0.000 |
| y |
0.000 |
-40.392 |
0.000 |
| z |
0.000 |
0.000 |
-34.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.125 |
0.000 |
0.000 |
| y |
0.000 |
1.482 |
0.000 |
| z |
0.000 |
0.000 |
10.644 |
|
| Polar |
|---|
| 3z2-r2 | 21.288 |
|---|
| x2-y2 | -9.071 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.140 |
0.000 |
0.000 |
| y |
0.000 |
11.566 |
0.000 |
| z |
0.000 |
0.000 |
20.322 |
<r2> (average value of r
2) Å
2
| <r2> |
269.185 |
| (<r2>)1/2 |
16.407 |