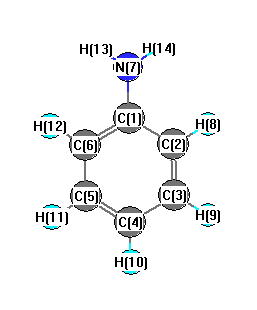
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3575 |
3421 |
13.81 |
|
|
|
| 2 |
A' |
3222 |
3083 |
17.02 |
|
|
|
| 3 |
A' |
3199 |
3060 |
3.62 |
|
|
|
| 4 |
A' |
3181 |
3044 |
19.03 |
|
|
|
| 5 |
A' |
1700 |
1626 |
142.56 |
|
|
|
| 6 |
A' |
1678 |
1605 |
16.52 |
|
|
|
| 7 |
A' |
1556 |
1488 |
60.59 |
|
|
|
| 8 |
A' |
1329 |
1271 |
49.62 |
|
|
|
| 9 |
A' |
1212 |
1159 |
7.55 |
|
|
|
| 10 |
A' |
1062 |
1016 |
2.67 |
|
|
|
| 11 |
A' |
1011 |
967 |
0.51 |
|
|
|
| 12 |
A' |
976 |
934 |
0.22 |
|
|
|
| 13 |
A' |
878 |
840 |
11.98 |
|
|
|
| 14 |
A' |
839 |
802 |
2.30 |
|
|
|
| 15 |
A' |
763 |
729 |
81.64 |
|
|
|
| 16 |
A' |
704 |
673 |
12.86 |
|
|
|
| 17 |
A' |
616 |
589 |
309.86 |
|
|
|
| 18 |
A' |
535 |
512 |
2.94 |
|
|
|
| 19 |
A' |
507 |
485 |
59.48 |
|
|
|
| 20 |
A' |
223 |
214 |
5.46 |
|
|
|
| 21 |
A" |
3678 |
3519 |
10.67 |
|
|
|
| 22 |
A" |
3204 |
3066 |
42.84 |
|
|
|
| 23 |
A" |
3183 |
3045 |
4.96 |
|
|
|
| 24 |
A" |
1658 |
1586 |
5.03 |
|
|
|
| 25 |
A" |
1521 |
1455 |
1.82 |
|
|
|
| 26 |
A" |
1390 |
1330 |
8.72 |
|
|
|
| 27 |
A" |
1372 |
1313 |
0.02 |
|
|
|
| 28 |
A" |
1189 |
1137 |
1.58 |
|
|
|
| 29 |
A" |
1151 |
1101 |
3.64 |
|
|
|
| 30 |
A" |
1082 |
1035 |
2.86 |
|
|
|
| 31 |
A" |
950 |
909 |
0.00 |
|
|
|
| 32 |
A" |
826 |
790 |
0.11 |
|
|
|
| 33 |
A" |
632 |
605 |
0.30 |
|
|
|
| 34 |
A" |
416 |
398 |
0.36 |
|
|
|
| 35 |
A" |
383 |
366 |
0.21 |
|
|
|
| 36 |
A" |
292 |
280 |
22.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25844.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24725.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.298 |
|
|
0.411 |
| 2 |
C |
-0.202 |
|
|
-0.293 |
| 3 |
C |
-0.169 |
|
|
-0.119 |
| 4 |
C |
-0.173 |
|
|
-0.148 |
| 5 |
C |
-0.169 |
|
|
-0.119 |
| 6 |
C |
-0.202 |
|
|
-0.293 |
| 7 |
N |
-0.821 |
|
|
-0.853 |
| 8 |
H |
0.148 |
|
|
0.158 |
| 9 |
H |
0.157 |
|
|
0.129 |
| 10 |
H |
0.154 |
|
|
0.118 |
| 11 |
H |
0.157 |
|
|
0.129 |
| 12 |
H |
0.148 |
|
|
0.158 |
| 13 |
H |
0.337 |
|
|
0.361 |
| 14 |
H |
0.337 |
|
|
0.361 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.163 |
1.323 |
0.000 |
1.761 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-1.283 |
1.308 |
0.000 |
1.832 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.404 |
-3.566 |
0.000 |
| y |
-3.566 |
-33.645 |
0.000 |
| z |
0.000 |
0.000 |
-35.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.725 |
-3.566 |
0.000 |
| y |
-3.566 |
6.914 |
0.000 |
| z |
0.000 |
0.000 |
3.811 |
|
| Polar |
|---|
| 3z2-r2 | 7.621 |
|---|
| x2-y2 | -11.759 |
|---|
| xy | -3.566 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.655 |
-0.101 |
0.000 |
| y |
-0.101 |
13.523 |
0.000 |
| z |
0.000 |
0.000 |
11.261 |
<r2> (average value of r
2) Å
2
| <r2> |
191.414 |
| (<r2>)1/2 |
13.835 |