return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5NH2 (aniline)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-287.273129
Energy at 298.15K-287.281110
HF Energy-286.967045
Nuclear repulsion energy271.082337
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3564 3383 10.84      
2 A' 3238 3074 17.63      
3 A' 3215 3052 2.68      
4 A' 3198 3035 17.65      
5 A' 1712 1625 96.63      
6 A' 1679 1594 40.82      
7 A' 1561 1482 55.32      
8 A' 1325 1257 48.92      
9 A' 1222 1160 6.31      
10 A' 1063 1009 2.01      
11 A' 1018 967 0.45      
12 A' 942 894 0.15      
13 A' 858 814 9.49      
14 A' 839 797 2.89      
15 A' 751 713 135.90      
16 A' 684 650 163.86      
17 A' 653 619 128.55      
18 A' 538 511 2.38      
19 A' 503 478 34.96      
20 A' 224 212 5.05      
21 A" 3665 3479 8.08      
22 A" 3221 3058 40.24      
23 A" 3199 3036 5.66      
24 A" 1659 1575 3.59      
25 A" 1528 1450 2.66      
26 A" 1395 1324 8.02      
27 A" 1386 1316 0.24      
28 A" 1201 1140 1.59      
29 A" 1163 1104 2.54      
30 A" 1090 1034 2.97      
31 A" 929 882 0.00      
32 A" 821 780 0.15      
33 A" 638 605 0.25      
34 A" 413 392 0.41      
35 A" 383 364 0.23      
36 A" 274 260 25.01      

Unscaled Zero Point Vibrational Energy (zpe) 25875.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24561.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.18732 0.08637 0.05922

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.002 0.938 0.000
C2 0.005 0.222 1.209
C3 0.005 -1.172 1.204
C4 0.003 -1.882 0.000
C5 0.005 -1.172 -1.204
C6 0.005 0.222 -1.209
N7 0.063 2.339 0.000
H8 0.012 0.763 2.152
H9 0.004 -1.706 2.150
H10 0.002 -2.967 0.000
H11 0.004 -1.706 -2.150
H12 0.012 0.763 -2.152
H13 -0.311 2.773 -0.834
H14 -0.311 2.773 0.834

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14
C11.40482.42942.82032.42941.40481.40242.15883.40773.90543.40772.15882.04022.0402
C21.40481.39442.42682.78642.41752.43841.08732.14543.41083.87283.40383.28362.5979
C32.42941.39441.39742.40722.78643.71242.15551.08642.16123.39583.87374.45203.9755
C42.82032.42681.39741.39742.42684.22173.41022.15731.08512.15733.41024.74024.7402
C52.42942.78642.40721.39741.39443.71243.87373.39582.16121.08642.15553.97554.4520
C61.40482.41752.78642.42681.39442.43843.40383.87283.41082.14541.08732.59793.2836
N71.40242.43843.71244.22173.71242.43842.66744.58115.30684.58112.66741.01221.0122
H82.15881.08732.15553.41023.87373.40382.66742.46924.30684.96014.30353.61392.4250
H93.40772.14541.08642.15733.39583.87284.58112.46922.49294.30014.96015.39144.6790
H103.90543.41082.16121.08512.16123.41085.30684.30682.49292.49294.30685.80945.8094
H113.40773.87283.39582.15731.08642.14544.58114.96014.30012.49292.46924.67905.3914
H122.15883.40383.87373.41022.15551.08732.66744.30354.96014.30682.46922.42503.6139
H132.04023.28364.45204.74023.97552.59791.01223.61395.39145.80944.67902.42501.6683
H142.04022.59793.97554.74024.45203.28361.01222.42504.67905.80945.39143.61391.6683

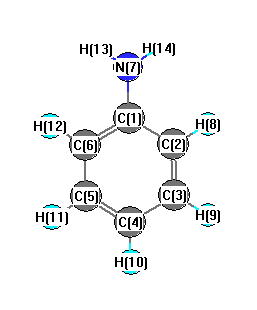
picture of aniline state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.425 C1 C2 H8 119.507
C1 C6 C5 120.425 C1 C6 H12 119.507
C1 N7 H13 114.362 C1 N7 H14 114.362
C2 C1 C6 118.728 C2 C1 N7 120.598
C2 C3 C4 120.745 C2 C3 H9 119.195
C3 C2 H8 120.067 C3 C4 C5 118.932
C3 C4 H10 120.534 C4 C3 H9 120.060
C4 C5 C6 120.745 C4 C5 H11 120.060
C5 C4 H10 120.534 C5 C6 H12 120.067
C6 C1 N7 120.598 C6 C5 H11 119.195
H13 N7 H14 110.994
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.296      
2 C -0.204      
3 C -0.156      
4 C -0.177      
5 C -0.156      
6 C -0.204      
7 N -0.829      
8 H 0.148      
9 H 0.157      
10 H 0.152      
11 H 0.157      
12 H 0.148      
13 H 0.334      
14 H 0.334      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 3.611 -0.124 0.000
y -0.124 13.343 0.000
z 0.000 0.000 11.193


<r2> (average value of r2) Å2
<r2> 192.128
(<r2>)1/2 13.861