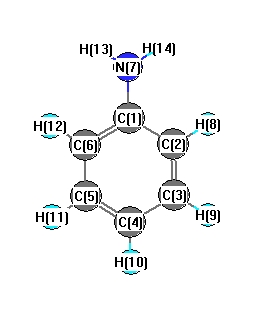
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3592 |
3416 |
14.75 |
|
|
|
| 2 |
A' |
3239 |
3081 |
15.88 |
|
|
|
| 3 |
A' |
3215 |
3058 |
2.49 |
|
|
|
| 4 |
A' |
3197 |
3040 |
18.95 |
|
|
|
| 5 |
A' |
1709 |
1625 |
157.00 |
|
|
|
| 6 |
A' |
1687 |
1605 |
7.65 |
|
|
|
| 7 |
A' |
1563 |
1487 |
62.45 |
|
|
|
| 8 |
A' |
1339 |
1273 |
46.91 |
|
|
|
| 9 |
A' |
1214 |
1154 |
7.40 |
|
|
|
| 10 |
A' |
1068 |
1016 |
2.66 |
|
|
|
| 11 |
A' |
1015 |
966 |
0.44 |
|
|
|
| 12 |
A' |
982 |
934 |
0.23 |
|
|
|
| 13 |
A' |
883 |
840 |
12.47 |
|
|
|
| 14 |
A' |
844 |
803 |
2.22 |
|
|
|
| 15 |
A' |
767 |
729 |
84.37 |
|
|
|
| 16 |
A' |
707 |
673 |
12.72 |
|
|
|
| 17 |
A' |
624 |
593 |
311.18 |
|
|
|
| 18 |
A' |
538 |
511 |
2.28 |
|
|
|
| 19 |
A' |
509 |
484 |
57.69 |
|
|
|
| 20 |
A' |
225 |
214 |
5.53 |
|
|
|
| 21 |
A" |
3696 |
3515 |
11.45 |
|
|
|
| 22 |
A" |
3221 |
3063 |
38.64 |
|
|
|
| 23 |
A" |
3198 |
3041 |
5.43 |
|
|
|
| 24 |
A" |
1671 |
1589 |
5.15 |
|
|
|
| 25 |
A" |
1528 |
1453 |
1.99 |
|
|
|
| 26 |
A" |
1399 |
1331 |
9.30 |
|
|
|
| 27 |
A" |
1375 |
1308 |
0.00 |
|
|
|
| 28 |
A" |
1191 |
1132 |
1.50 |
|
|
|
| 29 |
A" |
1155 |
1098 |
3.67 |
|
|
|
| 30 |
A" |
1088 |
1035 |
3.00 |
|
|
|
| 31 |
A" |
956 |
909 |
0.00 |
|
|
|
| 32 |
A" |
831 |
790 |
0.12 |
|
|
|
| 33 |
A" |
635 |
604 |
0.30 |
|
|
|
| 34 |
A" |
418 |
398 |
0.36 |
|
|
|
| 35 |
A" |
384 |
365 |
0.20 |
|
|
|
| 36 |
A" |
293 |
279 |
22.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25977.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 24704.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.291 |
|
|
|
| 2 |
C |
-0.204 |
|
|
|
| 3 |
C |
-0.171 |
|
|
|
| 4 |
C |
-0.177 |
|
|
|
| 5 |
C |
-0.171 |
|
|
|
| 6 |
C |
-0.204 |
|
|
|
| 7 |
N |
-0.822 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.152 |
|
|
|
| 13 |
H |
0.339 |
|
|
|
| 14 |
H |
0.339 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.171 |
1.303 |
0.000 |
1.752 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.508 |
-3.586 |
0.000 |
| y |
-3.586 |
-33.769 |
0.000 |
| z |
0.000 |
0.000 |
-35.771 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-10.738 |
-3.586 |
0.000 |
| y |
-3.586 |
6.871 |
0.000 |
| z |
0.000 |
0.000 |
3.867 |
|
| Polar |
|---|
| 3z2-r2 | 7.735 |
|---|
| x2-y2 | -11.739 |
|---|
| xy | -3.586 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.654 |
-0.104 |
0.000 |
| y |
-0.104 |
13.437 |
0.000 |
| z |
0.000 |
0.000 |
11.208 |
<r2> (average value of r
2) Å
2
| <r2> |
190.996 |
| (<r2>)1/2 |
13.820 |