return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5NH2 (aniline)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-286.681766
Energy at 298.15K-286.689621
HF Energy-285.729316
Nuclear repulsion energy271.184005
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3589 3369 9.56      
2 A' 3258 3058 14.55      
3 A' 3235 3037 2.95      
4 A' 3219 3022 14.27      
5 A' 1737 1631 120.00      
6 A' 1715 1609 12.78      
7 A' 1586 1489 59.02      
8 A' 1338 1256 39.26      
9 A' 1239 1163 4.17      
10 A' 1081 1014 2.64      
11 A' 1033 970 0.38      
12 A' 899 844 3.75      
13 A' 855 802 18.52      
14 A' 834 783 0.77      
15 A' 757 710 330.78      
16 A' 717 673 100.64      
17 A' 544 511 2.73      
18 A' 537 504 16.66      
19 A' 480 451 17.97      
20 A' 222 209 5.08      
21 A" 3685 3459 7.58      
22 A" 3242 3043 33.97      
23 A" 3220 3022 3.35      
24 A" 1700 1596 2.43      
25 A" 1551 1456 4.42      
26 A" 1400 1314 0.06      
27 A" 1344 1262 9.22      
28 A" 1212 1138 1.93      
29 A" 1174 1101 6.39      
30 A" 1108 1040 2.39      
31 A" 900 845 0.01      
32 A" 814 764 0.22      
33 A" 644 605 0.23      
34 A" 404 379 0.44      
35 A" 386 362 0.34      
36 A" 239 224 29.37      

Unscaled Zero Point Vibrational Energy (zpe) 25948.3 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 24355.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.18750 0.08643 0.05929

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.003 0.934 0.000
C2 0.007 0.222 1.207
C3 0.007 -1.171 1.202
C4 0.008 -1.878 0.000
C5 0.007 -1.171 -1.202
C6 0.007 0.222 -1.207
N7 0.065 2.340 0.000
H8 0.011 0.765 2.149
H9 0.009 -1.706 2.148
H10 0.008 -2.964 0.000
H11 0.009 -1.706 -2.148
H12 0.011 0.765 -2.149
H13 -0.345 2.756 -0.828
H14 -0.345 2.756 0.828

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 N7 H8 H9 H10 H11 H12 H13 H14
C11.40072.42332.81162.42331.40071.40832.15583.40283.89733.40282.15582.03062.0306
C21.40071.39272.42212.78232.41312.43831.08782.14533.40663.86903.39933.26852.5857
C32.42331.39271.39472.40412.78233.71132.15501.08682.15873.39253.87004.43423.9597
C42.81162.42211.39471.39472.42214.21863.40662.15491.08572.15493.40664.72044.7204
C52.42332.78232.40411.39471.39273.71133.87003.39252.15871.08682.15503.95974.4342
C61.40072.41312.78232.42211.39272.43833.39933.86903.40662.14531.08782.58573.2685
N71.40832.43833.71134.21863.71132.43832.66514.58095.30424.58092.66511.01321.0132
H82.15581.08782.15503.40663.87003.39932.66512.47064.30384.95674.29833.59932.4155
H93.40282.14531.08682.15493.39253.86904.58092.47062.48944.29604.95675.37464.6658
H103.89733.40662.15871.08572.15873.40665.30424.30382.48942.48944.30385.78985.7898
H113.40283.86903.39252.15491.08682.14534.58094.95674.29602.48942.47064.66585.3746
H122.15583.39933.87003.40662.15501.08782.66514.29834.95674.30382.47062.41553.5993
H132.03063.26854.43424.72043.95972.58571.01323.59935.37465.78984.66582.41551.6565
H142.03062.58573.95974.72044.43423.26851.01322.41554.66585.78985.37463.59931.6565

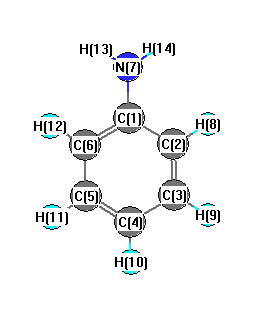
picture of aniline state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.338 C1 C2 H8 119.532
C1 C6 C5 120.338 C1 C6 H12 119.532
C1 N7 H13 112.961 C1 N7 H14 112.961
C2 C1 C6 118.944 C2 C1 N7 120.463
C2 C3 C4 120.664 C2 C3 H9 119.300
C3 C2 H8 120.130 C3 C4 C5 119.045
C3 C4 H10 120.477 C4 C3 H9 120.036
C4 C5 C6 120.664 C4 C5 H11 120.036
C5 C4 H10 120.477 C5 C6 H12 120.130
C6 C1 N7 120.463 C6 C5 H11 119.300
H13 N7 H14 109.659
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability