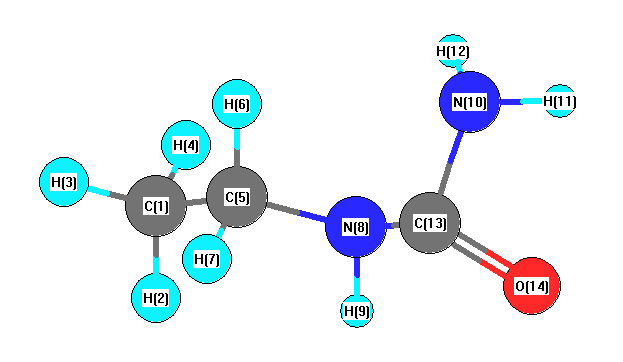
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3732 |
3546 |
33.19 |
|
|
|
| 2 |
A |
3657 |
3475 |
30.45 |
|
|
|
| 3 |
A |
3613 |
3434 |
20.61 |
|
|
|
| 4 |
A |
3171 |
3014 |
18.90 |
|
|
|
| 5 |
A |
3156 |
2999 |
26.36 |
|
|
|
| 6 |
A |
3078 |
2925 |
17.87 |
|
|
|
| 7 |
A |
3065 |
2912 |
32.45 |
|
|
|
| 8 |
A |
3032 |
2881 |
41.01 |
|
|
|
| 9 |
A |
1858 |
1765 |
467.69 |
|
|
|
| 10 |
A |
1664 |
1581 |
167.31 |
|
|
|
| 11 |
A |
1556 |
1479 |
2.31 |
|
|
|
| 12 |
A |
1531 |
1455 |
4.27 |
|
|
|
| 13 |
A |
1521 |
1445 |
8.16 |
|
|
|
| 14 |
A |
1484 |
1410 |
87.00 |
|
|
|
| 15 |
A |
1472 |
1399 |
192.64 |
|
|
|
| 16 |
A |
1440 |
1368 |
20.66 |
|
|
|
| 17 |
A |
1375 |
1307 |
17.13 |
|
|
|
| 18 |
A |
1316 |
1251 |
3.60 |
|
|
|
| 19 |
A |
1230 |
1169 |
20.55 |
|
|
|
| 20 |
A |
1180 |
1121 |
1.53 |
|
|
|
| 21 |
A |
1150 |
1093 |
20.93 |
|
|
|
| 22 |
A |
1090 |
1035 |
14.26 |
|
|
|
| 23 |
A |
1016 |
965 |
1.62 |
|
|
|
| 24 |
A |
916 |
871 |
2.27 |
|
|
|
| 25 |
A |
830 |
789 |
0.74 |
|
|
|
| 26 |
A |
787 |
748 |
75.49 |
|
|
|
| 27 |
A |
609 |
578 |
62.58 |
|
|
|
| 28 |
A |
576 |
547 |
108.46 |
|
|
|
| 29 |
A |
556 |
529 |
52.66 |
|
|
|
| 30 |
A |
499 |
474 |
42.71 |
|
|
|
| 31 |
A |
400 |
380 |
110.95 |
|
|
|
| 32 |
A |
365 |
347 |
0.28 |
|
|
|
| 33 |
A |
277 |
263 |
2.91 |
|
|
|
| 34 |
A |
197 |
188 |
2.81 |
|
|
|
| 35 |
A |
115 |
110 |
7.61 |
|
|
|
| 36 |
A |
51 |
49 |
0.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26781.3 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 25450.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.556 |
|
|
|
| 2 |
H |
0.185 |
|
|
|
| 3 |
H |
0.189 |
|
|
|
| 4 |
H |
0.192 |
|
|
|
| 5 |
C |
-0.168 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.188 |
|
|
|
| 8 |
N |
-0.641 |
|
|
|
| 9 |
H |
0.357 |
|
|
|
| 10 |
N |
-0.802 |
|
|
|
| 11 |
H |
0.362 |
|
|
|
| 12 |
H |
0.346 |
|
|
|
| 13 |
C |
0.714 |
|
|
|
| 14 |
O |
-0.533 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.051 |
2.596 |
-0.655 |
4.059 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.631 |
6.862 |
-1.744 |
| y |
6.862 |
-34.429 |
-1.932 |
| z |
-1.744 |
-1.932 |
-37.628 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.602 |
6.862 |
-1.744 |
| y |
6.862 |
3.201 |
-1.932 |
| z |
-1.744 |
-1.932 |
-1.598 |
|
| Polar |
|---|
| 3z2-r2 | -3.196 |
|---|
| x2-y2 | -3.202 |
|---|
| xy | 6.862 |
|---|
| xz | -1.744 |
|---|
| yz | -1.932 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.845 |
-0.185 |
0.164 |
| y |
-0.185 |
7.332 |
-0.104 |
| z |
0.164 |
-0.104 |
4.863 |
<r2> (average value of r
2) Å
2
| <r2> |
201.247 |
| (<r2>)1/2 |
14.186 |