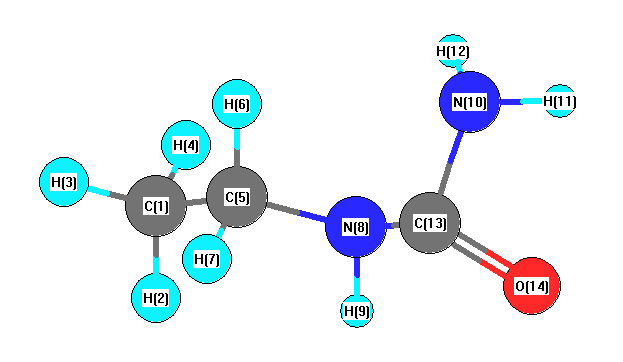
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3727 |
3545 |
32.61 |
|
|
|
| 2 |
A |
3651 |
3472 |
29.29 |
|
|
|
| 3 |
A |
3609 |
3432 |
19.56 |
|
|
|
| 4 |
A |
3169 |
3014 |
19.87 |
|
|
|
| 5 |
A |
3153 |
2999 |
27.68 |
|
|
|
| 6 |
A |
3076 |
2925 |
18.51 |
|
|
|
| 7 |
A |
3062 |
2912 |
33.67 |
|
|
|
| 8 |
A |
3030 |
2881 |
41.98 |
|
|
|
| 9 |
A |
1855 |
1764 |
465.80 |
|
|
|
| 10 |
A |
1663 |
1582 |
166.20 |
|
|
|
| 11 |
A |
1556 |
1480 |
2.38 |
|
|
|
| 12 |
A |
1531 |
1456 |
4.35 |
|
|
|
| 13 |
A |
1521 |
1447 |
7.98 |
|
|
|
| 14 |
A |
1483 |
1410 |
72.04 |
|
|
|
| 15 |
A |
1471 |
1399 |
205.85 |
|
|
|
| 16 |
A |
1440 |
1369 |
20.89 |
|
|
|
| 17 |
A |
1375 |
1308 |
17.85 |
|
|
|
| 18 |
A |
1316 |
1251 |
3.65 |
|
|
|
| 19 |
A |
1228 |
1168 |
20.34 |
|
|
|
| 20 |
A |
1180 |
1122 |
1.55 |
|
|
|
| 21 |
A |
1149 |
1093 |
21.82 |
|
|
|
| 22 |
A |
1089 |
1035 |
14.23 |
|
|
|
| 23 |
A |
1015 |
965 |
1.66 |
|
|
|
| 24 |
A |
916 |
871 |
2.15 |
|
|
|
| 25 |
A |
830 |
789 |
0.71 |
|
|
|
| 26 |
A |
786 |
748 |
75.14 |
|
|
|
| 27 |
A |
609 |
579 |
61.88 |
|
|
|
| 28 |
A |
575 |
547 |
105.21 |
|
|
|
| 29 |
A |
556 |
529 |
56.14 |
|
|
|
| 30 |
A |
499 |
474 |
42.79 |
|
|
|
| 31 |
A |
400 |
380 |
110.57 |
|
|
|
| 32 |
A |
365 |
347 |
0.26 |
|
|
|
| 33 |
A |
277 |
263 |
2.88 |
|
|
|
| 34 |
A |
197 |
188 |
2.79 |
|
|
|
| 35 |
A |
115 |
110 |
7.64 |
|
|
|
| 36 |
A |
49 |
47 |
0.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26760.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 25449.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.549 |
|
|
|
| 2 |
H |
0.183 |
|
|
|
| 3 |
H |
0.187 |
|
|
|
| 4 |
H |
0.189 |
|
|
|
| 5 |
C |
-0.164 |
|
|
|
| 6 |
H |
0.164 |
|
|
|
| 7 |
H |
0.185 |
|
|
|
| 8 |
N |
-0.634 |
|
|
|
| 9 |
H |
0.355 |
|
|
|
| 10 |
N |
-0.797 |
|
|
|
| 11 |
H |
0.361 |
|
|
|
| 12 |
H |
0.344 |
|
|
|
| 13 |
C |
0.709 |
|
|
|
| 14 |
O |
-0.532 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.046 |
2.595 |
-0.650 |
4.054 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.647 |
6.866 |
-1.744 |
| y |
6.866 |
-34.440 |
-1.926 |
| z |
-1.744 |
-1.926 |
-37.627 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.614 |
6.866 |
-1.744 |
| y |
6.866 |
3.197 |
-1.926 |
| z |
-1.744 |
-1.926 |
-1.584 |
|
| Polar |
|---|
| 3z2-r2 | -3.167 |
|---|
| x2-y2 | -3.208 |
|---|
| xy | 6.866 |
|---|
| xz | -1.744 |
|---|
| yz | -1.926 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.861 |
-0.183 |
0.164 |
| y |
-0.183 |
7.346 |
-0.106 |
| z |
0.164 |
-0.106 |
4.868 |
<r2> (average value of r
2) Å
2
| <r2> |
201.209 |
| (<r2>)1/2 |
14.185 |