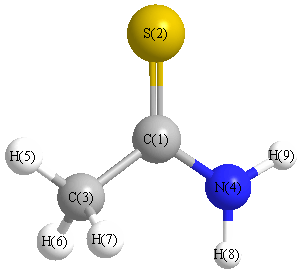
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3608 |
3548 |
26.56 |
|
|
|
| 2 |
A |
3478 |
3420 |
28.64 |
|
|
|
| 3 |
A |
3100 |
3049 |
3.25 |
|
|
|
| 4 |
A |
3063 |
3012 |
14.00 |
|
|
|
| 5 |
A |
2986 |
2937 |
21.02 |
|
|
|
| 6 |
A |
1612 |
1586 |
161.25 |
|
|
|
| 7 |
A |
1480 |
1455 |
11.24 |
|
|
|
| 8 |
A |
1448 |
1424 |
10.20 |
|
|
|
| 9 |
A |
1379 |
1357 |
114.50 |
|
|
|
| 10 |
A |
1377 |
1354 |
61.84 |
|
|
|
| 11 |
A |
1302 |
1281 |
67.48 |
|
|
|
| 12 |
A |
1016 |
999 |
1.39 |
|
|
|
| 13 |
A |
991 |
975 |
5.60 |
|
|
|
| 14 |
A |
968 |
952 |
18.61 |
|
|
|
| 15 |
A |
732 |
720 |
6.12 |
|
|
|
| 16 |
A |
627 |
617 |
15.53 |
|
|
|
| 17 |
A |
487 |
479 |
10.17 |
|
|
|
| 18 |
A |
425 |
418 |
1.83 |
|
|
|
| 19 |
A |
361 |
355 |
2.62 |
|
|
|
| 20 |
A |
285 |
280 |
192.50 |
|
|
|
| 21 |
A |
30 |
29 |
2.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15377.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 15123.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.159 |
|
|
|
| 2 |
S |
-0.240 |
|
|
|
| 3 |
C |
-0.542 |
|
|
|
| 4 |
N |
-0.653 |
|
|
|
| 5 |
H |
0.213 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
H |
0.210 |
|
|
|
| 8 |
H |
0.340 |
|
|
|
| 9 |
H |
0.357 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.486 |
1.528 |
0.064 |
4.739 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.326 |
1.174 |
0.090 |
| y |
1.174 |
-27.173 |
0.015 |
| z |
0.090 |
0.015 |
-33.226 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.126 |
1.174 |
0.090 |
| y |
1.174 |
5.103 |
0.015 |
| z |
0.090 |
0.015 |
-3.977 |
|
| Polar |
|---|
| 3z2-r2 | -7.954 |
|---|
| x2-y2 | -4.153 |
|---|
| xy | 1.174 |
|---|
| xz | 0.090 |
|---|
| yz | 0.015 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.756 |
0.788 |
-0.015 |
| y |
0.788 |
6.725 |
-0.006 |
| z |
-0.015 |
-0.006 |
3.866 |
<r2> (average value of r
2) Å
2
| <r2> |
109.471 |
| (<r2>)1/2 |
10.463 |