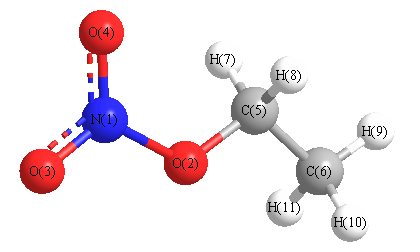
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3176 |
3018 |
12.07 |
|
|
|
| 2 |
A' |
3104 |
2949 |
10.34 |
|
|
|
| 3 |
A' |
3091 |
2938 |
9.98 |
|
|
|
| 4 |
A' |
1821 |
1731 |
381.70 |
|
|
|
| 5 |
A' |
1545 |
1468 |
2.27 |
|
|
|
| 6 |
A' |
1524 |
1448 |
5.66 |
|
|
|
| 7 |
A' |
1455 |
1383 |
22.33 |
|
|
|
| 8 |
A' |
1430 |
1359 |
108.63 |
|
|
|
| 9 |
A' |
1368 |
1300 |
166.95 |
|
|
|
| 10 |
A' |
1168 |
1110 |
20.39 |
|
|
|
| 11 |
A' |
1079 |
1025 |
73.05 |
|
|
|
| 12 |
A' |
967 |
919 |
143.06 |
|
|
|
| 13 |
A' |
918 |
872 |
107.82 |
|
|
|
| 14 |
A' |
740 |
703 |
6.50 |
|
|
|
| 15 |
A' |
590 |
561 |
1.70 |
|
|
|
| 16 |
A' |
389 |
369 |
1.89 |
|
|
|
| 17 |
A' |
231 |
220 |
0.41 |
|
|
|
| 18 |
A" |
3185 |
3027 |
21.39 |
|
|
|
| 19 |
A" |
3155 |
2998 |
6.07 |
|
|
|
| 20 |
A" |
1512 |
1437 |
8.13 |
|
|
|
| 21 |
A" |
1305 |
1240 |
0.50 |
|
|
|
| 22 |
A" |
1193 |
1134 |
5.03 |
|
|
|
| 23 |
A" |
832 |
790 |
0.04 |
|
|
|
| 24 |
A" |
796 |
756 |
18.28 |
|
|
|
| 25 |
A" |
262 |
249 |
0.40 |
|
|
|
| 26 |
A" |
137 |
131 |
0.90 |
|
|
|
| 27 |
A" |
88 |
83 |
0.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18528.8 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 17607.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
0.776 |
|
|
|
| 2 |
O |
-0.380 |
|
|
|
| 3 |
O |
-0.367 |
|
|
|
| 4 |
O |
-0.381 |
|
|
|
| 5 |
C |
-0.097 |
|
|
|
| 6 |
C |
-0.551 |
|
|
|
| 7 |
H |
0.201 |
|
|
|
| 8 |
H |
0.201 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
H |
0.202 |
|
|
|
| 11 |
H |
0.202 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.470 |
0.246 |
0.000 |
3.479 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.018 |
-0.127 |
0.000 |
| y |
-0.127 |
-37.336 |
0.000 |
| z |
0.000 |
0.000 |
-32.795 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.047 |
-0.127 |
0.000 |
| y |
-0.127 |
-3.429 |
0.000 |
| z |
0.000 |
0.000 |
3.382 |
|
| Polar |
|---|
| 3z2-r2 | 6.763 |
|---|
| x2-y2 | 2.318 |
|---|
| xy | -0.127 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.857 |
-0.099 |
0.000 |
| y |
-0.099 |
6.028 |
0.000 |
| z |
0.000 |
0.000 |
4.182 |
<r2> (average value of r
2) Å
2
| <r2> |
176.092 |
| (<r2>)1/2 |
13.270 |