return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8OS (s-Ethyl thioacetate)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-630.232598
Energy at 298.15K-630.241050
HF Energy-629.957874
Nuclear repulsion energy292.017373
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3202 3040 6.08      
2 A 3179 3018 26.35      
3 A 3166 3005 5.30      
4 A 3166 3005 19.95      
5 A 3155 2995 0.60      
6 A 3106 2948 19.17      
7 A 3093 2936 0.86      
8 A 3091 2934 21.09      
9 A 1778 1687 254.93      
10 A 1553 1474 2.61      
11 A 1544 1466 8.65      
12 A 1536 1458 2.29      
13 A 1524 1446 13.60      
14 A 1511 1435 20.48      
15 A 1462 1388 3.50      
16 A 1427 1355 18.79      
17 A 1344 1276 28.14      
18 A 1308 1241 0.01      
19 A 1174 1114 151.15      
20 A 1101 1045 5.99      
21 A 1082 1027 0.16      
22 A 1051 997 2.10      
23 A 1016 964 3.29      
24 A 967 918 49.30      
25 A 815 774 4.72      
26 A 708 672 0.25      
27 A 647 614 62.69      
28 A 514 488 2.75      
29 A 450 427 3.29      
30 A 378 359 1.42      
31 A 309 293 0.99      
32 A 261 248 0.10      
33 A 162 154 1.06      
34 A 149 141 0.10      
35 A 86 81 0.27      
36 A 57 54 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 25534.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24237.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.21266 0.05412 0.04421

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -2.057 -0.961 -0.001
C2 -1.150 -0.118 -0.001
S3 0.199 -0.570 0.001
H4 -1.146 1.790 0.886
H5 -2.623 1.369 -0.001
H6 -1.143 1.793 -0.885
C7 -1.534 1.292 0.002
H8 1.113 1.176 -0.892
H9 1.111 1.174 0.891
C10 1.220 0.555 -0.002
H11 2.771 -0.666 0.889
H12 3.377 0.753 0.000
H13 2.773 -0.664 -0.888
C14 2.635 -0.048 0.000

Atom - Atom Distances (Å)
  O1 C2 S3 H4 H5 H6 C7 H8 H9 C10 H11 H12 H13 C14
O11.23872.29013.03092.39773.03342.31263.92533.92343.61094.91805.69794.92014.7804
C21.23871.42292.10402.09322.10591.46112.75442.75242.46354.05704.60964.05893.7856
S32.29011.42292.85763.42452.85932.54422.16382.16031.51932.72193.44212.72472.4913
H43.03092.10402.85761.77361.77181.08722.93942.34002.81334.62294.72444.95294.2968
H52.39772.09323.42451.77361.77531.09153.84523.84433.92855.83296.03155.83425.4458
H63.03342.10592.85931.77181.77531.09122.33882.93662.81074.95124.72234.62344.2956
C72.31261.46112.54421.08721.09151.09122.79582.79362.85134.81154.94074.81354.3794
H83.92532.75442.16382.93943.84522.33882.79581.78311.09033.05202.47032.47862.1478
H93.92342.75242.16032.34003.84432.93662.79361.78311.09252.47742.47083.05052.1470
C103.61092.46351.51932.81333.92852.81072.85131.09031.09252.16532.16582.16371.5381
H114.91804.05702.72194.62295.83294.95124.81153.05202.47742.16531.78081.77761.0913
H125.69794.60963.44214.72446.03154.72234.94072.47032.47082.16581.78081.77761.0915
H134.92014.05892.72474.95295.83424.62344.81352.47863.05052.16371.77761.77761.0897
C144.78043.78562.49134.29685.44584.29564.37942.14782.14701.53811.09131.09151.0897

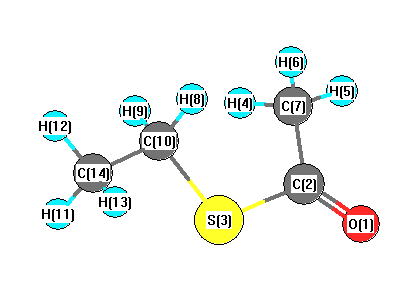
picture of s-Ethyl thioacetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 S3 118.558 O1 C2 C7 117.633
C2 S3 C10 113.665 C2 C7 H4 110.449
C2 C7 H5 109.319 C2 C7 H6 110.357
S3 C2 C7 123.808 S3 C10 H8 110.955
S3 C10 H9 110.537 S3 C10 C14 109.139
H4 C7 H5 108.990 H4 C7 H6 108.847
H5 C7 H6 108.845 H8 C10 H9 109.539
H8 C10 C14 108.393 H9 C10 C14 108.207
C10 C14 H11 109.708 C10 C14 H12 109.736
C10 C14 H13 109.671 H11 C14 H12 109.347
H11 C14 H13 109.189 H12 C14 H13 109.174
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 O -0.460      
2 C 0.313      
3 S 0.145      
4 H 0.187      
5 H 0.206      
6 H 0.187      
7 C -0.545      
8 H 0.186      
9 H 0.186      
10 C -0.453      
11 H 0.179      
12 H 0.168      
13 H 0.179      
14 C -0.477      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.474 -0.160 0.000
y -0.160 8.650 0.000
z 0.000 0.000 6.410


<r2> (average value of r2) Å2
<r2> 252.630
(<r2>)1/2 15.894