return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1-pentyne)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-194.624562
Energy at 298.15K-194.631579
HF Energy-193.931743
Nuclear repulsion energy153.253764
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3492 3325 44.55      
2 A' 3141 2991 33.64      
3 A' 3074 2927 27.54      
4 A' 3061 2915 30.11      
5 A' 3055 2909 7.49      
6 A' 2226 2120 0.51      
7 A' 1559 1485 4.91      
8 A' 1545 1471 0.89      
9 A' 1534 1461 1.03      
10 A' 1467 1397 1.92      
11 A' 1436 1368 1.93      
12 A' 1339 1275 5.83      
13 A' 1147 1092 1.64      
14 A' 1089 1037 0.08      
15 A' 990 942 2.35      
16 A' 902 859 3.13      
17 A' 560 533 49.50      
18 A' 474 451 2.23      
19 A' 341 325 0.02      
20 A' 148 141 0.06      
21 A" 3137 2987 61.17      
22 A" 3113 2965 6.48      
23 A" 3089 2942 4.08      
24 A" 1549 1475 6.93      
25 A" 1357 1292 0.01      
26 A" 1296 1234 0.08      
27 A" 1155 1099 0.02      
28 A" 899 856 0.78      
29 A" 758 722 1.39      
30 A" 541 515 52.37      
31 A" 303 289 0.18      
32 A" 244 233 0.01      
33 A" 100 95 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 25058.6 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 23863.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.78443 0.07360 0.06992

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.653 0.218 0.000
C2 1.456 0.416 0.000
C3 0.000 0.638 0.000
C4 -0.802 -0.673 0.000
C5 -2.311 -0.421 0.000
H6 3.708 0.046 0.000
H7 -0.274 1.238 0.880
H8 -0.274 1.238 -0.880
H9 -0.518 -1.265 0.880
H10 -0.518 -1.265 -0.880
H11 -2.869 -1.365 0.000
H12 -2.615 0.150 0.887
H13 -2.615 0.150 -0.887

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.21392.68643.56855.00491.06883.22243.22243.60993.60995.74445.34305.3430
C21.21391.47252.50643.85802.28282.10762.10762.73752.73754.67654.17474.1747
C32.68641.47251.53682.54183.75521.09971.09972.15972.15973.49882.80442.8044
C43.56852.50641.53681.52934.56732.16862.16861.09831.09832.17932.17952.1795
C55.00493.85802.54181.52936.03692.77002.77002.16802.16801.09681.09791.0979
H61.06882.28283.75524.56736.03694.24894.24894.51164.51166.72646.38606.3860
H73.22242.10761.09972.16862.77004.24891.76072.51413.06933.77892.58143.1286
H83.22242.10761.09972.16862.77004.24891.76073.06932.51413.77893.12862.5814
H93.60992.73752.15971.09832.16804.51162.51413.06931.76062.51192.52943.0858
H103.60992.73752.15971.09832.16804.51163.06932.51411.76062.51193.08582.5294
H115.74444.67653.49882.17931.09686.72643.77893.77892.51192.51191.77381.7738
H125.34304.17472.80442.17951.09796.38602.58143.12862.52943.08581.77381.7744
H135.34304.17472.80442.17951.09796.38603.12862.58143.08582.52941.77381.7744

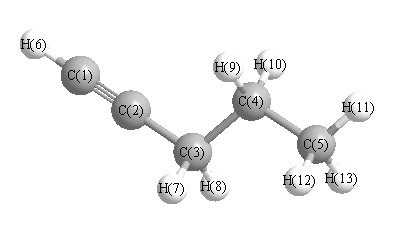
picture of 1-pentyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.325 C2 C1 H6 179.902
C2 C3 C4 112.775 C2 C3 H7 109.189
C2 C3 H8 109.189 C3 C4 C5 111.988
C3 C4 H9 108.950 C3 C4 H10 108.950
C4 C3 H7 109.561 C4 C3 H8 109.561
C4 C5 H11 111.100 C4 C5 H12 111.045
C4 C5 H13 111.045 C5 C4 H9 110.117
C5 C4 H10 110.117 H7 C3 H8 106.361
H9 C4 H10 106.558 H11 C5 H12 107.841
H11 C5 H13 107.841 H12 C5 H13 107.817
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability