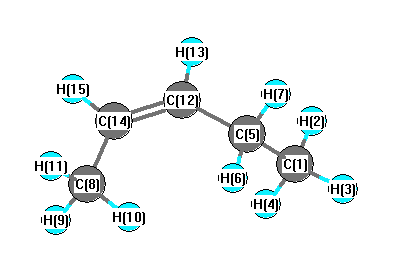
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3166 |
3029 |
63.78 |
|
|
|
| 2 |
A |
3149 |
3012 |
10.16 |
|
|
|
| 3 |
A |
3139 |
3003 |
39.57 |
|
|
|
| 4 |
A |
3136 |
3001 |
20.44 |
|
|
|
| 5 |
A |
3130 |
2994 |
24.71 |
|
|
|
| 6 |
A |
3100 |
2966 |
7.29 |
|
|
|
| 7 |
A |
3095 |
2961 |
22.91 |
|
|
|
| 8 |
A |
3055 |
2923 |
33.31 |
|
|
|
| 9 |
A |
3044 |
2912 |
30.38 |
|
|
|
| 10 |
A |
3034 |
2902 |
31.27 |
|
|
|
| 11 |
A |
1759 |
1683 |
2.14 |
|
|
|
| 12 |
A |
1532 |
1466 |
6.36 |
|
|
|
| 13 |
A |
1523 |
1457 |
6.80 |
|
|
|
| 14 |
A |
1520 |
1454 |
3.99 |
|
|
|
| 15 |
A |
1508 |
1443 |
6.19 |
|
|
|
| 16 |
A |
1505 |
1439 |
0.99 |
|
|
|
| 17 |
A |
1456 |
1393 |
7.72 |
|
|
|
| 18 |
A |
1430 |
1368 |
1.55 |
|
|
|
| 19 |
A |
1425 |
1364 |
1.90 |
|
|
|
| 20 |
A |
1348 |
1290 |
7.00 |
|
|
|
| 21 |
A |
1314 |
1257 |
0.26 |
|
|
|
| 22 |
A |
1290 |
1234 |
0.30 |
|
|
|
| 23 |
A |
1182 |
1131 |
0.84 |
|
|
|
| 24 |
A |
1102 |
1055 |
3.64 |
|
|
|
| 25 |
A |
1070 |
1024 |
0.36 |
|
|
|
| 26 |
A |
1059 |
1013 |
0.14 |
|
|
|
| 27 |
A |
1049 |
1004 |
3.39 |
|
|
|
| 28 |
A |
1012 |
969 |
0.27 |
|
|
|
| 29 |
A |
953 |
912 |
11.78 |
|
|
|
| 30 |
A |
877 |
839 |
1.81 |
|
|
|
| 31 |
A |
798 |
764 |
3.53 |
|
|
|
| 32 |
A |
725 |
693 |
32.02 |
|
|
|
| 33 |
A |
571 |
547 |
7.14 |
|
|
|
| 34 |
A |
471 |
451 |
1.42 |
|
|
|
| 35 |
A |
304 |
290 |
0.02 |
|
|
|
| 36 |
A |
258 |
247 |
0.02 |
|
|
|
| 37 |
A |
219 |
210 |
0.01 |
|
|
|
| 38 |
A |
143 |
137 |
0.37 |
|
|
|
| 39 |
A |
57 |
54 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30254.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28944.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.506 |
|
|
-0.380 |
| 2 |
H |
0.166 |
|
|
0.094 |
| 3 |
H |
0.166 |
|
|
0.070 |
| 4 |
H |
0.174 |
|
|
0.106 |
| 5 |
C |
-0.356 |
|
|
0.380 |
| 6 |
H |
0.165 |
|
|
-0.036 |
| 7 |
H |
0.170 |
|
|
-0.029 |
| 8 |
C |
-0.556 |
|
|
-0.241 |
| 9 |
H |
0.178 |
|
|
0.083 |
| 10 |
H |
0.175 |
|
|
0.094 |
| 11 |
H |
0.179 |
|
|
0.077 |
| 12 |
C |
-0.116 |
|
|
-0.450 |
| 13 |
H |
0.145 |
|
|
0.160 |
| 14 |
C |
-0.131 |
|
|
-0.023 |
| 15 |
H |
0.148 |
|
|
0.093 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.024 |
-0.194 |
0.035 |
0.199 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.041 |
-0.220 |
0.019 |
0.225 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.388 |
0.312 |
0.832 |
| y |
0.312 |
-31.992 |
-0.331 |
| z |
0.832 |
-0.331 |
-33.851 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.534 |
0.312 |
0.832 |
| y |
0.312 |
0.628 |
-0.331 |
| z |
0.832 |
-0.331 |
-2.161 |
|
| Polar |
|---|
| 3z2-r2 | -4.323 |
|---|
| x2-y2 | 0.604 |
|---|
| xy | 0.312 |
|---|
| xz | 0.832 |
|---|
| yz | -0.331 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
165.637 |
| (<r2>)1/2 |
12.870 |