return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (Z)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-196.271023
Energy at 298.15K-196.281090
HF Energy-196.056723
Nuclear repulsion energy170.990934
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3188 3026 63.22      
2 A 3171 3010 8.16      
3 A 3161 3000 39.19      
4 A 3157 2997 22.86      
5 A 3151 2991 26.09      
6 A 3123 2965 7.66      
7 A 3121 2962 24.35      
8 A 3078 2922 30.63      
9 A 3070 2914 26.78      
10 A 3061 2906 31.13      
11 A 1753 1664 1.51      
12 A 1558 1479 5.51      
13 A 1549 1470 5.52      
14 A 1547 1468 2.50      
15 A 1535 1457 4.87      
16 A 1532 1455 1.04      
17 A 1477 1402 5.44      
18 A 1456 1382 1.09      
19 A 1450 1376 2.08      
20 A 1371 1302 8.17      
21 A 1330 1262 0.16      
22 A 1309 1242 0.18      
23 A 1197 1136 0.74      
24 A 1118 1062 3.56      
25 A 1089 1034 0.50      
26 A 1075 1020 0.06      
27 A 1054 1000 2.76      
28 A 1013 962 0.22      
29 A 959 910 8.93      
30 A 881 836 1.29      
31 A 809 768 2.91      
32 A 732 695 28.81      
33 A 574 545 6.03      
34 A 475 451 1.16      
35 A 310 294 0.03      
36 A 261 248 0.02      
37 A 225 214 0.01      
38 A 135 128 0.29      
39 A 56 53 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 30555.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29002.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.36891 0.08566 0.07836

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.156 -0.275 -0.421
H2 2.636 0.695 -0.519
H3 2.900 -1.000 -0.086
H4 1.776 -0.589 -1.392
C5 1.000 -0.191 0.585
H6 0.553 -1.168 0.680
H7 1.391 0.109 1.554
C8 -2.051 -0.646 -0.080
H9 -2.455 -0.886 -1.066
H10 -1.447 -1.482 0.249
H11 -2.888 -0.530 0.610
C12 0.013 0.792 0.135
H13 0.365 1.751 0.022
C14 -1.288 0.598 -0.150
H15 -1.807 1.426 -0.456

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 H11 C12 H13 C14 H15
C11.08681.09221.08881.53482.14002.15294.23674.69533.85835.15462.45692.73973.56314.3123
H21.08681.76901.77542.16373.04122.48854.89525.35924.69105.77032.70502.56263.94334.5038
H31.09221.76901.77212.17172.47492.48914.96405.44534.38675.84913.40483.74254.48355.3085
H41.08881.77541.77212.16082.47493.05244.04574.25343.72515.07582.71013.07653.51274.2156
C51.53482.16372.17172.16081.07841.08803.15543.89152.78713.90281.46312.11912.52933.4023
H62.14003.04122.47492.47491.07841.75982.76253.48942.07003.50062.10432.99802.68283.6861
H72.15292.48852.48913.05241.08801.75983.88514.75953.50644.42912.09302.46983.21364.0012
C84.23674.89524.96404.04573.15542.76253.88511.09221.08321.09132.52513.40501.46072.1194
H94.69535.35925.44534.25343.89153.48944.75951.09221.76081.76763.21754.01152.09832.4772
H103.85834.69104.38673.72512.78712.07003.50641.08321.76081.76522.70543.71372.12433.0140
H115.15465.77035.84915.07583.90283.50064.42911.09131.76761.76523.22374.01652.09982.4756
C122.45692.70503.40482.71011.46312.10432.09302.52513.21752.70543.22371.02781.34682.0165
H132.73972.56263.74253.07652.11912.99802.46983.40504.01153.71374.01651.02782.02322.2480
C143.56313.94334.48353.51272.52932.68283.21361.46072.09832.12432.09981.34682.02321.0237
H154.31234.50385.30854.21563.40233.68614.00122.11942.47723.01402.47562.01652.24801.0237

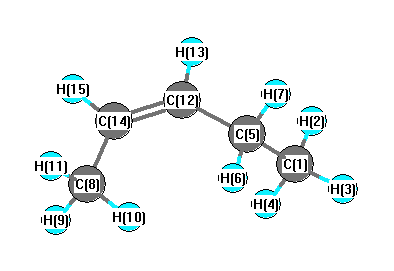
picture of 2-Pentene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 108.697 C1 C5 H7 109.158
C1 C5 C12 110.053 H2 C1 H3 108.556
H2 C1 H4 109.386 H2 C1 C5 110.076
H3 C1 H4 108.684 H3 C1 C5 110.382
H4 C1 C5 109.725 C5 C12 H13 115.470
C5 C12 C14 128.311 H6 C5 H7 108.645
H6 C5 C12 110.876 H7 C5 C12 109.377
C8 C14 C12 128.116 C8 C14 H15 115.983
H9 C8 H10 108.081 H9 C8 H11 108.105
H9 C8 C14 109.713 H10 C8 H11 108.541
H10 C8 C14 112.401 H11 C8 C14 109.888
C12 C14 H15 115.901 H13 C12 C14 116.219
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.457      
2 H 0.150      
3 H 0.150      
4 H 0.157      
5 C -0.318      
6 H 0.152      
7 H 0.153      
8 C -0.502      
9 H 0.161      
10 H 0.162      
11 H 0.162      
12 C -0.121      
13 H 0.139      
14 C -0.130      
15 H 0.142      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.429 0.315 0.674
y 0.315 7.208 -0.176
z 0.674 -0.176 5.928


<r2> (average value of r2) Å2
<r2> 165.326
(<r2>)1/2 12.858