Jump to
S1C2
Vibrational Frequencies calculated at MP4=FULL/6-31G*
Geometric Data calculated at MP4=FULL/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4=FULL/6-31G*
| | hartrees |
|---|
| Energy at 0K | -553.155137 |
| Energy at 298.15K | -553.160575 |
| HF Energy | -552.416289 |
| Nuclear repulsion energy | 201.489517 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3268 |
3142 |
|
|
|
|
| 2 |
A |
3264 |
3139 |
|
|
|
|
| 3 |
A |
3197 |
3074 |
|
|
|
|
| 4 |
A |
3187 |
3064 |
|
|
|
|
| 5 |
A |
3176 |
3054 |
|
|
|
|
| 6 |
A |
3170 |
3048 |
|
|
|
|
| 7 |
A |
1663 |
1599 |
|
|
|
|
| 8 |
A |
1654 |
1591 |
|
|
|
|
| 9 |
A |
1449 |
1393 |
|
|
|
|
| 10 |
A |
1445 |
1390 |
|
|
|
|
| 11 |
A |
1324 |
1274 |
|
|
|
|
| 12 |
A |
1318 |
1267 |
|
|
|
|
| 13 |
A |
1080 |
1039 |
|
|
|
|
| 14 |
A |
1054 |
1014 |
|
|
|
|
| 15 |
A |
1000 |
962 |
|
|
|
|
| 16 |
A |
987 |
949 |
|
|
|
|
| 17 |
A |
912 |
877 |
|
|
|
|
| 18 |
A |
877 |
844 |
|
|
|
|
| 19 |
A |
757 |
728 |
|
|
|
|
| 20 |
A |
709 |
682 |
|
|
|
|
| 21 |
A |
633 |
609 |
|
|
|
|
| 22 |
A |
606 |
582 |
|
|
|
|
| 23 |
A |
461 |
443 |
|
|
|
|
| 24 |
A |
374 |
360 |
|
|
|
|
| 25 |
A |
229 |
220 |
|
|
|
|
| 26 |
A |
142 |
137 |
|
|
|
|
| 27 |
A |
85 |
82 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19010.3 cm
-1
Scaled (by 0.9616) Zero Point Vibrational Energy (zpe) 18280.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.008 |
-1.001 |
-0.007 |
| C2 |
1.075 |
0.351 |
0.397 |
| C3 |
-1.562 |
-0.185 |
-0.023 |
| C4 |
2.244 |
0.571 |
-0.226 |
| C5 |
-1.788 |
1.136 |
-0.130 |
| H6 |
0.756 |
0.956 |
1.247 |
| H7 |
-2.390 |
-0.887 |
0.075 |
| H8 |
2.922 |
1.349 |
0.121 |
| H9 |
2.560 |
-0.022 |
-1.082 |
| H10 |
-0.986 |
1.859 |
-0.256 |
| H11 |
-2.806 |
1.520 |
-0.100 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7689 | 1.7692 | 2.7423 | 2.7941 | 2.4415 | 2.4019 | 3.7456 | 2.9375 | 3.0378 | 3.7792 |
C2 | 1.7689 | | 2.7232 | 1.3429 | 3.0157 | 1.0913 | 3.6936 | 2.1172 | 2.1290 | 2.6362 | 4.0838 | C3 | 1.7692 | 2.7232 | | 3.8857 | 1.3441 | 2.8784 | 1.0904 | 4.7408 | 4.2592 | 2.1362 | 2.1122 | C4 | 2.7423 | 1.3429 | 3.8857 | | 4.0733 | 2.1291 | 4.8676 | 1.0887 | 1.0885 | 3.4776 | 5.1402 | C5 | 2.7941 | 3.0157 | 1.3441 | 4.0733 | | 2.8988 | 2.1205 | 4.7217 | 4.6000 | 1.0879 | 1.0880 | H6 | 2.4415 | 1.0913 | 2.8784 | 2.1291 | 2.8988 | | 3.8297 | 2.4723 | 3.1045 | 2.4721 | 3.8498 | H7 | 2.4019 | 3.6936 | 1.0904 | 4.8676 | 2.1205 | 3.8297 | | 5.7635 | 5.1569 | 3.1023 | 2.4497 | H8 | 3.7456 | 2.1172 | 4.7408 | 1.0887 | 4.7217 | 2.4723 | 5.7635 | | 1.8602 | 3.9587 | 5.7343 | H9 | 2.9375 | 2.1290 | 4.2592 | 1.0885 | 4.6000 | 3.1045 | 5.1569 | 1.8602 | | 4.0982 | 5.6691 | H10 | 3.0378 | 2.6362 | 2.1362 | 3.4776 | 1.0879 | 2.4721 | 3.1023 | 3.9587 | 4.0982 | | 1.8578 | H11 | 3.7792 | 4.0838 | 2.1122 | 5.1402 | 1.0880 | 3.8498 | 2.4497 | 5.7343 | 5.6691 | 1.8578 | |
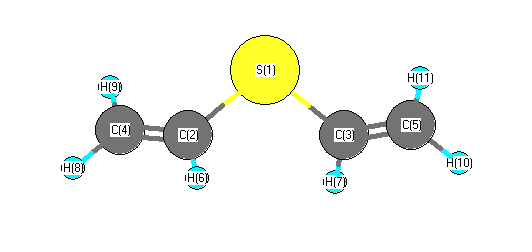 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
123.005 |
|
S1 |
C2 |
H6 |
115.158 |
| S1 |
C3 |
C5 |
127.124 |
|
S1 |
C3 |
H7 |
112.078 |
| C2 |
S1 |
C3 |
100.654 |
|
C2 |
C4 |
H8 |
120.726 |
| C2 |
C4 |
H9 |
121.884 |
|
C3 |
C5 |
H10 |
122.549 |
| C3 |
C5 |
H11 |
120.197 |
|
C4 |
C2 |
H6 |
121.671 |
| C5 |
C3 |
H7 |
120.797 |
|
H8 |
C4 |
H9 |
117.385 |
| H10 |
C5 |
H11 |
117.252 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
