return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (Butane, 1-methoxy-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-272.629487
Energy at 298.15K 
HF Energy-272.355693
Nuclear repulsion energy248.139181
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3174 3013 34.30      
2 A' 3153 2993 38.61      
3 A' 3080 2924 48.40      
4 A' 3076 2920 15.85      
5 A' 3061 2905 23.42      
6 A' 3024 2871 72.86      
7 A' 2996 2843 36.42      
8 A' 1582 1501 0.57      
9 A' 1561 1482 2.49      
10 A' 1555 1476 10.94      
11 A' 1547 1468 0.20      
12 A' 1540 1462 0.32      
13 A' 1521 1443 1.10      
14 A' 1466 1392 35.28      
15 A' 1460 1386 3.07      
16 A' 1423 1350 1.40      
17 A' 1325 1257 2.37      
18 A' 1250 1187 39.71      
19 A' 1184 1124 132.98      
20 A' 1154 1095 24.41      
21 A' 1096 1040 0.51      
22 A' 1056 1003 1.33      
23 A' 999 948 21.04      
24 A' 924 877 6.51      
25 A' 496 471 1.24      
26 A' 387 367 2.90      
27 A' 317 301 0.77      
28 A' 140 133 0.97      
29 A" 3150 2990 61.43      
30 A" 3127 2968 34.39      
31 A" 3092 2935 7.31      
32 A" 3073 2917 66.77      
33 A" 3027 2873 61.02      
34 A" 1550 1472 6.12      
35 A" 1536 1458 4.67      
36 A" 1355 1287 0.05      
37 A" 1343 1274 0.02      
38 A" 1282 1217 1.99      
39 A" 1231 1168 6.22      
40 A" 1196 1136 0.30      
41 A" 977 927 0.01      
42 A" 835 793 0.66      
43 A" 760 721 2.49      
44 A" 257 244 0.77      
45 A" 237 225 2.78      
46 A" 140 133 1.38      
47 A" 104 98 1.28      
48 A" 74 71 1.24      

Unscaled Zero Point Vibrational Energy (zpe) 36945.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35068.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.474 2.780 0.000
C2 1.429 1.252 0.000
C3 -1.548 -2.624 0.000
O4 -1.408 -1.219 0.000
C5 -0.050 -0.815 0.000
C6 0.000 0.704 0.000
H7 -2.617 -2.841 0.000
H8 2.503 3.149 0.000
H9 0.972 3.187 0.883
H10 0.972 3.187 -0.883
H11 1.966 0.871 0.877
H12 1.966 0.871 -0.877
H13 -1.090 -3.077 0.891
H14 -1.090 -3.077 -0.891
H15 -0.542 1.070 -0.879
H16 -0.542 1.070 0.879
H17 0.467 -1.218 -0.886
H18 0.467 -1.218 0.886

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.52916.19124.92953.90442.54606.95171.09371.09441.09442.15822.15826.45576.45572.78632.78634.21664.2166
C21.52914.88673.76222.54121.53035.75462.18052.17592.17591.09711.09715.08725.08722.16622.16622.79432.7943
C36.19124.88671.41142.34853.67021.09097.05246.39486.39485.03275.03271.09981.09983.92773.92772.61192.6119
O44.92953.76221.41141.41662.38372.02245.86365.08525.08524.06484.06482.08512.08512.60052.60052.07392.0739
C53.90442.54122.34851.41661.51953.26984.71504.22344.22342.77062.77062.64492.64492.13682.13681.10261.1026
C62.54601.53033.67022.38371.51954.40603.49932.80882.80882.15952.15954.03514.03511.09581.09582.16712.1671
H76.95175.75461.09092.02243.26984.40607.87997.07047.07045.96225.96221.78381.78384.51314.51313.59583.5958
H81.09372.18057.05245.86364.71503.49937.87991.76841.76842.49992.49997.24397.24393.79123.79124.89894.8989
H91.09442.17596.39485.08524.22342.80887.07041.76841.76672.52063.07476.59496.82953.14352.60304.77354.4333
H101.09442.17596.39485.08524.22342.80887.07041.76841.76673.07472.52066.82956.59492.60303.14354.43334.7735
H112.15821.09715.03274.06482.77062.15955.96222.49992.52063.07471.75504.99275.29673.06892.51653.11762.5706
H122.15821.09715.03274.06482.77062.15955.96222.49993.07472.52061.75505.29674.99272.51653.06892.57063.1176
H136.45575.08721.09982.08512.64494.03511.78387.24396.59496.82954.99275.29671.78244.54244.18323.00712.4255
H146.45575.08721.09982.08512.64494.03511.78387.24396.82956.59495.29674.99271.78244.18324.54242.42553.0071
H152.78632.16623.92772.60052.13681.09584.51313.79123.14352.60303.06892.51654.54244.18321.75822.50023.0607
H162.78632.16623.92772.60052.13681.09584.51313.79122.60303.14352.51653.06894.18324.54241.75823.06072.5002
H174.21662.79432.61192.07391.10262.16713.59584.89894.77354.43333.11762.57063.00712.42552.50023.06071.7728
H184.21662.79432.61192.07391.10262.16713.59584.89894.43334.77352.57063.11762.42553.00713.06072.50021.7728

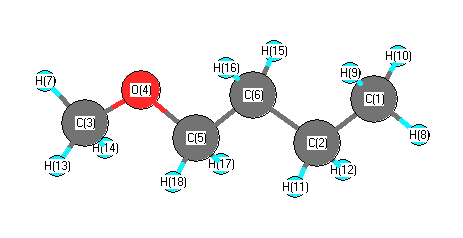
picture of Butane, 1-methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C6 112.645 C1 C2 H11 109.434
C1 C2 H12 109.434 C2 C1 H8 111.394
C2 C1 H9 110.986 C2 C1 H10 110.986
C2 C6 C5 112.862 C2 C6 H15 110.048
C2 C6 H16 110.048 C3 O4 C5 112.283
O4 C3 H7 107.142 O4 C3 H13 111.666
O4 C3 H14 111.666 O4 C5 C6 108.503
O4 C5 H17 110.193 O4 C5 H18 110.193
C5 C6 H15 108.493 C5 C6 H16 108.493
C6 C2 H11 109.448 C6 C2 H12 109.448
C6 C5 H17 110.470 C6 C5 H18 110.470
H7 C3 H13 109.023 H7 C3 H14 109.023
H8 C1 H9 107.839 H8 C1 H10 107.839
H9 C1 H10 107.635 H11 C2 H12 106.226
H13 C3 H14 108.263 H15 C6 H16 106.690
H17 C5 H18 107.015
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.458      
2 C -0.285      
3 C -0.180      
4 O -0.532      
5 C 0.010      
6 C -0.288      
7 H 0.166      
8 H 0.151      
9 H 0.151      
10 H 0.151      
11 H 0.142      
12 H 0.142      
13 H 0.135      
14 H 0.135      
15 H 0.155      
16 H 0.155      
17 H 0.124      
18 H 0.124      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.520 1.241 0.000
y 1.241 9.652 0.000
z 0.000 0.000 7.605


<r2> (average value of r2) Å2
<r2> 293.785
(<r2>)1/2 17.140