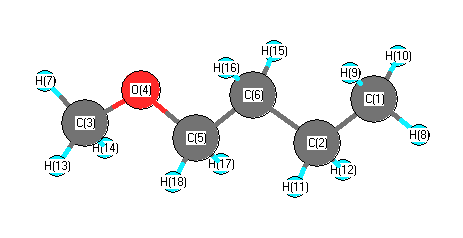
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3005 |
33.19 |
|
|
|
| 2 |
A' |
3149 |
2994 |
35.85 |
|
|
|
| 3 |
A' |
3069 |
2919 |
52.88 |
|
|
|
| 4 |
A' |
3065 |
2915 |
15.46 |
|
|
|
| 5 |
A' |
3050 |
2901 |
20.68 |
|
|
|
| 6 |
A' |
3004 |
2856 |
77.91 |
|
|
|
| 7 |
A' |
2975 |
2829 |
38.10 |
|
|
|
| 8 |
A' |
1561 |
1484 |
1.06 |
|
|
|
| 9 |
A' |
1538 |
1463 |
3.34 |
|
|
|
| 10 |
A' |
1533 |
1458 |
13.21 |
|
|
|
| 11 |
A' |
1523 |
1448 |
0.48 |
|
|
|
| 12 |
A' |
1516 |
1442 |
0.16 |
|
|
|
| 13 |
A' |
1504 |
1430 |
0.12 |
|
|
|
| 14 |
A' |
1453 |
1382 |
34.89 |
|
|
|
| 15 |
A' |
1438 |
1368 |
2.07 |
|
|
|
| 16 |
A' |
1406 |
1337 |
2.13 |
|
|
|
| 17 |
A' |
1308 |
1244 |
2.91 |
|
|
|
| 18 |
A' |
1248 |
1187 |
74.63 |
|
|
|
| 19 |
A' |
1207 |
1148 |
120.24 |
|
|
|
| 20 |
A' |
1150 |
1094 |
8.39 |
|
|
|
| 21 |
A' |
1099 |
1045 |
0.78 |
|
|
|
| 22 |
A' |
1065 |
1013 |
2.48 |
|
|
|
| 23 |
A' |
1016 |
966 |
21.52 |
|
|
|
| 24 |
A' |
924 |
879 |
5.95 |
|
|
|
| 25 |
A' |
494 |
470 |
1.19 |
|
|
|
| 26 |
A' |
387 |
368 |
2.82 |
|
|
|
| 27 |
A' |
315 |
300 |
0.75 |
|
|
|
| 28 |
A' |
138 |
131 |
0.93 |
|
|
|
| 29 |
A" |
3143 |
2989 |
55.99 |
|
|
|
| 30 |
A" |
3117 |
2965 |
34.23 |
|
|
|
| 31 |
A" |
3083 |
2932 |
5.47 |
|
|
|
| 32 |
A" |
3052 |
2902 |
65.46 |
|
|
|
| 33 |
A" |
3005 |
2858 |
66.22 |
|
|
|
| 34 |
A" |
1527 |
1452 |
7.71 |
|
|
|
| 35 |
A" |
1514 |
1439 |
6.26 |
|
|
|
| 36 |
A" |
1340 |
1274 |
0.04 |
|
|
|
| 37 |
A" |
1329 |
1264 |
0.00 |
|
|
|
| 38 |
A" |
1268 |
1206 |
2.46 |
|
|
|
| 39 |
A" |
1216 |
1157 |
5.87 |
|
|
|
| 40 |
A" |
1187 |
1129 |
0.03 |
|
|
|
| 41 |
A" |
964 |
916 |
0.00 |
|
|
|
| 42 |
A" |
823 |
783 |
0.67 |
|
|
|
| 43 |
A" |
752 |
715 |
3.57 |
|
|
|
| 44 |
A" |
256 |
244 |
1.00 |
|
|
|
| 45 |
A" |
238 |
227 |
2.30 |
|
|
|
| 46 |
A" |
140 |
133 |
1.48 |
|
|
|
| 47 |
A" |
103 |
98 |
1.32 |
|
|
|
| 48 |
A" |
75 |
71 |
0.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36711.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34913.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.512 |
|
|
|
| 2 |
C |
-0.321 |
|
|
|
| 3 |
C |
-0.256 |
|
|
|
| 4 |
O |
-0.463 |
|
|
|
| 5 |
C |
-0.050 |
|
|
|
| 6 |
C |
-0.326 |
|
|
|
| 7 |
H |
0.181 |
|
|
|
| 8 |
H |
0.169 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.161 |
|
|
|
| 12 |
H |
0.161 |
|
|
|
| 13 |
H |
0.148 |
|
|
|
| 14 |
H |
0.148 |
|
|
|
| 15 |
H |
0.173 |
|
|
|
| 16 |
H |
0.173 |
|
|
|
| 17 |
H |
0.137 |
|
|
|
| 18 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.928 |
-0.637 |
0.000 |
1.125 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.815 |
1.113 |
0.000 |
| y |
1.113 |
-36.088 |
0.000 |
| z |
0.000 |
0.000 |
-39.244 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.149 |
1.113 |
0.000 |
| y |
1.113 |
3.942 |
0.000 |
| z |
0.000 |
0.000 |
-0.793 |
|
| Polar |
|---|
| 3z2-r2 | -1.586 |
|---|
| x2-y2 | -4.727 |
|---|
| xy | 1.113 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.613 |
1.252 |
0.000 |
| y |
1.252 |
9.780 |
0.000 |
| z |
0.000 |
0.000 |
7.692 |
<r2> (average value of r
2) Å
2
| <r2> |
291.738 |
| (<r2>)1/2 |
17.080 |