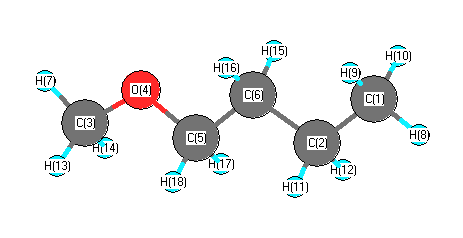
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3067 |
3016 |
32.43 |
|
|
|
| 2 |
A' |
3059 |
3008 |
35.67 |
|
|
|
| 3 |
A' |
2981 |
2932 |
49.16 |
|
|
|
| 4 |
A' |
2977 |
2927 |
23.01 |
|
|
|
| 5 |
A' |
2960 |
2911 |
18.12 |
|
|
|
| 6 |
A' |
2904 |
2857 |
86.28 |
|
|
|
| 7 |
A' |
2873 |
2825 |
41.81 |
|
|
|
| 8 |
A' |
1512 |
1487 |
0.64 |
|
|
|
| 9 |
A' |
1493 |
1468 |
2.61 |
|
|
|
| 10 |
A' |
1485 |
1460 |
12.98 |
|
|
|
| 11 |
A' |
1476 |
1452 |
0.59 |
|
|
|
| 12 |
A' |
1469 |
1445 |
0.31 |
|
|
|
| 13 |
A' |
1452 |
1428 |
0.72 |
|
|
|
| 14 |
A' |
1396 |
1373 |
28.48 |
|
|
|
| 15 |
A' |
1390 |
1367 |
7.63 |
|
|
|
| 16 |
A' |
1355 |
1333 |
0.47 |
|
|
|
| 17 |
A' |
1264 |
1243 |
1.84 |
|
|
|
| 18 |
A' |
1195 |
1175 |
30.76 |
|
|
|
| 19 |
A' |
1143 |
1124 |
146.87 |
|
|
|
| 20 |
A' |
1110 |
1092 |
14.87 |
|
|
|
| 21 |
A' |
1062 |
1045 |
0.84 |
|
|
|
| 22 |
A' |
1026 |
1009 |
1.31 |
|
|
|
| 23 |
A' |
968 |
952 |
20.10 |
|
|
|
| 24 |
A' |
894 |
879 |
6.65 |
|
|
|
| 25 |
A' |
478 |
470 |
1.08 |
|
|
|
| 26 |
A' |
375 |
368 |
2.78 |
|
|
|
| 27 |
A' |
307 |
302 |
0.63 |
|
|
|
| 28 |
A' |
135 |
133 |
0.79 |
|
|
|
| 29 |
A" |
3053 |
3002 |
52.60 |
|
|
|
| 30 |
A" |
3024 |
2974 |
36.89 |
|
|
|
| 31 |
A" |
2990 |
2941 |
5.17 |
|
|
|
| 32 |
A" |
2944 |
2896 |
67.93 |
|
|
|
| 33 |
A" |
2898 |
2850 |
73.13 |
|
|
|
| 34 |
A" |
1482 |
1458 |
7.11 |
|
|
|
| 35 |
A" |
1462 |
1438 |
5.64 |
|
|
|
| 36 |
A" |
1298 |
1276 |
0.07 |
|
|
|
| 37 |
A" |
1282 |
1261 |
0.01 |
|
|
|
| 38 |
A" |
1224 |
1204 |
1.68 |
|
|
|
| 39 |
A" |
1175 |
1155 |
5.06 |
|
|
|
| 40 |
A" |
1144 |
1125 |
0.01 |
|
|
|
| 41 |
A" |
936 |
920 |
0.01 |
|
|
|
| 42 |
A" |
800 |
787 |
0.58 |
|
|
|
| 43 |
A" |
733 |
721 |
3.99 |
|
|
|
| 44 |
A" |
253 |
249 |
1.19 |
|
|
|
| 45 |
A" |
238 |
234 |
1.63 |
|
|
|
| 46 |
A" |
139 |
137 |
1.31 |
|
|
|
| 47 |
A" |
101 |
99 |
1.09 |
|
|
|
| 48 |
A" |
74 |
73 |
0.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35524.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34937.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.481 |
|
|
|
| 2 |
C |
-0.288 |
|
|
|
| 3 |
C |
-0.248 |
|
|
|
| 4 |
O |
-0.414 |
|
|
|
| 5 |
C |
-0.052 |
|
|
|
| 6 |
C |
-0.295 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.157 |
|
|
|
| 9 |
H |
0.158 |
|
|
|
| 10 |
H |
0.158 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.148 |
|
|
|
| 13 |
H |
0.138 |
|
|
|
| 14 |
H |
0.138 |
|
|
|
| 15 |
H |
0.158 |
|
|
|
| 16 |
H |
0.158 |
|
|
|
| 17 |
H |
0.125 |
|
|
|
| 18 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.877 |
-0.554 |
0.000 |
1.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.840 |
1.111 |
0.000 |
| y |
1.111 |
-36.404 |
0.000 |
| z |
0.000 |
0.000 |
-39.441 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.918 |
1.111 |
0.000 |
| y |
1.111 |
3.736 |
0.000 |
| z |
0.000 |
0.000 |
-0.819 |
|
| Polar |
|---|
| 3z2-r2 | -1.637 |
|---|
| x2-y2 | -4.436 |
|---|
| xy | 1.111 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.957 |
1.435 |
0.000 |
| y |
1.435 |
10.332 |
0.000 |
| z |
0.000 |
0.000 |
7.885 |
<r2> (average value of r
2) Å
2
| <r2> |
295.852 |
| (<r2>)1/2 |
17.200 |