return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Butane, 1-(methylthio)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-595.519212
Energy at 298.15K-595.531964
HF Energy-595.261390
Nuclear repulsion energy295.050753
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3188 3026 8.41      
2 A' 3154 2993 37.18      
3 A' 3087 2930 26.60      
4 A' 3078 2922 73.48      
5 A' 3076 2920 27.72      
6 A' 3069 2913 1.34      
7 A' 3063 2907 8.25      
8 A' 1562 1482 3.18      
9 A' 1549 1470 1.07      
10 A' 1540 1461 0.55      
11 A' 1538 1460 1.11      
12 A' 1532 1455 13.42      
13 A' 1460 1386 1.60      
14 A' 1436 1363 3.37      
15 A' 1415 1343 7.33      
16 A' 1374 1304 6.25      
17 A' 1287 1221 20.03      
18 A' 1150 1092 2.38      
19 A' 1089 1033 1.02      
20 A' 1052 999 1.70      
21 A' 1010 958 6.02      
22 A' 921 874 0.88      
23 A' 783 743 2.10      
24 A' 735 697 1.85      
25 A' 427 405 0.53      
26 A' 319 303 0.60      
27 A' 240 227 0.00      
28 A' 118 112 0.96      
29 A" 3168 3007 12.74      
30 A" 3150 2990 62.15      
31 A" 3131 2972 39.71      
32 A" 3111 2953 1.33      
33 A" 3094 2937 4.21      
34 A" 1550 1471 6.26      
35 A" 1519 1442 8.27      
36 A" 1363 1294 0.26      
37 A" 1339 1271 0.34      
38 A" 1264 1200 0.11      
39 A" 1111 1054 0.64      
40 A" 1010 958 3.96      
41 A" 951 903 0.82      
42 A" 815 773 0.03      
43 A" 755 717 3.86      
44 A" 254 241 0.02      
45 A" 180 171 0.50      
46 A" 130 124 0.02      
47 A" 95 90 0.77      
48 A" 55 52 0.80      

Unscaled Zero Point Vibrational Energy (zpe) 36146.0 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 34309.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.36659 0.03150 0.02982

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.303 -2.530 0.000
C2 -1.918 -1.043 0.000
C3 1.769 2.262 0.000
S4 1.515 0.758 0.000
C5 0.000 0.592 0.000
C6 -0.394 -0.892 0.000
H7 2.843 2.461 0.000
H8 -3.390 -2.630 0.000
H9 -1.899 -3.025 0.865
H10 -1.899 -3.025 -0.865
H11 -2.328 -0.578 0.895
H12 -2.328 -0.578 -0.895
H13 1.337 2.738 0.907
H14 1.337 2.738 -0.907
H15 0.015 -1.357 -0.897
H16 0.015 -1.357 0.897
H17 -0.433 1.075 -0.872
H18 -0.433 1.075 0.872

Atom - Atom Distances (Å)
  C1 C2 C3 S4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.53516.28845.03883.87942.51487.16871.09151.07591.07592.14752.14756.46696.46692.74812.74814.15354.1535
C21.53514.95233.87742.52091.53155.91192.16432.16242.16241.08891.08895.07145.07142.15412.15412.73032.7303
C36.28844.95231.52502.43293.82511.09167.11016.49286.49285.06475.06471.11241.11244.12114.12112.64952.6495
S45.03883.87741.52501.52422.52392.15945.96185.16885.16884.16584.16582.18522.18522.74452.74452.15782.1578
C53.87942.52092.43291.52421.53603.40224.67724.17594.17592.75462.75462.68622.68622.14622.14621.08681.0868
C62.51481.53153.82512.52391.53604.66113.46342.74962.74962.15382.15384.12314.12311.09021.09022.15242.1524
H77.16875.91191.09162.15943.40224.66118.04817.30287.30286.06396.06391.77991.77994.83594.83593.66233.6623
H81.09152.16437.11015.96184.67723.46348.04811.76881.76882.47842.47847.21017.21013.74413.74414.82024.8202
H91.07592.16246.49285.16884.17592.74967.30281.76881.72972.48533.04506.60956.84293.08992.53844.68804.3544
H101.07592.16246.49285.16884.17592.74967.30281.76881.72973.04502.48536.84296.60952.53843.08994.35444.6880
H112.14751.08895.06474.16582.75462.15386.06392.47842.48533.04501.78964.94205.26043.05092.46913.07312.5144
H122.14751.08895.06474.16582.75462.15386.06392.47843.04502.48531.78965.26044.94202.46913.05092.51443.0731
H136.46695.07141.11242.18522.68624.12311.77997.21016.60956.84294.94205.26041.81494.66674.30363.01072.4288
H146.46695.07141.11242.18522.68624.12311.77997.21016.84296.60955.26044.94201.81494.30364.66672.42883.0107
H152.74812.15414.12112.74452.14621.09024.83593.74413.08992.53843.05092.46914.66674.30361.79442.47353.0410
H162.74812.15414.12112.74452.14621.09024.83593.74412.53843.08992.46913.05094.30364.66671.79443.04102.4735
H174.15352.73032.64952.15781.08682.15243.66234.82024.68804.35443.07312.51443.01072.42882.47353.04101.7441
H184.15352.73032.64952.15781.08682.15243.66234.82024.35444.68802.51443.07312.42883.01073.04102.47351.7441

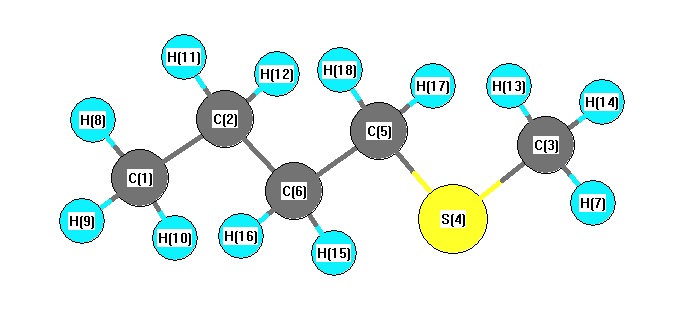
picture of Butane, 1-(methylthio)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C6 110.188 C1 C2 H11 108.657
C1 C2 H12 108.657 C2 C1 H8 109.818
C2 C1 H9 110.594 C2 C1 H10 110.594
C2 C6 C5 110.538 C2 C6 H15 109.345
C2 C6 H16 109.345 C3 S4 C5 105.856
S4 C3 H7 110.130 S4 C3 H13 110.929
S4 C3 H14 110.929 S4 C5 C6 111.131
S4 C5 H17 110.341 S4 C5 H18 110.341
C5 C6 H15 108.421 C5 C6 H16 108.421
C6 C2 H11 109.400 C6 C2 H12 109.400
C6 C5 H17 109.098 C6 C5 H18 109.098
H7 C3 H13 107.710 H7 C3 H14 107.710
H8 C1 H9 109.391 H8 C1 H10 109.391
H9 C1 H10 107.000 H11 C2 H12 110.526
H13 C3 H14 109.323 H15 C6 H16 110.761
H17 C5 H18 106.718
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.460      
2 C -0.275      
3 C -0.609      
4 S 0.101      
5 C -0.425      
6 C -0.289      
7 H 0.189      
8 H 0.153      
9 H 0.152      
10 H 0.152      
11 H 0.144      
12 H 0.144      
13 H 0.180      
14 H 0.180      
15 H 0.160      
16 H 0.160      
17 H 0.170      
18 H 0.170      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 12.186 1.880 0.000
y 1.880 10.686 0.000
z 0.000 0.000 8.417


<r2> (average value of r2) Å2
<r2> 371.989
(<r2>)1/2 19.287