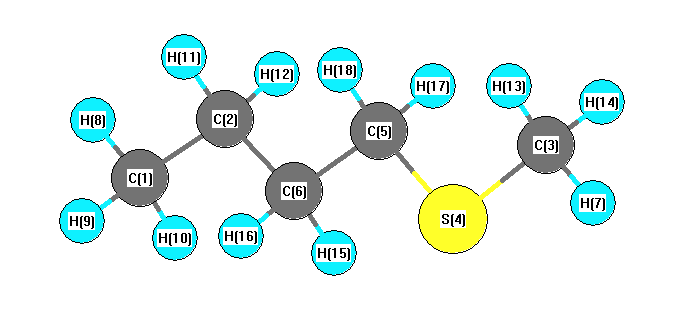
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3182 |
3026 |
6.48 |
|
|
|
| 2 |
A' |
3149 |
2995 |
34.18 |
|
|
|
| 3 |
A' |
3073 |
2923 |
25.16 |
|
|
|
| 4 |
A' |
3067 |
2917 |
73.35 |
|
|
|
| 5 |
A' |
3064 |
2914 |
28.55 |
|
|
|
| 6 |
A' |
3056 |
2906 |
4.76 |
|
|
|
| 7 |
A' |
3051 |
2902 |
7.64 |
|
|
|
| 8 |
A' |
1537 |
1462 |
5.10 |
|
|
|
| 9 |
A' |
1525 |
1450 |
1.13 |
|
|
|
| 10 |
A' |
1516 |
1442 |
0.29 |
|
|
|
| 11 |
A' |
1513 |
1439 |
2.06 |
|
|
|
| 12 |
A' |
1508 |
1434 |
14.10 |
|
|
|
| 13 |
A' |
1439 |
1369 |
2.92 |
|
|
|
| 14 |
A' |
1420 |
1351 |
4.11 |
|
|
|
| 15 |
A' |
1393 |
1325 |
3.80 |
|
|
|
| 16 |
A' |
1354 |
1287 |
6.39 |
|
|
|
| 17 |
A' |
1266 |
1204 |
16.99 |
|
|
|
| 18 |
A' |
1144 |
1088 |
2.97 |
|
|
|
| 19 |
A' |
1094 |
1040 |
0.88 |
|
|
|
| 20 |
A' |
1057 |
1006 |
0.98 |
|
|
|
| 21 |
A' |
1001 |
952 |
8.46 |
|
|
|
| 22 |
A' |
918 |
873 |
1.36 |
|
|
|
| 23 |
A' |
793 |
754 |
1.79 |
|
|
|
| 24 |
A' |
744 |
707 |
2.20 |
|
|
|
| 25 |
A' |
424 |
403 |
0.49 |
|
|
|
| 26 |
A' |
318 |
302 |
0.59 |
|
|
|
| 27 |
A' |
238 |
226 |
0.00 |
|
|
|
| 28 |
A' |
117 |
111 |
1.01 |
|
|
|
| 29 |
A" |
3158 |
3003 |
10.93 |
|
|
|
| 30 |
A" |
3142 |
2989 |
55.86 |
|
|
|
| 31 |
A" |
3121 |
2968 |
40.16 |
|
|
|
| 32 |
A" |
3100 |
2948 |
0.51 |
|
|
|
| 33 |
A" |
3084 |
2932 |
3.92 |
|
|
|
| 34 |
A" |
1527 |
1452 |
7.82 |
|
|
|
| 35 |
A" |
1497 |
1424 |
10.28 |
|
|
|
| 36 |
A" |
1348 |
1282 |
0.29 |
|
|
|
| 37 |
A" |
1322 |
1258 |
0.30 |
|
|
|
| 38 |
A" |
1244 |
1183 |
0.21 |
|
|
|
| 39 |
A" |
1096 |
1042 |
0.66 |
|
|
|
| 40 |
A" |
996 |
948 |
5.12 |
|
|
|
| 41 |
A" |
937 |
892 |
1.01 |
|
|
|
| 42 |
A" |
804 |
764 |
0.02 |
|
|
|
| 43 |
A" |
747 |
711 |
5.15 |
|
|
|
| 44 |
A" |
252 |
240 |
0.03 |
|
|
|
| 45 |
A" |
183 |
174 |
0.60 |
|
|
|
| 46 |
A" |
131 |
124 |
0.02 |
|
|
|
| 47 |
A" |
95 |
90 |
0.83 |
|
|
|
| 48 |
A" |
57 |
54 |
0.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35900.9 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 34141.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.514 |
|
|
|
| 2 |
C |
-0.312 |
|
|
|
| 3 |
C |
-0.668 |
|
|
|
| 4 |
S |
0.109 |
|
|
|
| 5 |
C |
-0.467 |
|
|
|
| 6 |
C |
-0.322 |
|
|
|
| 7 |
H |
0.208 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.170 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.198 |
|
|
|
| 14 |
H |
0.198 |
|
|
|
| 15 |
H |
0.179 |
|
|
|
| 16 |
H |
0.179 |
|
|
|
| 17 |
H |
0.188 |
|
|
|
| 18 |
H |
0.188 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.201 |
1.267 |
0.000 |
1.745 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.464 |
3.288 |
0.000 |
| y |
3.288 |
-43.895 |
0.000 |
| z |
0.000 |
0.000 |
-47.678 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.678 |
3.288 |
0.000 |
| y |
3.288 |
3.676 |
0.000 |
| z |
0.000 |
0.000 |
-1.998 |
|
| Polar |
|---|
| 3z2-r2 | -3.996 |
|---|
| x2-y2 | -3.569 |
|---|
| xy | 3.288 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.327 |
1.924 |
0.000 |
| y |
1.924 |
10.813 |
0.000 |
| z |
0.000 |
0.000 |
8.499 |
<r2> (average value of r
2) Å
2
| <r2> |
369.474 |
| (<r2>)1/2 |
19.222 |