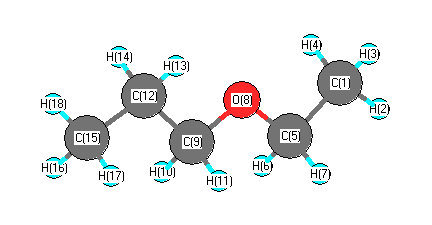
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3012 |
27.12 |
|
|
|
| 2 |
A' |
3136 |
3000 |
32.69 |
|
|
|
| 3 |
A' |
3068 |
2935 |
19.55 |
|
|
|
| 4 |
A' |
3064 |
2931 |
27.27 |
|
|
|
| 5 |
A' |
3053 |
2921 |
27.47 |
|
|
|
| 6 |
A' |
2978 |
2849 |
92.49 |
|
|
|
| 7 |
A' |
2962 |
2834 |
21.40 |
|
|
|
| 8 |
A' |
1564 |
1497 |
0.71 |
|
|
|
| 9 |
A' |
1543 |
1476 |
7.82 |
|
|
|
| 10 |
A' |
1533 |
1466 |
4.38 |
|
|
|
| 11 |
A' |
1526 |
1460 |
2.64 |
|
|
|
| 12 |
A' |
1519 |
1453 |
1.22 |
|
|
|
| 13 |
A' |
1472 |
1408 |
4.58 |
|
|
|
| 14 |
A' |
1437 |
1375 |
1.07 |
|
|
|
| 15 |
A' |
1431 |
1369 |
35.17 |
|
|
|
| 16 |
A' |
1406 |
1345 |
30.32 |
|
|
|
| 17 |
A' |
1342 |
1284 |
13.38 |
|
|
|
| 18 |
A' |
1198 |
1146 |
124.45 |
|
|
|
| 19 |
A' |
1184 |
1133 |
109.60 |
|
|
|
| 20 |
A' |
1127 |
1078 |
6.41 |
|
|
|
| 21 |
A' |
1079 |
1032 |
5.70 |
|
|
|
| 22 |
A' |
1051 |
1005 |
8.45 |
|
|
|
| 23 |
A' |
922 |
882 |
6.94 |
|
|
|
| 24 |
A' |
911 |
871 |
4.39 |
|
|
|
| 25 |
A' |
494 |
472 |
3.28 |
|
|
|
| 26 |
A' |
409 |
391 |
0.87 |
|
|
|
| 27 |
A' |
305 |
292 |
0.64 |
|
|
|
| 28 |
A' |
139 |
133 |
0.42 |
|
|
|
| 29 |
A" |
3153 |
3016 |
27.53 |
|
|
|
| 30 |
A" |
3128 |
2993 |
60.94 |
|
|
|
| 31 |
A" |
3102 |
2967 |
3.83 |
|
|
|
| 32 |
A" |
3002 |
2872 |
47.61 |
|
|
|
| 33 |
A" |
2992 |
2862 |
84.40 |
|
|
|
| 34 |
A" |
1527 |
1461 |
7.70 |
|
|
|
| 35 |
A" |
1509 |
1444 |
5.63 |
|
|
|
| 36 |
A" |
1328 |
1270 |
0.27 |
|
|
|
| 37 |
A" |
1312 |
1255 |
1.70 |
|
|
|
| 38 |
A" |
1281 |
1225 |
1.73 |
|
|
|
| 39 |
A" |
1214 |
1161 |
7.62 |
|
|
|
| 40 |
A" |
1186 |
1135 |
0.73 |
|
|
|
| 41 |
A" |
912 |
873 |
1.67 |
|
|
|
| 42 |
A" |
830 |
794 |
0.93 |
|
|
|
| 43 |
A" |
773 |
739 |
2.08 |
|
|
|
| 44 |
A" |
257 |
245 |
0.87 |
|
|
|
| 45 |
A" |
238 |
228 |
0.01 |
|
|
|
| 46 |
A" |
156 |
149 |
3.40 |
|
|
|
| 47 |
A" |
96 |
92 |
1.17 |
|
|
|
| 48 |
A" |
69 |
66 |
0.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36530.6 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 34948.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.523 |
|
|
-0.351 |
| 2 |
H |
0.165 |
|
|
0.078 |
| 3 |
H |
0.181 |
|
|
0.102 |
| 4 |
H |
0.181 |
|
|
0.102 |
| 5 |
C |
-0.050 |
|
|
0.376 |
| 6 |
H |
0.139 |
|
|
-0.032 |
| 7 |
H |
0.139 |
|
|
-0.032 |
| 8 |
O |
-0.472 |
|
|
-0.441 |
| 9 |
C |
-0.039 |
|
|
0.106 |
| 10 |
H |
0.135 |
|
|
0.012 |
| 11 |
H |
0.135 |
|
|
0.012 |
| 12 |
C |
-0.326 |
|
|
0.182 |
| 13 |
H |
0.173 |
|
|
-0.004 |
| 14 |
H |
0.173 |
|
|
-0.004 |
| 15 |
C |
-0.520 |
|
|
-0.377 |
| 16 |
H |
0.167 |
|
|
0.088 |
| 17 |
H |
0.167 |
|
|
0.088 |
| 18 |
H |
0.174 |
|
|
0.094 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.838 |
-0.489 |
0.000 |
0.971 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.846 |
-0.486 |
0.000 |
0.976 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.209 |
1.894 |
0.000 |
| y |
1.894 |
-38.121 |
0.000 |
| z |
0.000 |
0.000 |
-39.004 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.353 |
1.894 |
0.000 |
| y |
1.894 |
0.485 |
0.000 |
| z |
0.000 |
0.000 |
-0.838 |
|
| Polar |
|---|
| 3z2-r2 | -1.677 |
|---|
| x2-y2 | -0.088 |
|---|
| xy | 1.894 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
285.648 |
| (<r2>)1/2 |
16.901 |