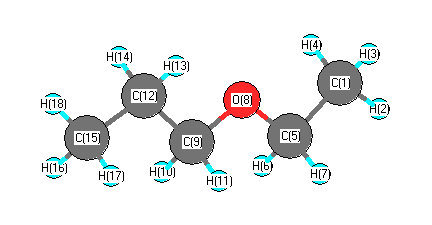
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3076 |
3025 |
24.36 |
|
|
|
| 2 |
A' |
3064 |
3014 |
30.27 |
|
|
|
| 3 |
A' |
2996 |
2947 |
18.20 |
|
|
|
| 4 |
A' |
2988 |
2938 |
34.06 |
|
|
|
| 5 |
A' |
2981 |
2932 |
19.92 |
|
|
|
| 6 |
A' |
2889 |
2842 |
94.41 |
|
|
|
| 7 |
A' |
2871 |
2824 |
24.53 |
|
|
|
| 8 |
A' |
1519 |
1494 |
0.62 |
|
|
|
| 9 |
A' |
1498 |
1474 |
6.46 |
|
|
|
| 10 |
A' |
1489 |
1464 |
4.11 |
|
|
|
| 11 |
A' |
1483 |
1458 |
3.03 |
|
|
|
| 12 |
A' |
1474 |
1450 |
1.84 |
|
|
|
| 13 |
A' |
1421 |
1397 |
4.47 |
|
|
|
| 14 |
A' |
1391 |
1368 |
3.86 |
|
|
|
| 15 |
A' |
1384 |
1361 |
23.84 |
|
|
|
| 16 |
A' |
1357 |
1335 |
32.28 |
|
|
|
| 17 |
A' |
1299 |
1277 |
14.17 |
|
|
|
| 18 |
A' |
1156 |
1137 |
31.35 |
|
|
|
| 19 |
A' |
1135 |
1116 |
193.87 |
|
|
|
| 20 |
A' |
1093 |
1075 |
8.47 |
|
|
|
| 21 |
A' |
1049 |
1031 |
4.34 |
|
|
|
| 22 |
A' |
1020 |
1003 |
10.56 |
|
|
|
| 23 |
A' |
896 |
882 |
5.09 |
|
|
|
| 24 |
A' |
881 |
866 |
6.63 |
|
|
|
| 25 |
A' |
482 |
474 |
3.07 |
|
|
|
| 26 |
A' |
399 |
393 |
0.83 |
|
|
|
| 27 |
A' |
299 |
294 |
0.58 |
|
|
|
| 28 |
A' |
135 |
133 |
0.38 |
|
|
|
| 29 |
A" |
3078 |
3027 |
25.50 |
|
|
|
| 30 |
A" |
3053 |
3003 |
54.38 |
|
|
|
| 31 |
A" |
3024 |
2974 |
6.13 |
|
|
|
| 32 |
A" |
2909 |
2861 |
45.21 |
|
|
|
| 33 |
A" |
2897 |
2850 |
91.52 |
|
|
|
| 34 |
A" |
1483 |
1459 |
7.48 |
|
|
|
| 35 |
A" |
1466 |
1442 |
5.76 |
|
|
|
| 36 |
A" |
1287 |
1265 |
0.11 |
|
|
|
| 37 |
A" |
1266 |
1245 |
1.27 |
|
|
|
| 38 |
A" |
1239 |
1219 |
1.48 |
|
|
|
| 39 |
A" |
1177 |
1157 |
6.84 |
|
|
|
| 40 |
A" |
1148 |
1129 |
0.76 |
|
|
|
| 41 |
A" |
886 |
871 |
1.64 |
|
|
|
| 42 |
A" |
807 |
794 |
1.11 |
|
|
|
| 43 |
A" |
754 |
742 |
2.36 |
|
|
|
| 44 |
A" |
254 |
250 |
0.71 |
|
|
|
| 45 |
A" |
232 |
228 |
0.00 |
|
|
|
| 46 |
A" |
153 |
150 |
2.79 |
|
|
|
| 47 |
A" |
96 |
95 |
1.11 |
|
|
|
| 48 |
A" |
68 |
67 |
0.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35499.9 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 34914.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.497 |
|
|
|
| 2 |
H |
0.156 |
|
|
|
| 3 |
H |
0.171 |
|
|
|
| 4 |
H |
0.171 |
|
|
|
| 5 |
C |
-0.050 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
H |
0.128 |
|
|
|
| 8 |
O |
-0.424 |
|
|
|
| 9 |
C |
-0.042 |
|
|
|
| 10 |
H |
0.124 |
|
|
|
| 11 |
H |
0.124 |
|
|
|
| 12 |
C |
-0.300 |
|
|
|
| 13 |
H |
0.161 |
|
|
|
| 14 |
H |
0.161 |
|
|
|
| 15 |
C |
-0.493 |
|
|
|
| 16 |
H |
0.158 |
|
|
|
| 17 |
H |
0.158 |
|
|
|
| 18 |
H |
0.164 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.747 |
-0.460 |
0.000 |
0.877 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.470 |
1.887 |
0.000 |
| y |
1.887 |
-38.309 |
0.000 |
| z |
0.000 |
0.000 |
-39.297 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.334 |
1.887 |
0.000 |
| y |
1.887 |
0.574 |
0.000 |
| z |
0.000 |
0.000 |
-0.908 |
|
| Polar |
|---|
| 3z2-r2 | -1.816 |
|---|
| x2-y2 | -0.160 |
|---|
| xy | 1.887 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.795 |
1.206 |
0.000 |
| y |
1.206 |
10.577 |
0.000 |
| z |
0.000 |
0.000 |
7.928 |
<r2> (average value of r
2) Å
2
| <r2> |
288.395 |
| (<r2>)1/2 |
16.982 |