return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9ClO (1-Chloro-2-ethoxyethane)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-692.814308
Energy at 298.15K-692.824694
HF Energy-692.541642
Nuclear repulsion energy289.892208
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3171 3010 24.87      
2 A' 3152 2992 14.79      
3 A' 3094 2937 15.87      
4 A' 3035 2881 51.29      
5 A' 3020 2866 30.11      
6 A' 1586 1506 0.79      
7 A' 1568 1489 5.37      
8 A' 1551 1472 3.99      
9 A' 1541 1463 3.22      
10 A' 1487 1412 14.06      
11 A' 1452 1379 9.10      
12 A' 1421 1349 33.48      
13 A' 1321 1254 32.00      
14 A' 1187 1127 120.42      
15 A' 1169 1110 123.49      
16 A' 1098 1042 1.27      
17 A' 1060 1006 14.82      
18 A' 920 873 15.40      
19 A' 777 737 47.28      
20 A' 484 460 0.66      
21 A' 382 362 2.76      
22 A' 269 256 3.33      
23 A' 124 118 1.41      
24 A" 3218 3054 7.78      
25 A" 3179 3018 25.94      
26 A" 3074 2918 39.16      
27 A" 3054 2899 63.04      
28 A" 1535 1457 5.14      
29 A" 1329 1262 1.35      
30 A" 1325 1258 0.73      
31 A" 1248 1184 6.16      
32 A" 1209 1148 3.27      
33 A" 1097 1041 2.53      
34 A" 843 800 0.53      
35 A" 822 780 0.23      
36 A" 261 248 1.01      
37 A" 157 149 8.22      
38 A" 87 83 2.21      
39 A" 61 58 0.27      

Unscaled Zero Point Vibrational Energy (zpe) 28684.1 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27227.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.55138 0.02882 0.02797

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.380 -0.230 0.000
Cl2 -2.655 1.016 0.000
C3 0.000 0.416 0.000
C4 3.227 -1.396 0.000
C5 2.308 -0.180 0.000
O6 0.954 -0.638 0.000
H7 4.271 -1.075 0.000
H8 -1.504 -0.855 0.885
H9 -1.504 -0.855 -0.885
H10 0.111 1.052 -0.889
H11 0.111 1.052 0.889
H12 3.048 -2.006 -0.888
H13 3.048 -2.006 0.888
H14 2.493 0.437 0.890
H15 2.493 0.437 -0.890

Atom - Atom Distances (Å)
  C1 Cl2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.78211.52404.75213.68882.36975.71421.08991.08992.15812.15814.85334.85334.03014.0301
Cl21.78212.72156.35665.10493.96967.23452.36732.36732.90522.90526.51486.51485.25635.2563
C31.52402.72153.70082.38401.42194.52402.15862.15861.09841.09843.99343.99342.64752.6475
C44.75216.35663.70081.52382.39561.09294.84304.84304.06084.06081.09161.09162.16552.1655
C53.68885.10492.38401.52381.42942.15733.97123.97122.67132.67132.16062.16061.09851.0985
O62.36973.96961.42192.39561.42943.34582.62142.62142.08722.08722.65432.65432.07762.0776
H75.71427.23454.52401.09292.15733.34585.84685.84684.75614.75611.77461.77462.49762.4976
H81.08992.36732.15864.84303.97122.62145.84681.76943.06412.49865.01934.69584.20094.5602
H91.08992.36732.15864.84303.97122.62145.84681.76942.49863.06414.69585.01934.56024.2009
H102.15812.90521.09844.06082.67132.08724.75613.06412.49861.77794.23994.59723.03602.4605
H112.15812.90521.09844.06082.67132.08724.75612.49863.06411.77794.59724.23992.46053.0360
H124.85336.51483.99341.09162.16062.65431.77465.01934.69584.23994.59721.77613.07152.5047
H134.85336.51483.99341.09162.16062.65431.77464.69585.01934.59724.23991.77612.50473.0715
H144.03015.25632.64752.16551.09852.07762.49764.20094.56023.03602.46053.07152.50471.7794
H154.03015.25632.64752.16551.09852.07762.49764.56024.20092.46053.03602.50473.07151.7794

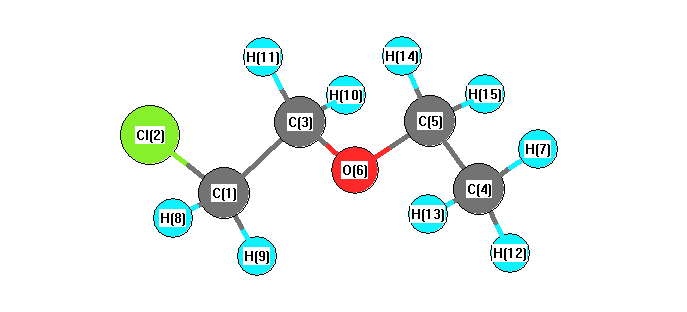
picture of 1-Chloro-2-ethoxyethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 O6 107.054 C1 C3 H10 109.691
C1 C3 H11 109.691 Cl2 C1 C3 110.560
Cl2 C1 H8 108.617 Cl2 C1 H9 108.617
C3 C1 H8 110.233 C3 C1 H9 110.233
C3 O6 C5 113.469 C4 C5 O6 108.385
C4 C5 H14 110.285 C4 C5 H15 110.285
C5 C4 H7 109.971 C5 C4 H12 110.312
C5 C4 H13 110.312 O6 C3 H10 111.176
O6 C3 H11 111.176 O6 C5 H14 109.853
O6 C5 H15 109.853 H7 C4 H12 108.660
H7 C4 H13 108.660 H8 C1 H9 108.525
H10 C3 H11 108.048 H12 C4 H13 108.882
H14 C5 H15 108.177
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.385      
2 Cl -0.098      
3 C 0.011      
4 C -0.472      
5 C 0.000      
6 O -0.546      
7 H 0.153      
8 H 0.213      
9 H 0.213      
10 H 0.151      
11 H 0.151      
12 H 0.168      
13 H 0.168      
14 H 0.135      
15 H 0.135      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.504 -1.442 0.000
y -1.442 7.802 0.000
z 0.000 0.000 6.807


<r2> (average value of r2) Å2
<r2> 366.342
(<r2>)1/2 19.140