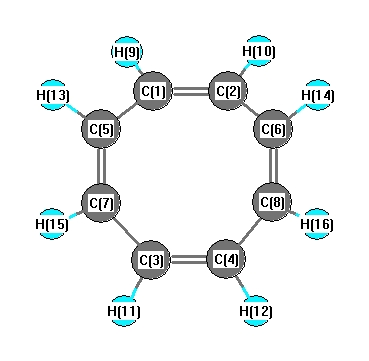
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3166 |
3029 |
0.00 |
|
|
|
| 2 |
A1 |
1753 |
1677 |
0.00 |
|
|
|
| 3 |
A1 |
1247 |
1193 |
0.00 |
|
|
|
| 4 |
A1 |
901 |
862 |
0.00 |
|
|
|
| 5 |
A1 |
785 |
751 |
0.00 |
|
|
|
| 6 |
A1 |
177 |
170 |
0.00 |
|
|
|
| 7 |
A2 |
3132 |
2997 |
0.00 |
|
|
|
| 8 |
A2 |
1391 |
1331 |
0.00 |
|
|
|
| 9 |
A2 |
1008 |
964 |
0.00 |
|
|
|
| 10 |
A2 |
938 |
898 |
0.00 |
|
|
|
| 11 |
A2 |
229 |
220 |
0.00 |
|
|
|
| 12 |
B1 |
3141 |
3005 |
0.00 |
|
|
|
| 13 |
B1 |
1488 |
1423 |
0.00 |
|
|
|
| 14 |
B1 |
1060 |
1014 |
0.00 |
|
|
|
| 15 |
B1 |
979 |
937 |
0.00 |
|
|
|
| 16 |
B1 |
671 |
642 |
0.00 |
|
|
|
| 17 |
B1 |
257 |
246 |
0.00 |
|
|
|
| 18 |
B2 |
3152 |
3016 |
41.84 |
|
|
|
| 19 |
B2 |
1727 |
1652 |
0.76 |
|
|
|
| 20 |
B2 |
1248 |
1194 |
0.88 |
|
|
|
| 21 |
B2 |
686 |
656 |
84.67 |
|
|
|
| 22 |
B2 |
288 |
276 |
1.74 |
|
|
|
| 23 |
E |
3159 |
3023 |
80.18 |
|
|
|
| 23 |
E |
3159 |
3023 |
80.18 |
|
|
|
| 24 |
E |
3136 |
3000 |
1.49 |
|
|
|
| 24 |
E |
3136 |
3000 |
1.49 |
|
|
|
| 25 |
E |
1726 |
1651 |
1.72 |
|
|
|
| 25 |
E |
1726 |
1651 |
1.72 |
|
|
|
| 26 |
E |
1444 |
1381 |
0.09 |
|
|
|
| 26 |
E |
1444 |
1381 |
0.09 |
|
|
|
| 27 |
E |
1262 |
1208 |
0.77 |
|
|
|
| 27 |
E |
1262 |
1208 |
0.77 |
|
|
|
| 28 |
E |
998 |
955 |
0.47 |
|
|
|
| 28 |
E |
998 |
955 |
0.47 |
|
|
|
| 29 |
E |
972 |
930 |
1.38 |
|
|
|
| 29 |
E |
972 |
930 |
1.38 |
|
|
|
| 30 |
E |
818 |
783 |
18.70 |
|
|
|
| 30 |
E |
818 |
783 |
18.70 |
|
|
|
| 31 |
E |
646 |
618 |
19.40 |
|
|
|
| 31 |
E |
646 |
618 |
19.40 |
|
|
|
| 32 |
E |
357 |
342 |
1.22 |
|
|
|
| 32 |
E |
357 |
342 |
1.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29231.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 27965.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.162 |
|
|
|
| 2 |
C |
-0.162 |
|
|
|
| 3 |
C |
-0.162 |
|
|
|
| 4 |
C |
-0.162 |
|
|
|
| 5 |
C |
-0.162 |
|
|
|
| 6 |
C |
-0.162 |
|
|
|
| 7 |
C |
-0.162 |
|
|
|
| 8 |
C |
-0.162 |
|
|
|
| 9 |
H |
0.162 |
|
|
|
| 10 |
H |
0.162 |
|
|
|
| 11 |
H |
0.162 |
|
|
|
| 12 |
H |
0.162 |
|
|
|
| 13 |
H |
0.162 |
|
|
|
| 14 |
H |
0.162 |
|
|
|
| 15 |
H |
0.162 |
|
|
|
| 16 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.678 |
0.000 |
0.000 |
| y |
0.000 |
-42.678 |
0.000 |
| z |
0.000 |
0.000 |
-48.247 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.785 |
0.000 |
0.000 |
| y |
0.000 |
2.785 |
0.000 |
| z |
0.000 |
0.000 |
-5.569 |
|
| Polar |
|---|
| 3z2-r2 | -11.138 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.852 |
0.000 |
0.000 |
| y |
0.000 |
13.852 |
0.000 |
| z |
0.000 |
0.000 |
6.704 |
<r2> (average value of r
2) Å
2
| <r2> |
235.655 |
| (<r2>)1/2 |
15.351 |