return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H8 (cyclooctatetraene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2D 1A1
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-309.219302
Energy at 298.15K-309.227048
HF Energy-308.882517
Nuclear repulsion energy323.560321
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3187 3025 0.00      
2 A1 1740 1652 0.00      
3 A1 1262 1198 0.00      
4 A1 905 859 0.00      
5 A1 784 744 0.00      
6 A1 181 171 0.00      
7 A2 3154 2994 0.00      
8 A2 1407 1336 0.00      
9 A2 999 948 0.00      
10 A2 943 895 0.00      
11 A2 228 217 0.00      
12 B1 3163 3002 0.00      
13 B1 1501 1425 0.00      
14 B1 1065 1011 0.00      
15 B1 969 920 0.00      
16 B1 666 632 0.00      
17 B1 254 241 0.00      
18 B2 3174 3013 43.18      
19 B2 1716 1629 1.08      
20 B2 1258 1194 0.56      
21 B2 692 657 77.01      
22 B2 293 278 1.31      
23 E 3181 3019 78.33      
24 E 3158 2997 1.75      
24 E 3158 2997 1.75      
25 E 1718 1631 1.30      
25 E 1718 1631 1.30      
26 E 1460 1386 0.01      
26 E 1460 1386 0.01      
27 E 1275 1210 0.76      
27 E 1275 1210 0.76      
28 E 992 942 0.13      
28 E 992 942 0.13      
29 E 977 927 2.22      
29 E 977 927 2.22      
30 E 822 780 17.07      
30 E 822 780 17.07      
31 E 648 615 18.06      
31 E 648 615 18.06      
32 E 359 341 1.24      
32 E 359 341 1.24      

Unscaled Zero Point Vibrational Energy (zpe) 29358.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 27867.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.08986 0.08986 0.05077

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is D2d

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.672 1.559 0.385
C2 -0.672 1.559 0.385
C3 0.672 -1.559 0.385
C4 -0.672 -1.559 0.385
C5 1.559 0.672 -0.385
C6 1.559 -0.672 -0.385
C7 -1.559 0.672 -0.385
C8 -1.559 -0.672 -0.385
H9 1.176 2.339 0.955
H10 -1.176 2.339 0.955
H11 1.176 -2.339 0.955
H12 -1.176 -2.339 0.955
H13 2.339 1.176 -0.955
H14 2.339 -1.176 -0.955
H15 -2.339 1.176 -0.955
H16 -2.339 -1.176 -0.955

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.34393.11883.39601.47212.52162.52163.24801.08972.08513.97224.35182.17313.47233.31804.2829
C21.34393.39603.11882.52163.24801.47212.52162.08511.08974.35183.97223.31804.28292.17313.4723
C33.11883.39601.34392.52161.47213.24802.52163.97224.35181.08972.08513.47232.17314.28293.3180
C43.39603.11881.34393.24802.52162.52161.47214.35183.97222.08511.08974.28293.31803.47232.1731
C51.47212.52162.52163.24801.34393.11883.39602.17313.47233.31804.28291.08972.08513.97224.3518
C62.52163.24801.47212.52161.34393.39603.11883.31804.28292.17313.47232.08511.08974.35183.9722
C72.52161.47213.24802.52163.11883.39601.34393.47232.17314.28293.31803.97224.35181.08972.0851
C83.24802.52162.52161.47213.39603.11881.34394.28293.31803.47232.17314.35183.97222.08511.0897
H91.08972.08513.97224.35182.17313.31803.47234.28292.35164.67825.23602.52134.16634.16635.3254
H102.08511.08974.35183.97223.47234.28292.17313.31802.35165.23604.67824.16635.32542.52134.1663
H113.97224.35181.08972.08513.31802.17314.28293.47234.67825.23602.35164.16632.52135.32544.1663
H124.35183.97222.08511.08974.28293.47233.31802.17315.23604.67822.35165.32544.16634.16632.5213
H132.17313.31803.47234.28291.08972.08513.97224.35182.52134.16634.16635.32542.35164.67825.2360
H143.47234.28292.17313.31802.08511.08974.35183.97224.16635.32542.52134.16632.35165.23604.6782
H153.31802.17314.28293.47233.97224.35181.08972.08514.16632.52135.32544.16634.67825.23602.3516
H164.28293.47233.31802.17314.35183.97222.08511.08975.32544.16634.16632.52135.23604.67822.3516

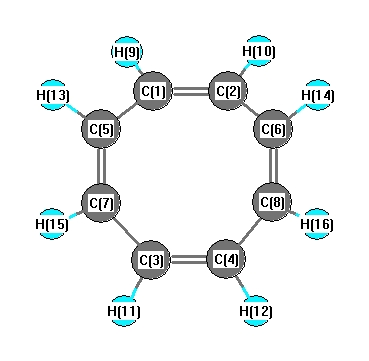
picture of cyclooctatetraene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C6 46.608 C1 C2 H10 117.543
C1 C5 C7 52.925 C1 C5 H13 115.230
C2 C1 C5 127.075 C2 C1 H9 117.543
C2 C6 C8 46.608 C2 C6 H14 158.947
C3 C4 C8 127.075 C3 C4 H12 117.543
C3 C7 C5 46.608 C3 C7 H15 158.947
C4 C3 C7 46.608 C4 C3 H11 117.543
C4 C8 C6 52.925 C4 C8 H16 115.230
C5 C1 H9 115.230 C5 C7 H15 135.685
C6 C2 H10 158.947 C6 C8 H16 135.685
C7 C3 H11 158.947 C7 C5 H13 135.685
C8 C4 H12 115.230 C8 C6 H14 135.685
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.156      
2 C -0.156      
3 C -0.156      
4 C -0.156      
5 C -0.156      
6 C -0.156      
7 C -0.156      
8 C -0.156      
9 H 0.156      
10 H 0.156      
11 H 0.156      
12 H 0.156      
13 H 0.156      
14 H 0.156      
15 H 0.156      
16 H 0.156      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 13.475 0.000 0.000
y 0.000 13.475 0.000
z 0.000 0.000 6.668


<r2> (average value of r2) Å2
<r2> 235.873
(<r2>)1/2 15.358