return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (E)-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-195.730522
Energy at 298.15K-195.740816
HF Energy-195.143355
Nuclear repulsion energy171.209026
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3283 3033 23.33      
2 A' 3249 3001 30.66      
3 A' 3239 2991 33.62      
4 A' 3235 2988 15.38      
5 A' 3162 2921 18.48      
6 A' 3148 2908 33.22      
7 A' 3137 2897 35.59      
8 A' 1861 1719 0.08      
9 A' 1604 1482 4.22      
10 A' 1591 1470 8.24      
11 A' 1584 1463 1.07      
12 A' 1520 1404 7.07      
13 A' 1511 1396 0.69      
14 A' 1470 1358 7.56      
15 A' 1414 1306 0.69      
16 A' 1403 1296 0.02      
17 A' 1213 1121 1.50      
18 A' 1152 1064 3.14      
19 A' 1096 1013 2.11      
20 A' 1007 930 5.27      
21 A' 909 839 0.02      
22 A' 608 562 0.50      
23 A' 383 354 0.18      
24 A' 218 202 0.25      
25 A" 3240 2993 36.03      
26 A" 3209 2964 27.22      
27 A" 3165 2924 22.91      
28 A" 1594 1472 6.01      
29 A" 1576 1455 6.01      
30 A" 1368 1263 0.18      
31 A" 1180 1090 2.20      
32 A" 1138 1051 0.85      
33 A" 1066 985 28.36      
34 A" 878 811 2.27      
35 A" 752 695 0.39      
36 A" 316 292 1.54      
37 A" 231 213 0.92      
38 A" 204 189 1.33      
39 A" 124 114 0.04      

Unscaled Zero Point Vibrational Energy (zpe) 31518.6 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 29113.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.45104 0.08235 0.07241

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.734 -1.227 0.000
C2 -0.975 -0.171 0.000
H3 -0.270 1.780 0.000
C4 0.000 0.728 0.000
H5 -2.935 -0.252 0.877
H6 -2.935 -0.252 -0.877
H7 -2.595 1.241 0.000
C8 -2.437 0.164 0.000
H9 1.921 0.950 -0.869
H10 1.921 0.950 0.869
C11 1.479 0.455 0.000
H12 2.963 -1.112 0.000
H13 1.497 -1.527 0.881
H14 1.497 -1.527 -0.881
C15 1.879 -1.014 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 H7 C8 H9 H10 C11 H12 H13 H14 C15
H11.08323.04262.08842.56252.56253.09122.19863.54193.54192.77943.69872.41702.41702.6212
C21.08322.07461.32652.14892.14892.14901.49933.22513.22512.53254.04922.95402.95402.9756
H33.04262.07461.08593.46453.46452.38722.70332.49862.49862.19354.33793.85163.85163.5242
C42.08841.32651.08593.21653.21652.64552.50122.12012.12011.50353.48822.84672.84672.5619
H52.56252.14893.46453.21651.75391.76441.09115.29895.00314.55546.02504.61184.93564.9521
H62.56252.14893.46453.21651.75391.76441.09115.00315.29894.55546.02504.93564.61184.9521
H73.09122.14902.38722.64551.76441.76441.08884.60844.60844.14896.03615.01835.01835.0099
C82.19861.49932.70332.50121.09111.09111.08884.51274.51273.92615.54854.37124.37124.4731
H93.54193.22512.49862.12015.29895.00314.60844.51271.73731.09342.46893.06292.51392.1480
H103.54193.22512.49862.12015.00315.29894.60844.51271.73731.09342.46892.51393.06292.1480
C112.77942.53252.19351.50354.55544.55544.14893.92611.09341.09342.15912.16982.16981.5226
H123.69874.04924.33793.48826.02506.02506.03615.54852.46892.46892.15911.76051.76051.0889
H132.41702.95403.85162.84674.61184.93565.01834.37123.06292.51392.16981.76051.76231.0892
H142.41702.95403.85162.84674.93564.61185.01834.37122.51393.06292.16981.76051.76231.0892
C152.62122.97563.52422.56194.95214.95215.00994.47312.14802.14801.52261.08891.08921.0892

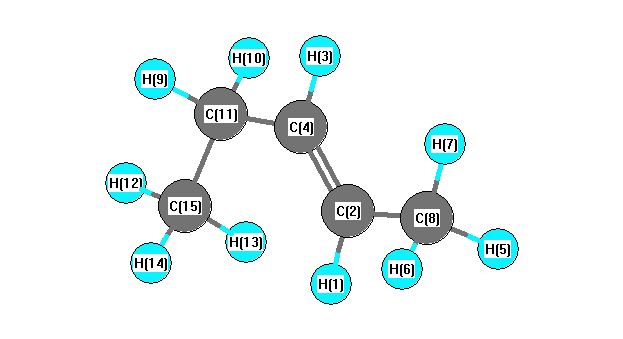
picture of 2-Pentene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 119.804 H1 C2 C8 115.774
C2 C4 H3 118.290 C2 C4 C11 126.870
C2 C8 H5 111.126 C2 C8 H6 111.126
C2 C8 H7 111.276 H3 C4 C11 114.840
C4 C2 C8 124.422 C4 C11 H9 108.418
C4 C11 H10 108.418 C4 C11 C15 115.687
H5 C8 H6 106.982 H5 C8 H7 108.078
H6 C8 H7 108.078 H9 C11 H10 105.209
H9 C11 C15 109.296 H10 C11 C15 109.296
C11 C15 H12 110.436 C11 C15 H13 111.280
C11 C15 H14 111.280 H12 C15 H13 107.852
H12 C15 H14 107.852 H13 C15 H14 107.998
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability