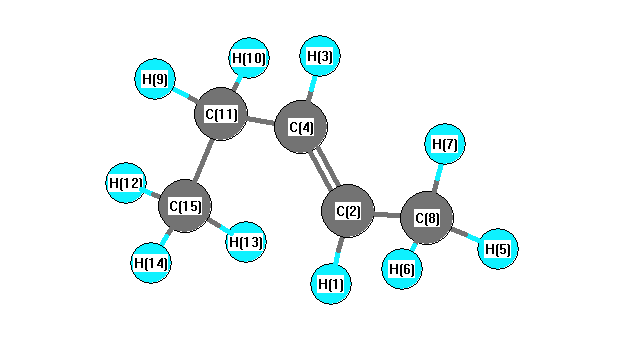
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3168 |
3031 |
31.61 |
|
|
|
| 2 |
A' |
3139 |
3003 |
28.63 |
|
|
|
| 3 |
A' |
3137 |
3001 |
33.92 |
|
|
|
| 4 |
A' |
3130 |
2994 |
12.78 |
|
|
|
| 5 |
A' |
3059 |
2926 |
21.88 |
|
|
|
| 6 |
A' |
3038 |
2906 |
39.12 |
|
|
|
| 7 |
A' |
3021 |
2891 |
36.37 |
|
|
|
| 8 |
A' |
1769 |
1692 |
0.18 |
|
|
|
| 9 |
A' |
1533 |
1467 |
5.22 |
|
|
|
| 10 |
A' |
1518 |
1452 |
9.34 |
|
|
|
| 11 |
A' |
1502 |
1437 |
2.12 |
|
|
|
| 12 |
A' |
1440 |
1378 |
7.21 |
|
|
|
| 13 |
A' |
1433 |
1371 |
0.60 |
|
|
|
| 14 |
A' |
1392 |
1332 |
9.94 |
|
|
|
| 15 |
A' |
1347 |
1288 |
0.82 |
|
|
|
| 16 |
A' |
1331 |
1273 |
0.03 |
|
|
|
| 17 |
A' |
1159 |
1108 |
1.31 |
|
|
|
| 18 |
A' |
1107 |
1059 |
5.53 |
|
|
|
| 19 |
A' |
1045 |
1000 |
2.68 |
|
|
|
| 20 |
A' |
966 |
924 |
8.34 |
|
|
|
| 21 |
A' |
872 |
834 |
0.05 |
|
|
|
| 22 |
A' |
584 |
559 |
0.94 |
|
|
|
| 23 |
A' |
364 |
349 |
0.21 |
|
|
|
| 24 |
A' |
202 |
194 |
0.30 |
|
|
|
| 25 |
A" |
3133 |
2997 |
34.04 |
|
|
|
| 26 |
A" |
3092 |
2959 |
25.47 |
|
|
|
| 27 |
A" |
3046 |
2914 |
22.48 |
|
|
|
| 28 |
A" |
1524 |
1458 |
6.86 |
|
|
|
| 29 |
A" |
1504 |
1439 |
6.77 |
|
|
|
| 30 |
A" |
1302 |
1246 |
0.06 |
|
|
|
| 31 |
A" |
1114 |
1065 |
1.38 |
|
|
|
| 32 |
A" |
1073 |
1027 |
0.00 |
|
|
|
| 33 |
A" |
1016 |
972 |
27.09 |
|
|
|
| 34 |
A" |
832 |
796 |
3.30 |
|
|
|
| 35 |
A" |
712 |
682 |
0.56 |
|
|
|
| 36 |
A" |
302 |
289 |
1.95 |
|
|
|
| 37 |
A" |
226 |
216 |
0.19 |
|
|
|
| 38 |
A" |
201 |
193 |
2.48 |
|
|
|
| 39 |
A" |
126 |
120 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30228.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 28919.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.148 |
|
|
0.100 |
| 2 |
C |
-0.116 |
|
|
-0.009 |
| 3 |
H |
0.141 |
|
|
0.127 |
| 4 |
C |
-0.128 |
|
|
-0.262 |
| 5 |
H |
0.175 |
|
|
0.116 |
| 6 |
H |
0.175 |
|
|
0.116 |
| 7 |
H |
0.173 |
|
|
0.119 |
| 8 |
C |
-0.556 |
|
|
-0.391 |
| 9 |
H |
0.170 |
|
|
0.033 |
| 10 |
H |
0.170 |
|
|
0.033 |
| 11 |
C |
-0.346 |
|
|
0.068 |
| 12 |
H |
0.168 |
|
|
0.067 |
| 13 |
H |
0.172 |
|
|
0.068 |
| 14 |
H |
0.172 |
|
|
0.068 |
| 15 |
C |
-0.518 |
|
|
-0.254 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.087 |
0.008 |
0.000 |
0.087 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.077 |
0.032 |
0.000 |
0.083 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.294 |
0.482 |
0.000 |
| y |
0.482 |
-31.815 |
0.000 |
| z |
0.000 |
0.000 |
-33.898 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.563 |
0.482 |
0.000 |
| y |
0.482 |
0.781 |
0.000 |
| z |
0.000 |
0.000 |
-2.344 |
|
| Polar |
|---|
| 3z2-r2 | -4.687 |
|---|
| x2-y2 | 0.521 |
|---|
| xy | 0.482 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
171.637 |
| (<r2>)1/2 |
13.101 |