return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10 (2-Pentene, (E)-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-196.272408
Energy at 298.15K-196.282545
HF Energy-196.058093
Nuclear repulsion energy170.556212
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3194 3032 27.95      
2 A' 3159 2999 31.67      
3 A' 3158 2997 34.40      
4 A' 3150 2990 12.73      
5 A' 3083 2927 19.98      
6 A' 3065 2909 35.96      
7 A' 3049 2894 35.11      
8 A' 1764 1674 0.16      
9 A' 1559 1480 4.20      
10 A' 1545 1467 7.67      
11 A' 1532 1454 1.07      
12 A' 1465 1390 5.05      
13 A' 1458 1384 1.16      
14 A' 1414 1343 11.40      
15 A' 1363 1294 0.79      
16 A' 1351 1283 0.00      
17 A' 1171 1112 1.40      
18 A' 1111 1055 3.93      
19 A' 1060 1006 2.17      
20 A' 974 924 6.11      
21 A' 877 832 0.03      
22 A' 588 558 0.67      
23 A' 369 351 0.17      
24 A' 209 199 0.25      
25 A" 3157 2997 35.14      
26 A" 3120 2961 26.96      
27 A" 3073 2917 23.78      
28 A" 1549 1471 5.59      
29 A" 1531 1453 5.49      
30 A" 1322 1255 0.11      
31 A" 1135 1077 1.41      
32 A" 1091 1036 0.07      
33 A" 1023 971 24.34      
34 A" 841 798 2.54      
35 A" 720 683 0.31      
36 A" 306 290 1.62      
37 A" 225 214 0.34      
38 A" 202 191 1.94      
39 A" 123 117 0.07      

Unscaled Zero Point Vibrational Energy (zpe) 30542.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 28991.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.44644 0.08185 0.07192

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -0.767 -1.201 0.000
C2 -0.989 -0.187 0.000
H3 -0.283 1.723 0.000
C4 0.000 0.720 0.000
H5 -2.917 -0.192 0.885
H6 -2.917 -0.192 -0.885
H7 -2.501 1.296 0.000
C8 -2.407 0.212 0.000
H9 1.894 0.923 -0.875
H10 1.894 0.923 0.875
C11 1.458 0.442 0.000
H12 2.968 -1.112 0.000
H13 1.494 -1.544 0.876
H14 1.494 -1.544 -0.876
C15 1.878 -1.033 0.000

Atom - Atom Distances (Å)
  H1 C2 H3 C4 H5 H6 H7 C8 H9 H10 C11 H12 H13 H14 C15
H11.03792.96342.06802.53422.53423.03942.16433.51533.51532.76593.73622.44962.44962.6509
C21.03792.03591.34152.12172.12172.11761.47313.21063.21062.52654.06352.96262.96262.9893
H32.96342.03591.04243.37503.37502.25872.60662.47882.47882.16174.31323.82123.82123.5019
C42.06801.34151.04243.18213.18212.56652.46002.09602.09601.48433.48742.85062.85062.5686
H52.53422.12173.37503.18211.76941.78061.09845.24294.93894.50886.02194.61414.93874.9483
H62.53422.12173.37503.18211.76941.78061.09844.93895.24294.50886.02194.93874.61414.9483
H73.03942.11762.25872.56651.78061.78061.08794.49664.49664.05025.97534.97954.97954.9597
C82.16431.47312.60662.46001.09841.09841.08794.44634.44633.87205.53554.36724.36724.4623
H93.51533.21062.47882.09605.24294.93894.49664.44631.74911.08922.46133.05172.49952.1422
H103.51533.21062.47882.09604.93895.24294.49664.44631.74911.08922.46132.49953.05172.1422
C112.76592.52652.16171.48434.50884.50884.05023.87201.08921.08922.16632.17112.17111.5330
H123.73624.06354.31323.48746.02196.02195.97535.53552.46132.46132.16631.76821.76821.0925
H132.44962.96263.82122.85064.61414.93874.97954.36723.05172.49952.17111.76821.75261.0851
H142.44962.96263.82122.85064.93874.61414.97954.36722.49953.05172.17111.76821.75261.0851
C152.65092.98933.50192.56864.94834.94834.95974.46232.14222.14221.53301.09251.08511.0851

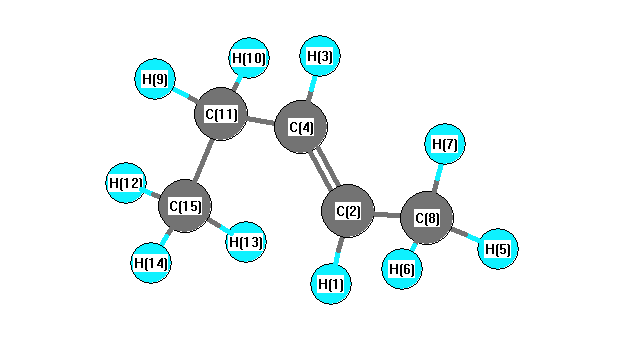
picture of 2-Pentene, (E)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C2 C4 120.176 H1 C2 C8 118.033
C2 C4 H3 116.749 C2 C4 C11 126.705
C2 C8 H5 110.341 C2 C8 H6 110.341
C2 C8 H7 110.656 H3 C4 C11 116.546
C4 C2 C8 121.791 C4 C11 H9 108.085
C4 C11 H10 108.085 C4 C11 C15 116.695
H5 C8 H6 107.301 H5 C8 H7 109.062
H6 C8 H7 109.062 H9 C11 H10 106.820
H9 C11 C15 108.372 H10 C11 C15 108.372
C11 C15 H12 110.070 C11 C15 H13 110.895
C11 C15 H14 110.895 H12 C15 H13 108.586
H12 C15 H14 108.586 H13 C15 H14 107.727
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.144      
2 C -0.121      
3 H 0.136      
4 C -0.132      
5 H 0.159      
6 H 0.159      
7 H 0.158      
8 C -0.498      
9 H 0.153      
10 H 0.153      
11 C -0.304      
12 H 0.152      
13 H 0.156      
14 H 0.156      
15 C -0.469      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.090 0.645 0.000
y 0.645 7.713 0.000
z 0.000 0.000 5.658


<r2> (average value of r2) Å2
<r2> 171.980
(<r2>)1/2 13.114