Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B3PW91/6-31G*
Geometric Data calculated at B3PW91/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/6-31G*
| | hartrees |
|---|
| Energy at 0K | -268.244635 |
| Energy at 298.15K | -268.252867 |
| Nuclear repulsion energy | 194.705082 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3150 |
3013 |
27.73 |
|
|
|
| 2 |
A |
3116 |
2982 |
39.61 |
|
|
|
| 3 |
A |
3091 |
2957 |
67.76 |
|
|
|
| 4 |
A |
3042 |
2910 |
66.13 |
|
|
|
| 5 |
A |
3020 |
2889 |
50.29 |
|
|
|
| 6 |
A |
2991 |
2861 |
118.18 |
|
|
|
| 7 |
A |
1579 |
1511 |
4.95 |
|
|
|
| 8 |
A |
1548 |
1481 |
2.75 |
|
|
|
| 9 |
A |
1533 |
1467 |
1.68 |
|
|
|
| 10 |
A |
1451 |
1388 |
12.45 |
|
|
|
| 11 |
A |
1398 |
1338 |
7.01 |
|
|
|
| 12 |
A |
1357 |
1298 |
0.30 |
|
|
|
| 13 |
A |
1274 |
1218 |
5.51 |
|
|
|
| 14 |
A |
1251 |
1197 |
12.48 |
|
|
|
| 15 |
A |
1213 |
1161 |
72.96 |
|
|
|
| 16 |
A |
1190 |
1139 |
43.05 |
|
|
|
| 17 |
A |
1161 |
1111 |
55.67 |
|
|
|
| 18 |
A |
1151 |
1101 |
63.20 |
|
|
|
| 19 |
A |
1095 |
1048 |
39.38 |
|
|
|
| 20 |
A |
1004 |
960 |
26.11 |
|
|
|
| 21 |
A |
975 |
933 |
40.01 |
|
|
|
| 22 |
A |
963 |
921 |
15.25 |
|
|
|
| 23 |
A |
909 |
870 |
21.03 |
|
|
|
| 24 |
A |
741 |
709 |
0.30 |
|
|
|
| 25 |
A |
655 |
627 |
4.22 |
|
|
|
| 26 |
A |
270 |
258 |
5.11 |
|
|
|
| 27 |
A |
37 |
36 |
10.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20581.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 19690.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.188 |
-0.067 |
0.078 |
| C2 |
-0.864 |
0.812 |
0.198 |
| C3 |
-1.011 |
-0.669 |
-0.136 |
| O4 |
0.454 |
1.083 |
-0.251 |
| O5 |
0.303 |
-1.174 |
0.058 |
| H6 |
1.981 |
-0.190 |
-0.669 |
| H7 |
1.629 |
0.013 |
1.086 |
| H8 |
-0.958 |
0.990 |
1.281 |
| H9 |
-1.328 |
-0.814 |
-1.179 |
| H10 |
-1.702 |
-1.202 |
0.526 |
| H11 |
-1.553 |
1.469 |
-0.339 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2356 | 2.2901 | 1.4034 | 1.4171 | 1.0965 | 1.1037 | 2.6777 | 2.9096 | 3.1367 | 3.1691 |
C2 | 2.2356 | | 1.5248 | 1.4183 | 2.3074 | 3.1389 | 2.7645 | 1.1018 | 2.1805 | 2.2057 | 1.0926 | C3 | 2.2901 | 1.5248 | | 2.2866 | 1.4213 | 3.0773 | 2.9882 | 2.1819 | 1.0994 | 1.0950 | 2.2144 | O4 | 1.4034 | 1.4183 | 2.2866 | | 2.2827 | 2.0322 | 2.0768 | 2.0853 | 2.7632 | 3.2358 | 2.0450 | O5 | 1.4171 | 2.3074 | 1.4213 | 2.2827 | | 2.0767 | 2.0552 | 2.7868 | 2.0780 | 2.0587 | 3.2533 | H6 | 1.0965 | 3.1389 | 3.0773 | 2.0322 | 2.0767 | | 1.8018 | 3.7196 | 3.4058 | 4.0020 | 3.9180 | H7 | 1.1037 | 2.7645 | 2.9882 | 2.0768 | 2.0552 | 1.8018 | | 2.7721 | 3.8153 | 3.5893 | 3.7779 | H8 | 2.6777 | 1.1018 | 2.1819 | 2.0853 | 2.7868 | 3.7196 | 2.7721 | | 3.0727 | 2.4345 | 1.7909 | H9 | 2.9096 | 2.1805 | 1.0994 | 2.7632 | 2.0780 | 3.4058 | 3.8153 | 3.0727 | | 1.7877 | 2.4432 | H10 | 3.1367 | 2.2057 | 1.0950 | 3.2358 | 2.0587 | 4.0020 | 3.5893 | 2.4345 | 1.7877 | | 2.8115 | H11 | 3.1691 | 1.0926 | 2.2144 | 2.0450 | 3.2533 | 3.9180 | 3.7779 | 1.7909 | 2.4432 | 2.8115 | |
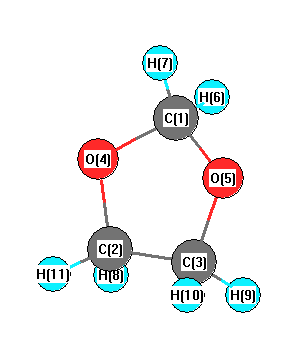 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
104.800 |
|
C1 |
O5 |
C3 |
107.571 |
| C2 |
C3 |
O5 |
103.052 |
|
C2 |
C3 |
H9 |
111.346 |
| C2 |
C3 |
H10 |
113.677 |
|
C3 |
C2 |
O4 |
101.897 |
| C3 |
C2 |
H8 |
111.323 |
|
C3 |
C2 |
H11 |
114.557 |
| O4 |
C1 |
O5 |
108.063 |
|
O4 |
C1 |
H6 |
108.137 |
| O4 |
C1 |
H7 |
111.306 |
|
O4 |
C2 |
H8 |
111.056 |
| O4 |
C2 |
H11 |
108.373 |
|
O5 |
C1 |
H6 |
110.773 |
| O5 |
C1 |
H7 |
108.596 |
|
O5 |
C3 |
H9 |
110.395 |
| O5 |
C3 |
H10 |
109.110 |
|
H6 |
C1 |
H7 |
109.958 |
| H8 |
C2 |
H11 |
109.402 |
|
H9 |
C3 |
H10 |
109.105 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.168 |
|
|
|
| 2 |
C |
-0.121 |
|
|
|
| 3 |
C |
-0.078 |
|
|
|
| 4 |
O |
-0.472 |
|
|
|
| 5 |
O |
-0.496 |
|
|
|
| 6 |
H |
0.168 |
|
|
|
| 7 |
H |
0.147 |
|
|
|
| 8 |
H |
0.163 |
|
|
|
| 9 |
H |
0.165 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.184 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.185 |
0.328 |
0.427 |
1.302 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.563 |
-0.598 |
0.189 |
| y |
-0.598 |
-33.923 |
0.881 |
| z |
0.189 |
0.881 |
-29.295 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.046 |
-0.598 |
0.189 |
| y |
-0.598 |
-6.494 |
0.881 |
| z |
0.189 |
0.881 |
0.449 |
|
| Polar |
|---|
| 3z2-r2 | 0.897 |
|---|
| x2-y2 | 8.360 |
|---|
| xy | -0.598 |
|---|
| xz | 0.189 |
|---|
| yz | 0.881 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.357 |
-0.075 |
0.058 |
| y |
-0.075 |
4.869 |
0.097 |
| z |
0.058 |
0.097 |
5.002 |
<r2> (average value of r
2) Å
2
| <r2> |
92.830 |
| (<r2>)1/2 |
9.635 |
