return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O2 (1,3-Dioxolane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-267.567351
Energy at 298.15K-267.575604
HF Energy-266.789976
Nuclear repulsion energy194.033044
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3150 3000 19.43      
2 A 3054 2908 20.90      
3 A 3048 2902 94.07      
4 A 1607 1531 6.40      
5 A 1568 1493 1.86      
6 A 1421 1353 12.13      
7 A 1281 1220 5.32      
8 A 1239 1180 72.32      
9 A 1190 1133 54.72      
10 A 1173 1117 96.05      
11 A 994 947 0.14      
12 A 982 935 8.16      
13 A 739 703 0.30      
14 A 288 274 0.87      
15 A 3155 3005 38.70      
16 A 3096 2949 70.14      
17 A 3056 2911 67.91      
18 A 1562 1487 1.08      
19 A 1462 1392 11.12      
20 A 1369 1304 0.02      
21 A 1253 1193 4.00      
22 A 1181 1125 20.63      
23 A 1121 1067 37.83      
24 A 991 943 46.91      
25 A 932 888 28.48      
26 A 653 622 5.28      
27 A 24 22 20.16      

Unscaled Zero Point Vibrational Energy (zpe) 20793.2 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 19801.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.25854 0.25333 0.14352

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.207
C2 -0.313 0.692 -0.945
C3 0.313 -0.692 -0.945
O4 0.000 1.150 0.372
O5 0.000 -1.150 0.372
H6 0.901 0.043 1.835
H7 -0.901 -0.043 1.835
H8 -1.402 0.637 -1.098
H9 1.402 -0.637 -1.098
H10 -0.125 -1.394 -1.661
H11 0.125 1.394 -1.661

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9 H10 H11
C12.28202.28201.42131.42131.09931.09932.77162.77163.19113.1911
C22.28201.51981.42882.28623.10222.93491.10102.17512.21411.0943
C32.28201.51982.28621.42882.93493.10222.17511.10101.09432.2141
O41.42131.42882.28622.30022.04412.09202.09502.70503.25912.0510
O51.42132.28621.42882.30022.09202.04412.70502.09502.05103.2591
H61.09933.10222.93492.04412.09201.80463.77593.05163.91653.8272
H71.09932.93493.10222.09202.04411.80463.05163.77593.82723.9165
H82.77161.10102.17512.09502.70503.77593.05163.07952.46411.7956
H92.77162.17511.10102.70502.09503.05163.77593.07951.79562.4641
H103.19112.21411.09433.25912.05103.91653.82722.46411.79562.8002
H113.19111.09432.21412.05103.25913.82723.91651.79562.46412.8002

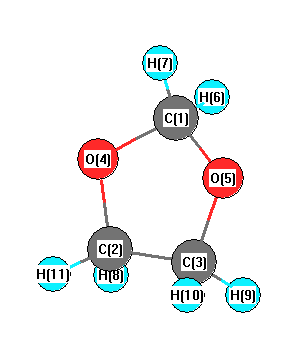
picture of 1,3-Dioxolane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O4 C2 106.392 C1 O5 C3 106.392
C2 C3 O5 101.630 C2 C3 H9 111.184
C2 C3 H10 114.801 C3 C2 O4 101.630
C3 C2 H8 111.184 C3 C2 H11 114.801
O4 C1 O5 108.037 O4 C1 H6 107.695
O4 C1 H7 111.548 O4 C2 H8 111.152
O4 C2 H11 108.026 O5 C1 H6 111.548
O5 C1 H7 107.695 O5 C3 H9 111.152
O5 C3 H10 108.026 H6 C1 H7 110.327
H8 C2 H11 109.760 H9 C3 H10 109.760
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability