Jump to
S1C2
S1C3
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -268.026716 |
| Energy at 298.15K | -268.034818 |
| HF Energy | -268.026716 |
| Nuclear repulsion energy | 193.236300 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3055 |
3004 |
22.41 |
|
|
|
| 2 |
A |
2932 |
2884 |
0.08 |
|
|
|
| 3 |
A |
2921 |
2873 |
148.40 |
|
|
|
| 4 |
A |
1529 |
1504 |
5.14 |
|
|
|
| 5 |
A |
1488 |
1464 |
2.37 |
|
|
|
| 6 |
A |
1348 |
1326 |
10.15 |
|
|
|
| 7 |
A |
1223 |
1203 |
8.63 |
|
|
|
| 8 |
A |
1175 |
1156 |
68.22 |
|
|
|
| 9 |
A |
1129 |
1110 |
25.06 |
|
|
|
| 10 |
A |
1103 |
1085 |
114.75 |
|
|
|
| 11 |
A |
951 |
935 |
4.79 |
|
|
|
| 12 |
A |
933 |
917 |
2.49 |
|
|
|
| 13 |
A |
709 |
698 |
0.92 |
|
|
|
| 14 |
A |
275 |
270 |
0.60 |
|
|
|
| 15 |
B |
3058 |
3007 |
36.14 |
|
|
|
| 16 |
B |
2955 |
2907 |
109.19 |
|
|
|
| 17 |
B |
2942 |
2893 |
67.85 |
|
|
|
| 18 |
B |
1485 |
1461 |
1.45 |
|
|
|
| 19 |
B |
1399 |
1376 |
10.87 |
|
|
|
| 20 |
B |
1304 |
1283 |
0.09 |
|
|
|
| 21 |
B |
1193 |
1173 |
1.76 |
|
|
|
| 22 |
B |
1122 |
1103 |
13.10 |
|
|
|
| 23 |
B |
1035 |
1018 |
25.84 |
|
|
|
| 24 |
B |
935 |
920 |
56.84 |
|
|
|
| 25 |
B |
889 |
874 |
26.72 |
|
|
|
| 26 |
B |
624 |
613 |
3.46 |
|
|
|
| 27 |
B |
25 |
25 |
15.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19867.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 19539.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
1.203 |
| C2 |
-0.285 |
0.713 |
-0.946 |
| C3 |
0.285 |
-0.713 |
-0.946 |
| O4 |
0.000 |
1.154 |
0.378 |
| O5 |
0.000 |
-1.154 |
0.378 |
| H6 |
0.905 |
0.045 |
1.845 |
| H7 |
-0.905 |
-0.045 |
1.845 |
| H8 |
-1.376 |
0.706 |
-1.153 |
| H9 |
1.376 |
-0.706 |
-1.153 |
| H10 |
-0.212 |
-1.396 |
-1.653 |
| H11 |
0.212 |
1.396 |
-1.653 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 2.2816 | 2.2816 | 1.4186 | 1.4186 | 1.1105 | 1.1105 | 2.8179 | 2.8179 | 3.1861 | 3.1861 |
C2 | 2.2816 | | 1.5356 | 1.4241 | 2.3063 | 3.1067 | 2.9577 | 1.1103 | 2.1943 | 2.2253 | 1.1018 | C3 | 2.2816 | 1.5356 | | 2.3063 | 1.4241 | 2.9577 | 3.1067 | 2.1943 | 1.1103 | 1.1018 | 2.2253 | O4 | 1.4186 | 1.4241 | 2.3063 | | 2.3083 | 2.0499 | 2.0999 | 2.1065 | 2.7742 | 3.2668 | 2.0566 | O5 | 1.4186 | 2.3063 | 1.4241 | 2.3083 | | 2.0999 | 2.0499 | 2.7742 | 2.1065 | 2.0566 | 3.2668 | H6 | 1.1105 | 3.1067 | 2.9577 | 2.0499 | 2.0999 | | 1.8118 | 3.8245 | 3.1264 | 3.9450 | 3.8138 | H7 | 1.1105 | 2.9577 | 3.1067 | 2.0999 | 2.0499 | 1.8118 | | 3.1264 | 3.8245 | 3.8138 | 3.9450 | H8 | 2.8179 | 1.1103 | 2.1943 | 2.1065 | 2.7742 | 3.8245 | 3.1264 | | 3.0928 | 2.4539 | 1.8019 | H9 | 2.8179 | 2.1943 | 1.1103 | 2.7742 | 2.1065 | 3.1264 | 3.8245 | 3.0928 | | 1.8019 | 2.4539 | H10 | 3.1861 | 2.2253 | 1.1018 | 3.2668 | 2.0566 | 3.9450 | 3.8138 | 2.4539 | 1.8019 | | 2.8231 | H11 | 3.1861 | 1.1018 | 2.2253 | 2.0566 | 3.2668 | 3.8138 | 3.9450 | 1.8019 | 2.4539 | 2.8231 | |
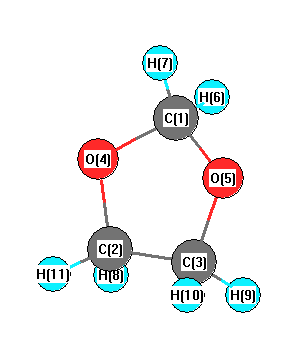 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
106.763 |
|
C1 |
O5 |
C3 |
106.763 |
| C2 |
C3 |
O5 |
102.318 |
|
C2 |
C3 |
H9 |
111.039 |
| C2 |
C3 |
H10 |
114.066 |
|
C3 |
C2 |
O4 |
102.318 |
| C3 |
C2 |
H8 |
111.039 |
|
C3 |
C2 |
H11 |
114.066 |
| O4 |
C1 |
O5 |
108.888 |
|
O4 |
C1 |
H6 |
107.664 |
| O4 |
C1 |
H7 |
111.673 |
|
O4 |
C2 |
H8 |
111.844 |
| O4 |
C2 |
H11 |
108.347 |
|
O5 |
C1 |
H6 |
111.673 |
| O5 |
C1 |
H7 |
107.664 |
|
O5 |
C3 |
H9 |
111.844 |
| O5 |
C3 |
H10 |
108.347 |
|
H6 |
C1 |
H7 |
109.316 |
| H8 |
C2 |
H11 |
109.095 |
|
H9 |
C3 |
H10 |
109.095 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.147 |
|
|
|
| 2 |
C |
-0.097 |
|
|
|
| 3 |
C |
-0.097 |
|
|
|
| 4 |
O |
-0.438 |
|
|
|
| 5 |
O |
-0.438 |
|
|
|
| 6 |
H |
0.146 |
|
|
|
| 7 |
H |
0.146 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.151 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.165 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.119 |
1.119 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.324 |
-0.263 |
0.001 |
| y |
-0.263 |
-34.077 |
-0.000 |
| z |
0.001 |
-0.000 |
-25.962 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.696 |
-0.263 |
0.001 |
| y |
-0.263 |
-6.435 |
-0.000 |
| z |
0.001 |
-0.000 |
5.739 |
|
| Polar |
|---|
| 3z2-r2 | 11.478 |
|---|
| x2-y2 | 4.754 |
|---|
| xy | -0.263 |
|---|
| xz | 0.001 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.229 |
-0.063 |
0.001 |
| y |
-0.063 |
5.052 |
-0.000 |
| z |
0.001 |
-0.000 |
6.694 |
<r2> (average value of r
2) Å
2
| <r2> |
93.967 |
| (<r2>)1/2 |
9.694 |
Jump to
S1C1
S1C3
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -268.026716 |
| Energy at 298.15K | -268.034818 |
| HF Energy | -268.026716 |
| Nuclear repulsion energy | 193.236300 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C2
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -268.026730 |
| Energy at 298.15K | -268.034833 |
| Nuclear repulsion energy | 193.232064 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3068 |
3018 |
25.35 |
|
|
|
| 2 |
A |
3033 |
2983 |
37.30 |
|
|
|
| 3 |
A |
2992 |
2942 |
71.98 |
|
|
|
| 4 |
A |
2948 |
2899 |
76.46 |
|
|
|
| 5 |
A |
2929 |
2881 |
44.66 |
|
|
|
| 6 |
A |
2894 |
2846 |
124.30 |
|
|
|
| 7 |
A |
1527 |
1502 |
4.68 |
|
|
|
| 8 |
A |
1497 |
1472 |
2.89 |
|
|
|
| 9 |
A |
1482 |
1458 |
1.66 |
|
|
|
| 10 |
A |
1398 |
1375 |
11.18 |
|
|
|
| 11 |
A |
1349 |
1326 |
7.75 |
|
|
|
| 12 |
A |
1306 |
1285 |
0.18 |
|
|
|
| 13 |
A |
1229 |
1209 |
5.22 |
|
|
|
| 14 |
A |
1206 |
1186 |
8.64 |
|
|
|
| 15 |
A |
1163 |
1144 |
57.96 |
|
|
|
| 16 |
A |
1143 |
1124 |
12.73 |
|
|
|
| 17 |
A |
1113 |
1095 |
23.54 |
|
|
|
| 18 |
A |
1104 |
1086 |
111.99 |
|
|
|
| 19 |
A |
1033 |
1016 |
29.48 |
|
|
|
| 20 |
A |
962 |
946 |
24.41 |
|
|
|
| 21 |
A |
936 |
920 |
8.75 |
|
|
|
| 22 |
A |
923 |
907 |
45.86 |
|
|
|
| 23 |
A |
880 |
865 |
25.57 |
|
|
|
| 24 |
A |
712 |
701 |
0.55 |
|
|
|
| 25 |
A |
629 |
619 |
3.46 |
|
|
|
| 26 |
A |
274 |
270 |
4.22 |
|
|
|
| 27 |
A |
24 |
24 |
9.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19876.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 19548.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| H1 |
-1.730 |
-1.208 |
0.494 |
| H2 |
-1.318 |
-0.789 |
-1.211 |
| C3 |
-1.018 |
-0.664 |
-0.150 |
| H4 |
-0.950 |
0.973 |
1.306 |
| H5 |
-1.559 |
1.491 |
-0.314 |
| C6 |
-0.866 |
0.816 |
0.210 |
| O7 |
0.297 |
-1.188 |
0.071 |
| O8 |
0.460 |
1.093 |
-0.254 |
| H9 |
1.652 |
0.005 |
1.086 |
| H10 |
1.985 |
-0.197 |
-0.690 |
| C11 |
1.194 |
-0.072 |
0.073 |
Atom - Atom Distances (Å)
| |
H1 |
H2 |
C3 |
H4 |
H5 |
C6 |
O7 |
O8 |
H9 |
H10 |
C11 |
| H1 | | 1.8030 | 1.1034 | 2.4553 | 2.8230 | 2.2190 | 2.0711 | 3.2635 | 3.6410 | 4.0285 | 3.1655 |
H2 | 1.8030 | | 1.1091 | 3.0950 | 2.4626 | 2.1906 | 2.1003 | 2.7601 | 3.8370 | 3.3962 | 2.9112 | C3 | 1.1034 | 1.1091 | | 2.1927 | 2.2284 | 1.5305 | 1.4331 | 2.2984 | 3.0171 | 3.0873 | 2.3014 | H4 | 2.4553 | 3.0950 | 2.1927 | | 1.8069 | 1.1111 | 2.7840 | 2.1064 | 2.7846 | 3.7378 | 2.6852 | H5 | 2.8230 | 2.4626 | 2.2284 | 1.8069 | | 1.1009 | 3.2821 | 2.0586 | 3.8050 | 3.9441 | 3.1898 | C6 | 2.2190 | 2.1906 | 1.5305 | 1.1111 | 1.1009 | | 2.3204 | 1.4313 | 2.7858 | 3.1564 | 2.2470 | O7 | 2.0711 | 2.1003 | 1.4331 | 2.7840 | 3.2821 | 2.3204 | | 2.3095 | 2.0699 | 2.1001 | 1.4317 | O8 | 3.2635 | 2.7601 | 2.2984 | 2.1064 | 2.0586 | 1.4313 | 2.3095 | | 2.0978 | 2.0450 | 1.4155 | H9 | 3.6410 | 3.8370 | 3.0171 | 2.7846 | 3.8050 | 2.7858 | 2.0699 | 2.0978 | | 1.8182 | 1.1136 | H10 | 4.0285 | 3.3962 | 3.0873 | 3.7378 | 3.9441 | 3.1564 | 2.1001 | 2.0450 | 1.8182 | | 1.1064 | C11 | 3.1655 | 2.9112 | 2.3014 | 2.6852 | 3.1898 | 2.2470 | 1.4317 | 1.4155 | 1.1136 | 1.1064 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O4 |
C2 |
35.606 |
|
C1 |
O5 |
C3 |
21.357 |
| C2 |
C3 |
O5 |
88.477 |
|
C2 |
C3 |
H9 |
130.985 |
| C2 |
C3 |
H10 |
96.477 |
|
C3 |
C2 |
O4 |
29.061 |
| C3 |
C2 |
H8 |
54.384 |
|
C3 |
C2 |
H11 |
46.942 |
| O4 |
C1 |
O5 |
39.265 |
|
O4 |
C1 |
H6 |
26.896 |
| O4 |
C1 |
H7 |
75.380 |
|
O4 |
C2 |
H8 |
41.681 |
| O4 |
C2 |
H11 |
53.004 |
|
O5 |
C1 |
H6 |
21.192 |
| O5 |
C1 |
H7 |
82.696 |
|
O5 |
C3 |
H9 |
91.745 |
| O5 |
C3 |
H10 |
94.415 |
|
H6 |
C1 |
H7 |
65.381 |
| H8 |
C2 |
H11 |
28.748 |
|
H9 |
C3 |
H10 |
34.632 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.159 |
|
|
|
| 2 |
H |
0.152 |
|
|
|
| 3 |
C |
-0.079 |
|
|
|
| 4 |
H |
0.151 |
|
|
|
| 5 |
H |
0.170 |
|
|
|
| 6 |
C |
-0.117 |
|
|
|
| 7 |
O |
-0.447 |
|
|
|
| 8 |
O |
-0.425 |
|
|
|
| 9 |
H |
0.136 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
C |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.108 |
0.329 |
0.370 |
1.213 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.980 |
-0.603 |
0.141 |
| y |
-0.603 |
-33.845 |
0.846 |
| z |
0.141 |
0.846 |
-29.558 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.722 |
-0.603 |
0.141 |
| y |
-0.603 |
-6.076 |
0.846 |
| z |
0.141 |
0.846 |
0.355 |
|
| Polar |
|---|
| 3z2-r2 | 0.710 |
|---|
| x2-y2 | 7.865 |
|---|
| xy | -0.603 |
|---|
| xz | 0.141 |
|---|
| yz | 0.846 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.668 |
-0.091 |
0.056 |
| y |
-0.091 |
5.072 |
0.093 |
| z |
0.056 |
0.093 |
5.175 |
<r2> (average value of r
2) Å
2
| <r2> |
93.977 |
| (<r2>)1/2 |
9.694 |
