return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5COOH (benzoic acid)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-419.293873
Energy at 298.15K-419.301231
HF Energy-418.324847
Nuclear repulsion energy407.713457
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3931 3631 130.26      
2 A' 3371 3114 2.26      
3 A' 3366 3110 3.61      
4 A' 3349 3093 16.70      
5 A' 3338 3084 12.67      
6 A' 3325 3071 0.56      
7 A' 1964 1814 387.55      
8 A' 1781 1646 18.60      
9 A' 1759 1625 4.98      
10 A' 1636 1511 4.76      
11 A' 1584 1463 26.20      
12 A' 1491 1377 171.95      
13 A' 1432 1323 4.79      
14 A' 1369 1265 5.52      
15 A' 1308 1208 231.94      
16 A' 1276 1178 46.83      
17 A' 1244 1149 3.44      
18 A' 1204 1112 52.88      
19 A' 1170 1081 21.88      
20 A' 1113 1028 7.45      
21 A' 1071 990 0.63      
22 A' 825 762 16.52      
23 A' 675 623 62.03      
24 A' 661 611 0.06      
25 A' 525 485 5.05      
26 A' 404 373 6.28      
27 A' 230 213 1.52      
28 A" 1039 960 0.11      
29 A" 1031 952 0.08      
30 A" 1005 928 1.22      
31 A" 922 852 0.04      
32 A" 860 794 0.72      
33 A" 776 716 173.32      
34 A" 690 638 4.82      
35 A" 629 581 75.97      
36 A" 453 419 8.05      
37 A" 433 400 0.42      
38 A" 169 156 0.59      
39 A" 65 60 1.09      

Unscaled Zero Point Vibrational Energy (zpe) 26735.9 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 24695.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.13136 0.04141 0.03148

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.217 0.000
C2 1.269 -0.348 0.000
C3 1.411 -1.723 0.000
C4 0.287 -2.534 0.000
C5 -0.980 -1.971 0.000
C6 -1.127 -0.595 0.000
C7 -0.100 1.695 0.000
O8 0.838 2.445 0.000
O9 -1.364 2.140 0.000
H10 2.129 0.301 0.000
H11 2.396 -2.162 0.000
H12 0.398 -3.607 0.000
H13 -1.853 -2.604 0.000
H14 -2.108 -0.152 0.000
H15 -1.318 3.102 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 O9 H10 H11 H12 H13 H14 H15
C11.38932.39872.76592.39741.38931.48162.38022.35752.13073.37643.84473.37502.14023.1719
C21.38931.38232.39702.77382.40932.45932.82573.62231.07702.13573.37393.85213.38324.3122
C32.39871.38231.38682.40402.77773.73714.20684.75622.14721.07822.13973.38093.85415.5432
C42.76592.39701.38681.38592.39974.24695.00934.95703.38112.14181.07882.14063.37785.8600
C52.39742.77382.40401.38591.38393.77044.77534.12913.85063.38112.13871.07832.14075.0843
C61.38932.40932.77772.39971.38392.51023.61972.74553.37733.85593.37602.13591.07643.7020
C71.48162.45933.73714.24693.77042.51021.20041.33962.62914.59435.32564.64282.72851.8609
O82.38022.82574.20685.00934.77533.61971.20042.22212.50274.86336.06795.72103.92702.2538
O92.35753.62234.75624.95704.12912.74551.33962.22213.94725.71336.01114.76942.41010.9628
H102.13071.07702.14723.38113.85063.37732.62912.50273.94722.47754.27454.92894.26144.4417
H113.37642.13571.07822.14183.38113.85594.59434.86335.71332.47752.46604.27164.93236.4424
H123.84473.37392.13971.07882.13873.37605.32566.06796.01114.27452.46602.46394.26826.9250
H133.37503.85213.38092.14061.07832.13594.64285.72104.76944.92894.27162.46392.46545.7309
H142.14023.38323.85413.37782.14071.07642.72853.92702.41014.26144.93234.26822.46543.3483
H153.17194.31225.54325.86005.08433.70201.86092.25380.96284.44176.44246.92505.73093.3483

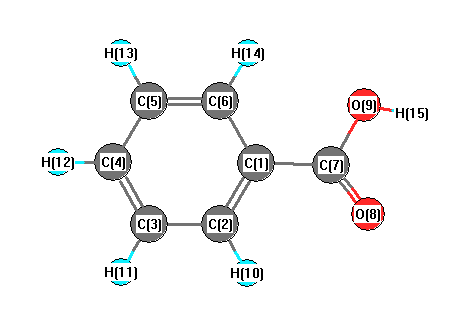
picture of benzoic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.876 C1 C2 H10 118.981
C1 C6 C5 119.654 C1 C6 H14 119.917
C1 C7 O8 124.776 C1 C7 O9 113.268
C2 C1 C6 120.249 C2 C1 C7 117.849
C2 C3 C4 119.914 C2 C3 H11 119.944
C3 C2 H10 121.143 C3 C4 C5 120.225
C3 C4 H12 119.901 C4 C3 H11 120.142
C4 C5 C6 120.082 C4 C5 H13 120.090
C5 C4 H12 119.874 C5 C6 H14 120.430
C6 C1 C7 121.902 C6 C5 H13 119.828
C7 O9 H15 106.709 O8 C7 O9 121.956
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability