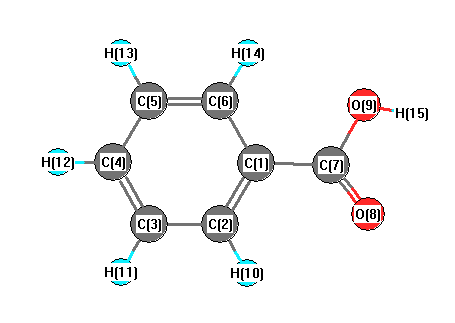
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3732 |
3571 |
81.15 |
|
|
|
| 2 |
A' |
3240 |
3100 |
3.13 |
|
|
|
| 3 |
A' |
3235 |
3095 |
5.45 |
|
|
|
| 4 |
A' |
3220 |
3081 |
18.18 |
|
|
|
| 5 |
A' |
3210 |
3071 |
13.20 |
|
|
|
| 6 |
A' |
3198 |
3059 |
0.39 |
|
|
|
| 7 |
A' |
1839 |
1759 |
302.34 |
|
|
|
| 8 |
A' |
1675 |
1602 |
18.24 |
|
|
|
| 9 |
A' |
1654 |
1583 |
4.34 |
|
|
|
| 10 |
A' |
1544 |
1477 |
2.63 |
|
|
|
| 11 |
A' |
1499 |
1434 |
18.36 |
|
|
|
| 12 |
A' |
1406 |
1345 |
140.19 |
|
|
|
| 13 |
A' |
1388 |
1328 |
4.43 |
|
|
|
| 14 |
A' |
1349 |
1290 |
5.17 |
|
|
|
| 15 |
A' |
1227 |
1174 |
157.63 |
|
|
|
| 16 |
A' |
1202 |
1150 |
107.63 |
|
|
|
| 17 |
A' |
1194 |
1143 |
0.50 |
|
|
|
| 18 |
A' |
1138 |
1088 |
49.45 |
|
|
|
| 19 |
A' |
1109 |
1061 |
39.99 |
|
|
|
| 20 |
A' |
1058 |
1012 |
9.78 |
|
|
|
| 21 |
A' |
1019 |
975 |
1.09 |
|
|
|
| 22 |
A' |
781 |
747 |
12.45 |
|
|
|
| 23 |
A' |
637 |
609 |
54.32 |
|
|
|
| 24 |
A' |
628 |
601 |
0.05 |
|
|
|
| 25 |
A' |
498 |
477 |
5.79 |
|
|
|
| 26 |
A' |
383 |
367 |
5.50 |
|
|
|
| 27 |
A' |
213 |
203 |
1.43 |
|
|
|
| 28 |
A" |
1011 |
967 |
0.10 |
|
|
|
| 29 |
A" |
988 |
946 |
0.12 |
|
|
|
| 30 |
A" |
957 |
916 |
1.55 |
|
|
|
| 31 |
A" |
868 |
830 |
0.06 |
|
|
|
| 32 |
A" |
820 |
784 |
0.00 |
|
|
|
| 33 |
A" |
732 |
700 |
142.07 |
|
|
|
| 34 |
A" |
705 |
674 |
5.18 |
|
|
|
| 35 |
A" |
618 |
591 |
62.39 |
|
|
|
| 36 |
A" |
434 |
415 |
7.24 |
|
|
|
| 37 |
A" |
413 |
395 |
0.41 |
|
|
|
| 38 |
A" |
159 |
152 |
0.62 |
|
|
|
| 39 |
A" |
68 |
65 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25525.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 24419.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.022 |
|
|
|
| 2 |
C |
-0.174 |
|
|
|
| 3 |
C |
-0.172 |
|
|
|
| 4 |
C |
-0.149 |
|
|
|
| 5 |
C |
-0.171 |
|
|
|
| 6 |
C |
-0.178 |
|
|
|
| 7 |
C |
0.562 |
|
|
|
| 8 |
O |
-0.480 |
|
|
|
| 9 |
O |
-0.595 |
|
|
|
| 10 |
H |
0.199 |
|
|
|
| 11 |
H |
0.173 |
|
|
|
| 12 |
H |
0.172 |
|
|
|
| 13 |
H |
0.172 |
|
|
|
| 14 |
H |
0.195 |
|
|
|
| 15 |
H |
0.426 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.217 |
-1.469 |
0.000 |
1.907 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.727 |
-5.802 |
0.000 |
| y |
-5.802 |
-43.811 |
0.000 |
| z |
0.000 |
0.000 |
-53.135 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.746 |
-5.802 |
0.000 |
| y |
-5.802 |
6.620 |
0.000 |
| z |
0.000 |
0.000 |
-7.366 |
|
| Polar |
|---|
| 3z2-r2 | -14.732 |
|---|
| x2-y2 | -3.917 |
|---|
| xy | -5.802 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.468 |
-0.220 |
0.000 |
| y |
-0.220 |
15.671 |
0.000 |
| z |
0.000 |
0.000 |
4.124 |
<r2> (average value of r
2) Å
2
| <r2> |
328.597 |
| (<r2>)1/2 |
18.127 |