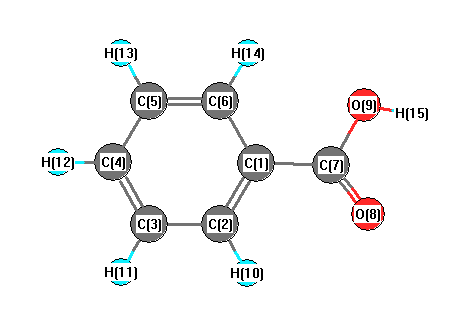
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3575 |
3516 |
52.33 |
|
|
|
| 2 |
A' |
3158 |
3106 |
3.54 |
|
|
|
| 3 |
A' |
3152 |
3100 |
7.22 |
|
|
|
| 4 |
A' |
3139 |
3088 |
17.99 |
|
|
|
| 5 |
A' |
3129 |
3078 |
12.93 |
|
|
|
| 6 |
A' |
3118 |
3066 |
0.29 |
|
|
|
| 7 |
A' |
1764 |
1735 |
253.45 |
|
|
|
| 8 |
A' |
1618 |
1592 |
16.34 |
|
|
|
| 9 |
A' |
1599 |
1572 |
3.89 |
|
|
|
| 10 |
A' |
1495 |
1470 |
1.81 |
|
|
|
| 11 |
A' |
1453 |
1429 |
14.68 |
|
|
|
| 12 |
A' |
1375 |
1353 |
3.11 |
|
|
|
| 13 |
A' |
1355 |
1332 |
105.84 |
|
|
|
| 14 |
A' |
1307 |
1285 |
5.69 |
|
|
|
| 15 |
A' |
1188 |
1168 |
89.24 |
|
|
|
| 16 |
A' |
1164 |
1145 |
123.21 |
|
|
|
| 17 |
A' |
1161 |
1142 |
11.91 |
|
|
|
| 18 |
A' |
1095 |
1077 |
39.69 |
|
|
|
| 19 |
A' |
1073 |
1055 |
79.45 |
|
|
|
| 20 |
A' |
1029 |
1012 |
13.69 |
|
|
|
| 21 |
A' |
990 |
974 |
1.74 |
|
|
|
| 22 |
A' |
756 |
743 |
8.37 |
|
|
|
| 23 |
A' |
614 |
604 |
47.06 |
|
|
|
| 24 |
A' |
611 |
601 |
1.74 |
|
|
|
| 25 |
A' |
482 |
474 |
6.12 |
|
|
|
| 26 |
A' |
373 |
367 |
5.42 |
|
|
|
| 27 |
A' |
207 |
204 |
1.29 |
|
|
|
| 28 |
A" |
968 |
952 |
0.15 |
|
|
|
| 29 |
A" |
943 |
928 |
0.18 |
|
|
|
| 30 |
A" |
917 |
902 |
1.53 |
|
|
|
| 31 |
A" |
836 |
822 |
0.10 |
|
|
|
| 32 |
A" |
788 |
775 |
0.37 |
|
|
|
| 33 |
A" |
705 |
694 |
122.77 |
|
|
|
| 34 |
A" |
684 |
672 |
9.34 |
|
|
|
| 35 |
A" |
606 |
596 |
49.99 |
|
|
|
| 36 |
A" |
420 |
413 |
6.40 |
|
|
|
| 37 |
A" |
399 |
393 |
0.49 |
|
|
|
| 38 |
A" |
154 |
151 |
0.65 |
|
|
|
| 39 |
A" |
69 |
68 |
0.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24733.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 24325.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.072 |
|
|
|
| 2 |
C |
-0.160 |
|
|
|
| 3 |
C |
-0.147 |
|
|
|
| 4 |
C |
-0.125 |
|
|
|
| 5 |
C |
-0.145 |
|
|
|
| 6 |
C |
-0.165 |
|
|
|
| 7 |
C |
0.483 |
|
|
|
| 8 |
O |
-0.443 |
|
|
|
| 9 |
O |
-0.547 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.148 |
|
|
|
| 12 |
H |
0.147 |
|
|
|
| 13 |
H |
0.147 |
|
|
|
| 14 |
H |
0.165 |
|
|
|
| 15 |
H |
0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.160 |
-1.406 |
0.000 |
1.823 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.984 |
-5.580 |
-0.010 |
| y |
-5.580 |
-44.078 |
-0.006 |
| z |
-0.010 |
-0.006 |
-53.170 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.640 |
-5.580 |
-0.010 |
| y |
-5.580 |
6.499 |
-0.006 |
| z |
-0.010 |
-0.006 |
-7.139 |
|
| Polar |
|---|
| 3z2-r2 | -14.278 |
|---|
| x2-y2 | -3.906 |
|---|
| xy | -5.580 |
|---|
| xz | -0.010 |
|---|
| yz | -0.006 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.872 |
-0.342 |
-0.002 |
| y |
-0.342 |
16.530 |
0.000 |
| z |
-0.002 |
0.000 |
4.168 |
<r2> (average value of r
2) Å
2
| <r2> |
332.344 |
| (<r2>)1/2 |
18.230 |