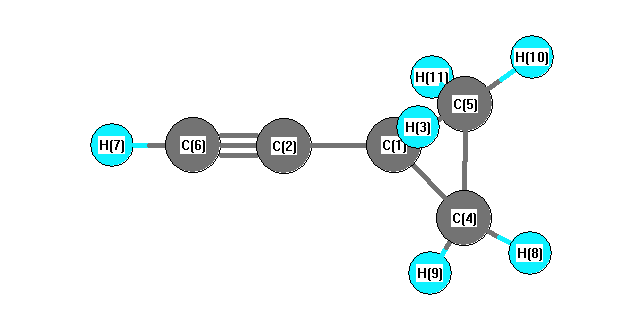
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3503 |
3351 |
64.88 |
|
|
|
| 2 |
A |
3265 |
3123 |
12.23 |
|
|
|
| 3 |
A |
3173 |
3036 |
8.34 |
|
|
|
| 4 |
A |
3168 |
3031 |
5.97 |
|
|
|
| 5 |
A |
2243 |
2145 |
16.05 |
|
|
|
| 6 |
A |
1527 |
1461 |
0.42 |
|
|
|
| 7 |
A |
1402 |
1341 |
4.02 |
|
|
|
| 8 |
A |
1238 |
1185 |
1.17 |
|
|
|
| 9 |
A |
1150 |
1100 |
0.67 |
|
|
|
| 10 |
A |
1072 |
1025 |
5.38 |
|
|
|
| 11 |
A |
989 |
946 |
22.79 |
|
|
|
| 12 |
A |
821 |
786 |
1.40 |
|
|
|
| 13 |
A |
769 |
736 |
2.20 |
|
|
|
| 14 |
A |
641 |
613 |
48.30 |
|
|
|
| 15 |
A |
511 |
488 |
5.16 |
|
|
|
| 16 |
A |
206 |
197 |
0.79 |
|
|
|
| 17 |
A |
3251 |
3110 |
0.08 |
|
|
|
| 18 |
A |
3167 |
3029 |
17.09 |
|
|
|
| 19 |
A |
1486 |
1422 |
2.27 |
|
|
|
| 20 |
A |
1211 |
1159 |
0.58 |
|
|
|
| 21 |
A |
1122 |
1073 |
1.76 |
|
|
|
| 22 |
A |
1081 |
1034 |
6.97 |
|
|
|
| 23 |
A |
919 |
879 |
2.07 |
|
|
|
| 24 |
A |
835 |
799 |
8.17 |
|
|
|
| 25 |
A |
575 |
550 |
56.36 |
|
|
|
| 26 |
A |
527 |
504 |
0.70 |
|
|
|
| 27 |
A |
213 |
204 |
0.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20030.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 19163.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.187 |
|
|
-0.053 |
| 2 |
C |
0.384 |
|
|
0.000 |
| 3 |
H |
0.221 |
|
|
0.150 |
| 4 |
C |
-0.365 |
|
|
-0.311 |
| 5 |
C |
-0.365 |
|
|
-0.311 |
| 6 |
C |
-0.683 |
|
|
-0.393 |
| 7 |
H |
0.231 |
|
|
0.271 |
| 8 |
H |
0.195 |
|
|
0.147 |
| 9 |
H |
0.186 |
|
|
0.175 |
| 10 |
H |
0.195 |
|
|
0.147 |
| 11 |
H |
0.186 |
|
|
0.175 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.699 |
-1.176 |
0.000 |
1.368 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.809 |
0.375 |
0.000 |
0.891 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.311 |
2.604 |
0.000 |
| y |
2.604 |
-26.723 |
0.000 |
| z |
0.000 |
0.000 |
-30.693 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.397 |
2.604 |
0.000 |
| y |
2.604 |
2.779 |
0.000 |
| z |
0.000 |
0.000 |
-3.176 |
|
| Polar |
|---|
| 3z2-r2 | -6.352 |
|---|
| x2-y2 | -1.588 |
|---|
| xy | 2.604 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
120.446 |
| (<r2>)1/2 |
10.975 |