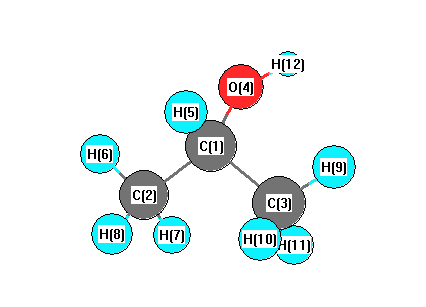
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3809 |
3620 |
11.10 |
|
|
|
| 2 |
A |
3170 |
3013 |
21.91 |
|
|
|
| 3 |
A |
3163 |
3005 |
45.22 |
|
|
|
| 4 |
A |
3156 |
2999 |
4.61 |
|
|
|
| 5 |
A |
3137 |
2981 |
31.59 |
|
|
|
| 6 |
A |
3079 |
2926 |
11.59 |
|
|
|
| 7 |
A |
3062 |
2910 |
21.70 |
|
|
|
| 8 |
A |
2998 |
2849 |
65.43 |
|
|
|
| 9 |
A |
1538 |
1461 |
5.85 |
|
|
|
| 10 |
A |
1526 |
1450 |
4.81 |
|
|
|
| 11 |
A |
1515 |
1439 |
2.08 |
|
|
|
| 12 |
A |
1509 |
1434 |
0.92 |
|
|
|
| 13 |
A |
1460 |
1388 |
24.55 |
|
|
|
| 14 |
A |
1437 |
1365 |
22.01 |
|
|
|
| 15 |
A |
1412 |
1342 |
1.67 |
|
|
|
| 16 |
A |
1395 |
1326 |
18.18 |
|
|
|
| 17 |
A |
1301 |
1236 |
59.89 |
|
|
|
| 18 |
A |
1212 |
1152 |
46.01 |
|
|
|
| 19 |
A |
1186 |
1127 |
15.21 |
|
|
|
| 20 |
A |
1110 |
1055 |
25.26 |
|
|
|
| 21 |
A |
997 |
948 |
32.88 |
|
|
|
| 22 |
A |
962 |
915 |
1.75 |
|
|
|
| 23 |
A |
937 |
890 |
0.51 |
|
|
|
| 24 |
A |
846 |
804 |
2.69 |
|
|
|
| 25 |
A |
484 |
460 |
8.20 |
|
|
|
| 26 |
A |
417 |
396 |
10.58 |
|
|
|
| 27 |
A |
361 |
343 |
3.04 |
|
|
|
| 28 |
A |
323 |
307 |
119.28 |
|
|
|
| 29 |
A |
273 |
259 |
1.20 |
|
|
|
| 30 |
A |
228 |
217 |
4.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24001.0 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 22808.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.092 |
|
|
|
| 2 |
C |
-0.534 |
|
|
|
| 3 |
C |
-0.516 |
|
|
|
| 4 |
O |
-0.630 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
| 7 |
H |
0.182 |
|
|
|
| 8 |
H |
0.171 |
|
|
|
| 9 |
H |
0.186 |
|
|
|
| 10 |
H |
0.166 |
|
|
|
| 11 |
H |
0.179 |
|
|
|
| 12 |
H |
0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.183 |
-0.640 |
0.817 |
1.574 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.899 |
-2.734 |
-0.271 |
| y |
-2.734 |
-26.401 |
1.090 |
| z |
-0.271 |
1.090 |
-26.401 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.502 |
-2.734 |
-0.271 |
| y |
-2.734 |
-0.752 |
1.090 |
| z |
-0.271 |
1.090 |
-0.751 |
|
| Polar |
|---|
| 3z2-r2 | -1.502 |
|---|
| x2-y2 | 1.503 |
|---|
| xy | -2.734 |
|---|
| xz | -0.271 |
|---|
| yz | 1.090 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.788 |
-0.175 |
-0.070 |
| y |
-0.175 |
5.522 |
-0.042 |
| z |
-0.070 |
-0.042 |
4.987 |
<r2> (average value of r
2) Å
2
| <r2> |
88.532 |
| (<r2>)1/2 |
9.409 |