Jump to
S2C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.942371 |
| Energy at 298.15K | -192.948266 |
| HF Energy | -192.942371 |
| Nuclear repulsion energy | 119.412411 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3198 |
3041 |
6.53 |
71.37 |
0.60 |
0.75 |
| 2 |
A |
3136 |
2982 |
0.00 |
10.12 |
0.75 |
0.86 |
| 3 |
A |
3074 |
2923 |
5.56 |
205.09 |
0.01 |
0.02 |
| 4 |
A |
1858 |
1767 |
160.72 |
6.70 |
0.46 |
0.63 |
| 5 |
A |
1495 |
1422 |
25.28 |
30.63 |
0.73 |
0.84 |
| 6 |
A |
1490 |
1417 |
0.00 |
30.77 |
0.75 |
0.86 |
| 7 |
A |
1407 |
1338 |
18.07 |
3.09 |
0.68 |
0.81 |
| 8 |
A |
1094 |
1041 |
0.07 |
4.08 |
0.56 |
0.72 |
| 9 |
A |
891 |
847 |
0.00 |
3.57 |
0.75 |
0.86 |
| 10 |
A |
807 |
767 |
0.80 |
10.96 |
0.16 |
0.28 |
| 11 |
A |
378 |
359 |
1.31 |
0.34 |
0.49 |
0.66 |
| 12 |
A |
41 |
39 |
0.00 |
0.07 |
0.75 |
0.86 |
| 13 |
B |
3197 |
3040 |
9.20 |
56.01 |
0.75 |
0.86 |
| 14 |
B |
3142 |
2988 |
16.72 |
105.97 |
0.75 |
0.86 |
| 15 |
B |
3067 |
2917 |
1.04 |
0.61 |
0.75 |
0.86 |
| 16 |
B |
1513 |
1439 |
23.33 |
0.36 |
0.75 |
0.86 |
| 17 |
B |
1485 |
1413 |
0.28 |
1.63 |
0.75 |
0.86 |
| 18 |
B |
1415 |
1346 |
78.28 |
5.08 |
0.75 |
0.86 |
| 19 |
B |
1258 |
1197 |
67.81 |
0.70 |
0.75 |
0.86 |
| 20 |
B |
1131 |
1075 |
5.30 |
0.01 |
0.75 |
0.86 |
| 21 |
B |
900 |
856 |
3.96 |
3.40 |
0.75 |
0.86 |
| 22 |
B |
534 |
508 |
14.85 |
1.65 |
0.75 |
0.86 |
| 23 |
B |
489 |
465 |
0.22 |
1.08 |
0.75 |
0.86 |
| 24 |
B |
138 |
131 |
0.14 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 18568.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 17658.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.185 |
| O2 |
0.000 |
0.000 |
1.397 |
| C3 |
0.000 |
1.285 |
-0.613 |
| C4 |
0.000 |
-1.285 |
-0.613 |
| H5 |
0.001 |
2.141 |
0.064 |
| H6 |
-0.001 |
-2.141 |
0.064 |
| H7 |
0.880 |
1.331 |
-1.266 |
| H8 |
-0.881 |
1.332 |
-1.265 |
| H9 |
-0.880 |
-1.331 |
-1.266 |
| H10 |
0.881 |
-1.332 |
-1.265 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.2118 | 1.5124 | 1.5124 | 2.1448 | 2.1448 | 2.1566 | 2.1565 | 2.1566 | 2.1565 |
O2 | 1.2118 | | 2.3853 | 2.3853 | 2.5225 | 2.5225 | 3.1041 | 3.1037 | 3.1041 | 3.1037 | C3 | 1.5124 | 2.3853 | | 2.5698 | 1.0913 | 3.4924 | 1.0966 | 1.0966 | 2.8362 | 2.8367 | C4 | 1.5124 | 2.3853 | 2.5698 | | 3.4924 | 1.0913 | 2.8362 | 2.8367 | 1.0966 | 1.0966 | H5 | 2.1448 | 2.5225 | 1.0913 | 3.4924 | | 4.2827 | 1.7879 | 1.7879 | 3.8210 | 3.8211 | H6 | 2.1448 | 2.5225 | 3.4924 | 1.0913 | 4.2827 | | 3.8210 | 3.8211 | 1.7879 | 1.7879 | H7 | 2.1566 | 3.1041 | 1.0966 | 2.8362 | 1.7879 | 3.8210 | | 1.7603 | 3.1912 | 2.6628 | H8 | 2.1565 | 3.1037 | 1.0966 | 2.8367 | 1.7879 | 3.8211 | 1.7603 | | 2.6628 | 3.1930 | H9 | 2.1566 | 3.1041 | 2.8362 | 1.0966 | 3.8210 | 1.7879 | 3.1912 | 2.6628 | | 1.7603 | H10 | 2.1565 | 3.1037 | 2.8367 | 1.0966 | 3.8211 | 1.7879 | 2.6628 | 3.1930 | 1.7603 | |
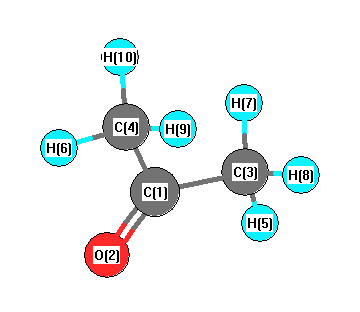 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
109.869 |
|
C1 |
C3 |
H7 |
110.481 |
| C1 |
C3 |
H8 |
110.479 |
|
C1 |
C4 |
H6 |
109.869 |
| C1 |
C4 |
H9 |
110.481 |
|
C1 |
C4 |
H10 |
110.479 |
| O2 |
C1 |
C3 |
121.835 |
|
O2 |
C1 |
C4 |
121.835 |
| C3 |
C1 |
C4 |
116.330 |
|
H5 |
C3 |
H7 |
109.601 |
| H5 |
C3 |
H8 |
109.601 |
|
H6 |
C4 |
H9 |
109.601 |
| H6 |
C4 |
H10 |
109.601 |
|
H7 |
C3 |
H8 |
106.757 |
| H9 |
C4 |
H10 |
106.757 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.453 |
|
|
|
| 2 |
O |
-0.427 |
|
|
|
| 3 |
C |
-0.604 |
|
|
|
| 4 |
C |
-0.604 |
|
|
|
| 5 |
H |
0.207 |
|
|
|
| 6 |
H |
0.207 |
|
|
|
| 7 |
H |
0.192 |
|
|
|
| 8 |
H |
0.192 |
|
|
|
| 9 |
H |
0.192 |
|
|
|
| 10 |
H |
0.192 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.855 |
2.855 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.826 |
-0.000 |
0.000 |
| y |
-0.000 |
-23.348 |
0.000 |
| z |
0.000 |
0.000 |
-28.072 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.884 |
-0.000 |
0.000 |
| y |
-0.000 |
2.601 |
0.000 |
| z |
0.000 |
0.000 |
-4.486 |
|
| Polar |
|---|
| 3z2-r2 | -8.971 |
|---|
| x2-y2 | -0.478 |
|---|
| xy | -0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.980 |
0.000 |
0.000 |
| y |
0.000 |
5.317 |
0.000 |
| z |
0.000 |
0.000 |
5.724 |
<r2> (average value of r
2) Å
2
| <r2> |
81.570 |
| (<r2>)1/2 |
9.032 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -192.824499 |
| Energy at 298.15K | -192.830384 |
| Nuclear repulsion energy | 117.686567 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3170 |
3014 |
11.48 |
|
|
|
| 2 |
A' |
3122 |
2969 |
24.40 |
|
|
|
| 3 |
A' |
3018 |
2870 |
48.87 |
|
|
|
| 4 |
A' |
1528 |
1453 |
8.77 |
|
|
|
| 5 |
A' |
1498 |
1424 |
8.53 |
|
|
|
| 6 |
A' |
1426 |
1356 |
1.74 |
|
|
|
| 7 |
A' |
1318 |
1254 |
6.44 |
|
|
|
| 8 |
A' |
1111 |
1057 |
0.46 |
|
|
|
| 9 |
A' |
1003 |
954 |
0.03 |
|
|
|
| 10 |
A' |
811 |
771 |
7.87 |
|
|
|
| 11 |
A' |
371 |
353 |
1.28 |
|
|
|
| 12 |
A' |
317 |
301 |
2.08 |
|
|
|
| 13 |
A' |
189 |
179 |
0.53 |
|
|
|
| 14 |
A" |
3169 |
3014 |
13.02 |
|
|
|
| 15 |
A" |
3123 |
2970 |
4.34 |
|
|
|
| 16 |
A" |
3013 |
2865 |
32.43 |
|
|
|
| 17 |
A" |
1504 |
1430 |
0.06 |
|
|
|
| 18 |
A" |
1485 |
1412 |
7.15 |
|
|
|
| 19 |
A" |
1398 |
1330 |
0.00 |
|
|
|
| 20 |
A" |
1169 |
1112 |
0.01 |
|
|
|
| 21 |
A" |
973 |
926 |
1.05 |
|
|
|
| 22 |
A" |
968 |
921 |
0.96 |
|
|
|
| 23 |
A" |
352 |
335 |
2.77 |
|
|
|
| 24 |
A" |
186 |
177 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18110.7 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 17223.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.181 |
0.270 |
0.000 |
| O2 |
-0.541 |
1.279 |
0.000 |
| C3 |
0.181 |
-0.591 |
1.293 |
| C4 |
0.181 |
-0.591 |
-1.293 |
| H5 |
-0.836 |
-0.676 |
1.690 |
| H6 |
-0.836 |
-0.676 |
-1.690 |
| H7 |
0.571 |
-1.587 |
1.122 |
| H8 |
0.803 |
-0.116 |
2.057 |
| H9 |
0.571 |
-1.587 |
-1.122 |
| H10 |
0.803 |
-0.116 |
-2.057 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.2401 | 1.5533 | 1.5533 | 2.1876 | 2.1876 | 2.2041 | 2.1836 | 2.2041 | 2.1836 |
O2 | 1.2401 | | 2.3846 | 2.3846 | 2.6012 | 2.6012 | 3.2717 | 2.8254 | 3.2717 | 2.8254 | C3 | 1.5533 | 2.3846 | | 2.5863 | 1.0946 | 3.1531 | 1.0832 | 1.0939 | 2.6417 | 3.4404 | C4 | 1.5533 | 2.3846 | 2.5863 | | 3.1531 | 1.0946 | 2.6417 | 3.4404 | 1.0832 | 1.0939 | H5 | 2.1876 | 2.6012 | 1.0946 | 3.1531 | | 3.3809 | 1.7692 | 1.7706 | 3.2739 | 4.1285 | H6 | 2.1876 | 2.6012 | 3.1531 | 1.0946 | 3.3809 | | 3.2739 | 4.1285 | 1.7692 | 1.7706 | H7 | 2.2041 | 3.2717 | 1.0832 | 2.6417 | 1.7692 | 3.2739 | | 1.7579 | 2.2445 | 3.5106 | H8 | 2.1836 | 2.8254 | 1.0939 | 3.4404 | 1.7706 | 4.1285 | 1.7579 | | 3.5106 | 4.1140 | H9 | 2.2041 | 3.2717 | 2.6417 | 1.0832 | 3.2739 | 1.7692 | 2.2445 | 3.5106 | | 1.7579 | H10 | 2.1836 | 2.8254 | 3.4404 | 1.0939 | 4.1285 | 1.7706 | 3.5106 | 4.1140 | 1.7579 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.244 |
|
|
|
| 2 |
O |
-0.255 |
|
|
|
| 3 |
C |
-0.560 |
|
|
|
| 4 |
C |
-0.560 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
H |
0.189 |
|
|
|
| 8 |
H |
0.197 |
|
|
|
| 9 |
H |
0.189 |
|
|
|
| 10 |
H |
0.197 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.481 |
-1.636 |
0.000 |
1.705 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.730 |
0.236 |
0.000 |
| y |
0.236 |
-26.395 |
0.000 |
| z |
0.000 |
0.000 |
-22.662 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.202 |
0.236 |
0.000 |
| y |
0.236 |
-2.199 |
0.000 |
| z |
0.000 |
0.000 |
3.401 |
|
| Polar |
|---|
| 3z2-r2 | 6.802 |
|---|
| x2-y2 | 0.665 |
|---|
| xy | 0.236 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.532 |
-0.517 |
0.000 |
| y |
-0.517 |
5.385 |
0.000 |
| z |
0.000 |
0.000 |
5.824 |
<r2> (average value of r
2) Å
2
| <r2> |
81.958 |
| (<r2>)1/2 |
9.053 |
