return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3PHCH3 (dimethylphosphine)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-420.942492
Energy at 298.15K-420.950295
HF Energy-420.525288
Nuclear repulsion energy112.664642
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3156 3006 15.78      
2 A' 3152 3001 30.28      
3 A' 3067 2921 16.37      
4 A' 2401 2286 116.34      
5 A' 1529 1456 1.53      
6 A' 1528 1456 14.33      
7 A' 1407 1340 1.38      
8 A' 1051 1001 35.53      
9 A' 1011 963 33.96      
10 A' 738 703 2.90      
11 A' 684 652 2.24      
12 A' 265 253 0.29      
13 A' 194 184 0.10      
14 A" 3156 3006 8.82      
15 A" 3152 3002 0.09      
16 A" 3068 2922 17.55      
17 A" 1521 1449 8.22      
18 A" 1515 1443 2.31      
19 A" 1391 1324 2.00      
20 A" 1052 1002 29.66      
21 A" 875 833 0.89      
22 A" 742 707 4.07      
23 A" 731 696 9.96      
24 A" 182 174 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 18784.6 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 17888.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.52826 0.23116 0.17794

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
P1 -0.038 -0.669 0.000
H2 1.360 -0.953 0.000
C3 -0.038 0.531 1.423
C4 -0.038 0.531 -1.423
H5 -1.029 0.992 1.505
H6 -1.029 0.992 -1.505
H7 0.156 -0.005 2.360
H8 0.156 -0.005 -2.360
H9 0.708 1.326 1.313
H10 0.708 1.326 -1.313

Atom - Atom Distances (Å)
  P1 H2 C3 C4 H5 H6 H7 H8 H9 H10
P11.42691.86171.86172.45142.45142.45922.45922.50282.5028
H21.42692.48662.48663.42953.42952.81392.81392.71062.7106
C31.86172.48662.84641.09663.12581.09623.82561.09632.9456
C41.86172.48662.84643.12581.09663.82561.09622.94561.0963
H52.45143.42951.09663.12583.01031.76924.16381.77953.3274
H62.45143.42953.12581.09663.01034.16381.76923.32741.7795
H72.45922.81391.09623.82561.76924.16384.71951.78113.9453
H82.45922.81393.82561.09624.16381.76924.71953.94531.7811
H92.50282.71061.09632.94561.77953.32741.78113.94532.6259
H102.50282.71062.94561.09633.32741.77953.94531.78112.6259

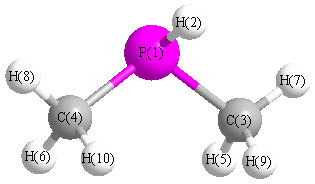
picture of dimethylphosphine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
P1 C3 H5 109.176 P1 C3 H7 109.770
P1 C3 H9 113.023 P1 C4 H6 109.176
P1 C4 H8 109.770 P1 C4 H10 113.023
H2 P1 C3 97.370 H2 P1 C4 97.370
C3 P1 C4 99.721 H5 C3 H7 107.579
H5 C3 H9 108.486 H6 C4 H8 107.579
H6 C4 H10 108.486 H7 C3 H9 108.655
H8 C4 H10 108.655
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability