Jump to
S1C2
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -498.726977 |
| Energy at 298.15K | |
| HF Energy | -498.460696 |
| Nuclear repulsion energy | 45.180115 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3219 |
3065 |
8.03 |
|
|
|
| 2 |
A' |
1469 |
1399 |
14.21 |
|
|
|
| 3 |
A' |
866 |
825 |
32.11 |
|
|
|
| 4 |
A' |
242 |
231 |
78.46 |
|
|
|
| 5 |
A" |
3363 |
3203 |
0.72 |
|
|
|
| 6 |
A" |
1037 |
987 |
0.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5098.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 4854.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.008 |
1.122 |
0.000 |
| Cl2 |
-0.008 |
-0.587 |
0.000 |
| H3 |
0.095 |
1.625 |
0.952 |
| H4 |
0.095 |
1.625 |
-0.952 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7098 | 1.0818 | 1.0818 |
Cl2 | 1.7098 | | 2.4107 | 2.4107 | H3 | 1.0818 | 2.4107 | | 1.9049 | H4 | 1.0818 | 2.4107 | 1.9049 | |
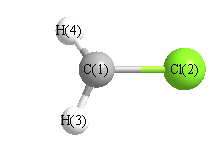 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
117.669 |
|
Br2 |
C1 |
H4 |
117.669 |
| H3 |
C1 |
H4 |
123.392 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -498.726960 |
| Energy at 298.15K | |
| HF Energy | -498.460569 |
| Nuclear repulsion energy | 45.195363 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3224 |
3070 |
7.11 |
|
|
|
| 2 |
A1 |
1468 |
1398 |
14.56 |
|
|
|
| 3 |
A1 |
868 |
827 |
31.22 |
|
|
|
| 4 |
B1 |
174i |
165i |
86.32 |
|
|
|
| 5 |
B2 |
3370 |
3209 |
0.44 |
|
|
|
| 6 |
B2 |
1033 |
984 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4894.4 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 4660.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.121 |
| Cl2 |
0.000 |
0.000 |
0.587 |
| H3 |
0.000 |
0.955 |
-1.629 |
| H4 |
0.000 |
-0.955 |
-1.629 |
Atom - Atom Distances (Å)
| |
C1 |
Cl2 |
H3 |
H4 |
| C1 | | 1.7086 | 1.0813 | 1.0813 |
Cl2 | 1.7086 | | 2.4133 | 2.4133 | H3 | 1.0813 | 2.4133 | | 1.9093 | H4 | 1.0813 | 2.4133 | 1.9093 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Br2 |
C1 |
H3 |
118.009 |
|
Br2 |
C1 |
H4 |
118.009 |
| H3 |
C1 |
H4 |
123.982 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
