return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (dimethylformamide)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-247.741383
Energy at 298.15K-247.749740
HF Energy-246.989320
Nuclear repulsion energy181.584589
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3239 3040 0.40      
2 A' 3195 2999 16.56      
3 A' 3094 2904 43.84      
4 A' 3086 2896 41.62      
5 A' 3051 2864 79.35      
6 A' 1864 1749 473.36      
7 A' 1600 1502 30.49      
8 A' 1567 1471 13.74      
9 A' 1528 1434 4.52      
10 A' 1505 1413 66.98      
11 A' 1489 1398 6.89      
12 A' 1476 1385 68.24      
13 A' 1343 1260 40.87      
14 A' 1139 1069 120.55      
15 A' 1118 1050 2.69      
16 A' 910 854 2.98      
17 A' 681 639 8.08      
18 A' 403 378 1.40      
19 A' 335 314 12.49      
20 A" 3153 2960 15.23      
21 A" 3146 2953 47.81      
22 A" 1558 1463 15.90      
23 A" 1536 1442 3.80      
24 A" 1212 1138 3.59      
25 A" 1164 1093 0.02      
26 A" 1052 988 0.75      
27 A" 332 312 22.03      
28 A" 229 215 2.75      
29 A" 164 154 0.55      
30 A" 83 77 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 23125.6 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 21705.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.29851 0.13972 0.09873

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.700 -0.820 0.000
O2 0.215 -1.937 0.000
N3 0.000 0.346 0.000
C4 -1.451 0.327 0.000
C5 0.658 1.636 0.000
H6 1.792 -0.655 0.000
H7 -1.775 -0.713 0.000
H8 -1.840 0.833 0.891
H9 -1.840 0.833 -0.891
H10 1.741 1.495 0.000
H11 0.381 2.214 -0.889
H12 0.381 2.214 0.889

Atom - Atom Distances (Å)
  C1 O2 N3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.21781.35972.43772.45591.10442.47803.15903.15902.53873.17743.1774
O21.21782.29272.81063.59992.03232.33663.56233.56233.75644.24814.2481
N31.35972.29271.45131.44802.05282.06722.10172.10172.08672.10372.1037
C42.43772.81061.45132.48233.38881.08911.09601.09603.39982.77642.7764
C52.45593.59991.44802.48232.55653.38202.77082.77081.09261.09641.0964
H61.10442.03232.05283.38882.55653.56824.02544.02542.15143.31893.3189
H72.47802.33662.06721.08913.38203.56821.78561.78564.15293.74263.7426
H83.15903.56232.10171.09602.77084.02541.78561.78133.74963.16342.6151
H93.15903.56232.10171.09602.77084.02541.78561.78133.74962.61513.1634
H102.53873.75642.08673.39981.09262.15144.15293.74963.74961.77721.7772
H113.17744.24812.10372.77641.09643.31893.74263.16342.61511.77721.7789
H123.17744.24812.10372.77641.09643.31893.74262.61513.16341.77721.7789

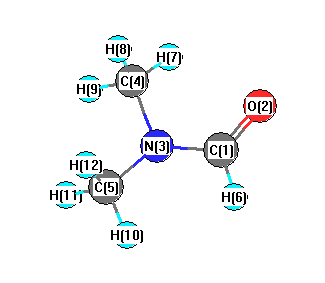
picture of dimethylformamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C4 120.239 C1 N3 C5 121.987
O2 C1 N3 125.529 O2 C1 H6 122.046
N3 C1 H6 112.426 N3 C4 H7 108.080
N3 C4 H8 110.416 N3 C4 H9 110.416
N3 C5 H10 109.645 N3 C5 H11 110.786
N3 C5 H12 110.786 C4 N3 C5 117.774
H7 C4 H8 109.606 H7 C4 H9 109.606
H8 C4 H9 108.706 H10 C5 H11 108.561
H10 C5 H12 108.561 H11 C5 H12 108.439
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability