Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -749.173744 |
| Energy at 298.15K | |
| HF Energy | -747.991005 |
| Nuclear repulsion energy | 566.377801 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
2028 |
1873 |
0.00 |
|
|
|
| 2 |
Ag |
1553 |
1434 |
0.00 |
|
|
|
| 3 |
Ag |
1453 |
1342 |
0.00 |
|
|
|
| 4 |
Ag |
1281 |
1184 |
0.00 |
|
|
|
| 5 |
Ag |
763 |
705 |
0.00 |
|
|
|
| 6 |
Ag |
619 |
572 |
0.00 |
|
|
|
| 7 |
Ag |
398 |
368 |
0.00 |
|
|
|
| 8 |
Ag |
370 |
342 |
0.00 |
|
|
|
| 9 |
Ag |
234 |
216 |
0.00 |
|
|
|
| 10 |
Au |
607 |
561 |
8.38 |
|
|
|
| 11 |
Au |
365 |
337 |
7.88 |
|
|
|
| 12 |
Au |
118 |
109 |
0.17 |
|
|
|
| 13 |
Au |
24i |
22i |
0.15 |
|
|
|
| 14 |
Bg |
647 |
598 |
0.00 |
|
|
|
| 15 |
Bg |
505 |
467 |
0.00 |
|
|
|
| 16 |
Bg |
194 |
179 |
0.00 |
|
|
|
| 17 |
Bu |
1953 |
1804 |
419.46 |
|
|
|
| 18 |
Bu |
1479 |
1366 |
420.49 |
|
|
|
| 19 |
Bu |
1318 |
1217 |
362.53 |
|
|
|
| 20 |
Bu |
1034 |
955 |
298.39 |
|
|
|
| 21 |
Bu |
657 |
607 |
6.19 |
|
|
|
| 22 |
Bu |
521 |
482 |
4.17 |
|
|
|
| 23 |
Bu |
303 |
280 |
7.62 |
|
|
|
| 24 |
Bu |
145 |
134 |
1.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9260.3 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 8553.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.156 |
1.840 |
0.000 |
| C2 |
0.465 |
0.559 |
0.000 |
| C3 |
-0.465 |
-0.559 |
0.000 |
| C4 |
-0.156 |
-1.840 |
0.000 |
| F5 |
1.058 |
2.785 |
0.000 |
| F6 |
-1.058 |
2.319 |
0.000 |
| F7 |
1.759 |
0.229 |
0.000 |
| F8 |
-1.759 |
-0.229 |
0.000 |
| F9 |
1.058 |
-2.319 |
0.000 |
| F10 |
-1.058 |
-2.785 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3179 | 2.4776 | 3.6930 | 1.3063 | 1.3046 | 2.2728 | 2.8187 | 4.2559 | 4.7810 |
C2 | 1.3179 | | 1.4535 | 2.4776 | 2.3035 | 2.3275 | 1.3355 | 2.3589 | 2.9385 | 3.6737 | C3 | 2.4776 | 1.4535 | | 1.3179 | 3.6737 | 2.9385 | 2.3589 | 1.3355 | 2.3275 | 2.3035 | C4 | 3.6930 | 2.4776 | 1.3179 | | 4.7810 | 4.2559 | 2.8187 | 2.2728 | 1.3046 | 1.3063 | F5 | 1.3063 | 2.3035 | 3.6737 | 4.7810 | | 2.1663 | 2.6500 | 4.1249 | 5.1037 | 5.9575 | F6 | 1.3046 | 2.3275 | 2.9385 | 4.2559 | 2.1663 | | 3.5075 | 2.6428 | 5.0981 | 5.1037 | F7 | 2.2728 | 1.3355 | 2.3589 | 2.8187 | 2.6500 | 3.5075 | | 3.5473 | 2.6428 | 4.1249 | F8 | 2.8187 | 2.3589 | 1.3355 | 2.2728 | 4.1249 | 2.6428 | 3.5473 | | 3.5075 | 2.6500 | F9 | 4.2559 | 2.9385 | 2.3275 | 1.3046 | 5.1037 | 5.0981 | 2.6428 | 3.5075 | | 2.1663 | F10 | 4.7810 | 3.6737 | 2.3035 | 1.3063 | 5.9575 | 5.1037 | 4.1249 | 2.6500 | 2.1663 | |
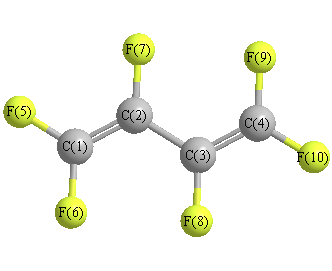 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.686 |
|
C1 |
C2 |
F7 |
117.865 |
| C2 |
C1 |
F5 |
122.746 |
|
C2 |
C1 |
F6 |
125.119 |
| C2 |
C3 |
C4 |
126.686 |
|
C2 |
C3 |
F8 |
115.449 |
| C3 |
C2 |
F7 |
115.449 |
|
C3 |
C4 |
F9 |
125.119 |
| C3 |
C4 |
F10 |
122.746 |
|
C4 |
C3 |
F8 |
117.865 |
| F5 |
C1 |
F6 |
112.135 |
|
F9 |
C4 |
F10 |
112.135 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -749.175284 |
| Energy at 298.15K | -749.176716 |
| HF Energy | -747.992148 |
| Nuclear repulsion energy | 570.618041 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2012 |
1858 |
75.58 |
|
|
|
| 2 |
A |
1526 |
1409 |
8.03 |
|
|
|
| 3 |
A |
1472 |
1359 |
208.71 |
|
|
|
| 4 |
A |
1241 |
1147 |
302.67 |
|
|
|
| 5 |
A |
763 |
704 |
0.27 |
|
|
|
| 6 |
A |
700 |
647 |
1.62 |
|
|
|
| 7 |
A |
568 |
524 |
0.08 |
|
|
|
| 8 |
A |
501 |
463 |
0.57 |
|
|
|
| 9 |
A |
399 |
368 |
1.10 |
|
|
|
| 10 |
A |
268 |
248 |
0.24 |
|
|
|
| 11 |
A |
200 |
185 |
0.39 |
|
|
|
| 12 |
A |
103 |
95 |
0.37 |
|
|
|
| 13 |
A |
55 |
51 |
0.00 |
|
|
|
| 14 |
B |
1986 |
1835 |
260.81 |
|
|
|
| 15 |
B |
1475 |
1362 |
280.74 |
|
|
|
| 16 |
B |
1316 |
1216 |
117.67 |
|
|
|
| 17 |
B |
1055 |
975 |
195.87 |
|
|
|
| 18 |
B |
661 |
610 |
6.92 |
|
|
|
| 19 |
B |
657 |
606 |
5.20 |
|
|
|
| 20 |
B |
596 |
551 |
5.14 |
|
|
|
| 21 |
B |
437 |
403 |
5.36 |
|
|
|
| 22 |
B |
311 |
287 |
5.12 |
|
|
|
| 23 |
B |
221 |
204 |
4.96 |
|
|
|
| 24 |
B |
115 |
107 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9317.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 8606.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.160 |
1.549 |
-0.373 |
| C2 |
0.146 |
0.710 |
0.593 |
| C3 |
-0.146 |
-0.710 |
0.593 |
| C4 |
0.160 |
-1.549 |
-0.373 |
| F5 |
0.146 |
2.816 |
-0.367 |
| F6 |
-0.788 |
1.198 |
-1.463 |
| F7 |
0.762 |
1.179 |
1.683 |
| F8 |
-0.762 |
-1.179 |
1.683 |
| F9 |
0.788 |
-1.198 |
-1.463 |
| F10 |
-0.146 |
-2.816 |
-0.367 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3150 | 2.4573 | 3.1148 | 1.3028 | 1.3069 | 2.2833 | 3.4687 | 3.1040 | 4.3649 |
C2 | 1.3150 | | 1.4506 | 2.4573 | 2.3136 | 2.3105 | 1.3371 | 2.3628 | 2.8777 | 3.6659 | C3 | 2.4573 | 1.4506 | | 1.3150 | 3.6659 | 2.8777 | 2.3628 | 1.3371 | 2.3105 | 2.3136 | C4 | 3.1148 | 2.4573 | 1.3150 | | 4.3649 | 3.1040 | 3.4687 | 2.2833 | 1.3069 | 1.3028 | F5 | 1.3028 | 2.3136 | 3.6659 | 4.3649 | | 2.1662 | 2.6947 | 4.5807 | 4.2099 | 5.6388 | F6 | 1.3069 | 2.3105 | 2.8777 | 3.1040 | 2.1662 | | 3.5078 | 3.9435 | 2.8681 | 4.2099 | F7 | 2.2833 | 1.3371 | 2.3628 | 3.4687 | 2.6947 | 3.5078 | | 2.8071 | 3.9435 | 4.5807 | F8 | 3.4687 | 2.3628 | 1.3371 | 2.2833 | 4.5807 | 3.9435 | 2.8071 | | 3.5078 | 2.6947 | F9 | 3.1040 | 2.8777 | 2.3105 | 1.3069 | 4.2099 | 2.8681 | 3.9435 | 3.5078 | | 2.1662 | F10 | 4.3649 | 3.6659 | 2.3136 | 1.3028 | 5.6388 | 4.2099 | 4.5807 | 2.6947 | 2.1662 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.301 |
|
C1 |
C2 |
F7 |
118.847 |
| C2 |
C1 |
F5 |
124.203 |
|
C2 |
C1 |
F6 |
123.580 |
| C2 |
C3 |
C4 |
125.301 |
|
C2 |
C3 |
F8 |
115.846 |
| C3 |
C2 |
F7 |
115.846 |
|
C3 |
C4 |
F9 |
123.580 |
| C3 |
C4 |
F10 |
124.203 |
|
C4 |
C3 |
F8 |
118.847 |
| F5 |
C1 |
F6 |
112.207 |
|
F9 |
C4 |
F10 |
112.207 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
