Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -750.702329 |
| Energy at 298.15K | -750.703371 |
| HF Energy | -750.702329 |
| Nuclear repulsion energy | 563.323818 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1877 |
1785 |
0.00 |
|
|
|
| 2 |
Ag |
1477 |
1404 |
0.00 |
|
|
|
| 3 |
Ag |
1383 |
1315 |
0.00 |
|
|
|
| 4 |
Ag |
1225 |
1165 |
0.00 |
|
|
|
| 5 |
Ag |
732 |
696 |
0.00 |
|
|
|
| 6 |
Ag |
592 |
563 |
0.00 |
|
|
|
| 7 |
Ag |
382 |
363 |
0.00 |
|
|
|
| 8 |
Ag |
351 |
334 |
0.00 |
|
|
|
| 9 |
Ag |
218 |
208 |
0.00 |
|
|
|
| 10 |
Au |
534 |
508 |
2.22 |
|
|
|
| 11 |
Au |
343 |
326 |
3.70 |
|
|
|
| 12 |
Au |
109 |
104 |
0.04 |
|
|
|
| 13 |
Au |
11 |
10 |
0.10 |
|
|
|
| 14 |
Bg |
590 |
561 |
0.00 |
|
|
|
| 15 |
Bg |
469 |
446 |
0.00 |
|
|
|
| 16 |
Bg |
179 |
171 |
0.00 |
|
|
|
| 17 |
Bu |
1823 |
1733 |
384.50 |
|
|
|
| 18 |
Bu |
1405 |
1336 |
400.06 |
|
|
|
| 19 |
Bu |
1262 |
1200 |
286.36 |
|
|
|
| 20 |
Bu |
985 |
937 |
273.10 |
|
|
|
| 21 |
Bu |
625 |
595 |
3.61 |
|
|
|
| 22 |
Bu |
495 |
470 |
2.83 |
|
|
|
| 23 |
Bu |
284 |
270 |
4.73 |
|
|
|
| 24 |
Bu |
136 |
129 |
0.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8742.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 8314.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.155 |
1.852 |
0.000 |
| C2 |
0.467 |
0.553 |
0.000 |
| C3 |
-0.467 |
-0.553 |
0.000 |
| C4 |
-0.155 |
-1.852 |
0.000 |
| F5 |
1.067 |
2.797 |
0.000 |
| F6 |
-1.067 |
2.330 |
0.000 |
| F7 |
1.767 |
0.223 |
0.000 |
| F8 |
-1.767 |
-0.223 |
0.000 |
| F9 |
1.067 |
-2.330 |
0.000 |
| F10 |
-1.067 |
-2.797 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3365 | 2.4835 | 3.7168 | 1.3135 | 1.3117 | 2.2916 | 2.8279 | 4.2801 | 4.8065 |
C2 | 1.3365 | | 1.4468 | 2.4835 | 2.3230 | 2.3477 | 1.3407 | 2.3644 | 2.9441 | 3.6838 | C3 | 2.4835 | 1.4468 | | 1.3365 | 3.6838 | 2.9441 | 2.3644 | 1.3407 | 2.3477 | 2.3230 | C4 | 3.7168 | 2.4835 | 1.3365 | | 4.8065 | 4.2801 | 2.8279 | 2.2916 | 1.3117 | 1.3135 | F5 | 1.3135 | 2.3230 | 3.6838 | 4.8065 | | 2.1846 | 2.6668 | 4.1412 | 5.1264 | 5.9866 | F6 | 1.3117 | 2.3477 | 2.9441 | 4.2801 | 2.1846 | | 3.5308 | 2.6471 | 5.1250 | 5.1264 | F7 | 2.2916 | 1.3407 | 2.3644 | 2.8279 | 2.6668 | 3.5308 | | 3.5613 | 2.6471 | 4.1412 | F8 | 2.8279 | 2.3644 | 1.3407 | 2.2916 | 4.1412 | 2.6471 | 3.5613 | | 3.5308 | 2.6668 | F9 | 4.2801 | 2.9441 | 2.3477 | 1.3117 | 5.1264 | 5.1250 | 2.6471 | 3.5308 | | 2.1846 | F10 | 4.8065 | 3.6838 | 2.3230 | 1.3135 | 5.9866 | 5.1264 | 4.1412 | 2.6668 | 2.1846 | |
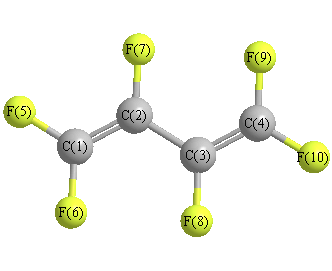 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
126.278 |
|
C1 |
C2 |
F7 |
117.736 |
| C2 |
C1 |
F5 |
122.474 |
|
C2 |
C1 |
F6 |
124.881 |
| C2 |
C3 |
C4 |
126.278 |
|
C2 |
C3 |
F8 |
115.986 |
| C3 |
C2 |
F7 |
115.986 |
|
C3 |
C4 |
F9 |
124.881 |
| C3 |
C4 |
F10 |
122.474 |
|
C4 |
C3 |
F8 |
117.736 |
| F5 |
C1 |
F6 |
112.646 |
|
F9 |
C4 |
F10 |
112.646 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.527 |
|
|
|
| 2 |
C |
0.197 |
|
|
|
| 3 |
C |
0.197 |
|
|
|
| 4 |
C |
0.527 |
|
|
|
| 5 |
F |
-0.227 |
|
|
|
| 6 |
F |
-0.222 |
|
|
|
| 7 |
F |
-0.274 |
|
|
|
| 8 |
F |
-0.274 |
|
|
|
| 9 |
F |
-0.222 |
|
|
|
| 10 |
F |
-0.227 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-53.390 |
-0.234 |
0.000 |
| y |
-0.234 |
-50.823 |
0.000 |
| z |
0.000 |
0.000 |
-48.339 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.809 |
-0.234 |
0.000 |
| y |
-0.234 |
0.041 |
0.000 |
| z |
0.000 |
0.000 |
3.768 |
|
| Polar |
|---|
| 3z2-r2 | 7.536 |
|---|
| x2-y2 | -2.567 |
|---|
| xy | -0.234 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.993 |
-0.106 |
0.000 |
| y |
-0.106 |
11.944 |
0.000 |
| z |
0.000 |
0.000 |
3.312 |
<r2> (average value of r
2) Å
2
| <r2> |
416.030 |
| (<r2>)1/2 |
20.397 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -750.702438 |
| Energy at 298.15K | -750.703523 |
| HF Energy | -750.702438 |
| Nuclear repulsion energy | 568.115681 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1868 |
1777 |
63.25 |
|
|
|
| 2 |
A |
1443 |
1372 |
6.57 |
|
|
|
| 3 |
A |
1398 |
1329 |
197.97 |
|
|
|
| 4 |
A |
1183 |
1125 |
268.80 |
|
|
|
| 5 |
A |
729 |
694 |
0.25 |
|
|
|
| 6 |
A |
633 |
602 |
1.37 |
|
|
|
| 7 |
A |
527 |
502 |
0.04 |
|
|
|
| 8 |
A |
472 |
449 |
0.08 |
|
|
|
| 9 |
A |
380 |
361 |
0.69 |
|
|
|
| 10 |
A |
253 |
240 |
0.42 |
|
|
|
| 11 |
A |
187 |
178 |
0.34 |
|
|
|
| 12 |
A |
99 |
94 |
0.20 |
|
|
|
| 13 |
A |
57 |
54 |
0.00 |
|
|
|
| 14 |
B |
1848 |
1758 |
270.38 |
|
|
|
| 15 |
B |
1397 |
1328 |
259.83 |
|
|
|
| 16 |
B |
1252 |
1191 |
84.80 |
|
|
|
| 17 |
B |
1006 |
957 |
181.30 |
|
|
|
| 18 |
B |
627 |
596 |
4.43 |
|
|
|
| 19 |
B |
595 |
566 |
1.24 |
|
|
|
| 20 |
B |
556 |
529 |
4.85 |
|
|
|
| 21 |
B |
406 |
386 |
3.30 |
|
|
|
| 22 |
B |
297 |
282 |
3.55 |
|
|
|
| 23 |
B |
206 |
196 |
3.12 |
|
|
|
| 24 |
B |
107 |
102 |
0.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8762.5 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 8333.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.172 |
1.544 |
-0.384 |
| C2 |
0.126 |
0.712 |
0.614 |
| C3 |
-0.126 |
-0.712 |
0.614 |
| C4 |
0.172 |
-1.544 |
-0.384 |
| F5 |
0.126 |
2.820 |
-0.383 |
| F6 |
-0.767 |
1.166 |
-1.492 |
| F7 |
0.693 |
1.218 |
1.722 |
| F8 |
-0.693 |
-1.218 |
1.722 |
| F9 |
0.767 |
-1.166 |
-1.492 |
| F10 |
-0.126 |
-2.820 |
-0.383 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3330 | 2.4668 | 3.1063 | 1.3103 | 1.3136 | 2.2999 | 3.5118 | 3.0748 | 4.3635 |
C2 | 1.3330 | | 1.4458 | 2.4668 | 2.3318 | 2.3329 | 1.3434 | 2.3711 | 2.8941 | 3.6782 | C3 | 2.4668 | 1.4458 | | 1.3330 | 3.6782 | 2.8941 | 2.3711 | 1.3434 | 2.3329 | 2.3318 | C4 | 3.1063 | 2.4668 | 1.3330 | | 4.3635 | 3.0748 | 3.5118 | 2.2999 | 1.3136 | 1.3103 | F5 | 1.3103 | 2.3318 | 3.6782 | 4.3635 | | 2.1823 | 2.7054 | 4.6264 | 4.1869 | 5.6449 | F6 | 1.3136 | 2.3329 | 2.8941 | 3.0748 | 2.1823 | | 3.5310 | 4.0028 | 2.7925 | 4.1869 | F7 | 2.2999 | 1.3434 | 2.3711 | 3.5118 | 2.7054 | 3.5310 | | 2.8023 | 4.0028 | 4.6264 | F8 | 3.5118 | 2.3711 | 1.3434 | 2.2999 | 4.6264 | 4.0028 | 2.8023 | | 3.5310 | 2.7054 | F9 | 3.0748 | 2.8941 | 2.3329 | 1.3136 | 4.1869 | 2.7925 | 4.0028 | 3.5310 | | 2.1823 | F10 | 4.3635 | 3.6782 | 2.3318 | 1.3103 | 5.6449 | 4.1869 | 4.6264 | 2.7054 | 2.1823 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.130 |
|
C1 |
C2 |
F7 |
118.476 |
| C2 |
C1 |
F5 |
123.805 |
|
C2 |
C1 |
F6 |
123.633 |
| C2 |
C3 |
C4 |
125.130 |
|
C2 |
C3 |
F8 |
116.390 |
| C3 |
C2 |
F7 |
116.390 |
|
C3 |
C4 |
F9 |
123.633 |
| C3 |
C4 |
F10 |
123.805 |
|
C4 |
C3 |
F8 |
118.476 |
| F5 |
C1 |
F6 |
112.542 |
|
F9 |
C4 |
F10 |
112.542 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.517 |
|
|
|
| 2 |
C |
0.201 |
|
|
|
| 3 |
C |
0.201 |
|
|
|
| 4 |
C |
0.517 |
|
|
|
| 5 |
F |
-0.221 |
|
|
|
| 6 |
F |
-0.225 |
|
|
|
| 7 |
F |
-0.272 |
|
|
|
| 8 |
F |
-0.272 |
|
|
|
| 9 |
F |
-0.225 |
|
|
|
| 10 |
F |
-0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.911 |
0.911 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.228 |
-1.119 |
0.000 |
| y |
-1.119 |
-51.185 |
0.000 |
| z |
0.000 |
0.000 |
-52.715 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.722 |
-1.119 |
0.000 |
| y |
-1.119 |
-0.213 |
0.000 |
| z |
0.000 |
0.000 |
-2.509 |
|
| Polar |
|---|
| 3z2-r2 | -5.018 |
|---|
| x2-y2 | 1.957 |
|---|
| xy | -1.119 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.986 |
-0.147 |
0.000 |
| y |
-0.147 |
9.024 |
0.000 |
| z |
0.000 |
0.000 |
6.821 |
<r2> (average value of r
2) Å
2
| <r2> |
383.322 |
| (<r2>)1/2 |
19.579 |
