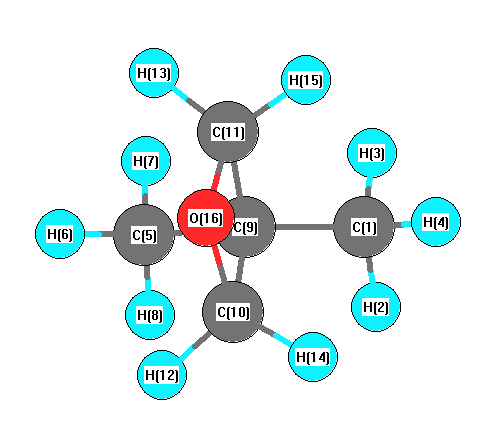
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3132 |
2996 |
30.35 |
|
|
|
| 2 |
A' |
3129 |
2993 |
29.20 |
|
|
|
| 3 |
A' |
3082 |
2949 |
85.70 |
|
|
|
| 4 |
A' |
3052 |
2920 |
20.81 |
|
|
|
| 5 |
A' |
3046 |
2914 |
25.74 |
|
|
|
| 6 |
A' |
3032 |
2901 |
24.93 |
|
|
|
| 7 |
A' |
1563 |
1495 |
1.58 |
|
|
|
| 8 |
A' |
1530 |
1464 |
4.00 |
|
|
|
| 9 |
A' |
1513 |
1448 |
3.81 |
|
|
|
| 10 |
A' |
1445 |
1382 |
3.16 |
|
|
|
| 11 |
A' |
1426 |
1365 |
8.07 |
|
|
|
| 12 |
A' |
1398 |
1337 |
0.19 |
|
|
|
| 13 |
A' |
1312 |
1255 |
0.66 |
|
|
|
| 14 |
A' |
1284 |
1229 |
10.39 |
|
|
|
| 15 |
A' |
1174 |
1123 |
0.67 |
|
|
|
| 16 |
A' |
1042 |
997 |
5.81 |
|
|
|
| 17 |
A' |
994 |
951 |
10.05 |
|
|
|
| 18 |
A' |
970 |
928 |
4.32 |
|
|
|
| 19 |
A' |
941 |
900 |
5.83 |
|
|
|
| 20 |
A' |
852 |
815 |
8.48 |
|
|
|
| 21 |
A' |
644 |
616 |
0.57 |
|
|
|
| 22 |
A' |
402 |
384 |
0.10 |
|
|
|
| 23 |
A' |
342 |
327 |
0.14 |
|
|
|
| 24 |
A' |
74 |
71 |
4.98 |
|
|
|
| 25 |
A" |
3124 |
2989 |
51.29 |
|
|
|
| 26 |
A" |
3120 |
2985 |
1.83 |
|
|
|
| 27 |
A" |
3081 |
2947 |
13.61 |
|
|
|
| 28 |
A" |
3021 |
2890 |
124.26 |
|
|
|
| 29 |
A" |
1543 |
1476 |
10.18 |
|
|
|
| 30 |
A" |
1526 |
1460 |
1.79 |
|
|
|
| 31 |
A" |
1513 |
1447 |
0.11 |
|
|
|
| 32 |
A" |
1315 |
1258 |
2.52 |
|
|
|
| 33 |
A" |
1238 |
1185 |
0.35 |
|
|
|
| 34 |
A" |
1169 |
1118 |
0.08 |
|
|
|
| 35 |
A" |
1069 |
1023 |
0.00 |
|
|
|
| 36 |
A" |
1066 |
1020 |
87.20 |
|
|
|
| 37 |
A" |
947 |
906 |
0.02 |
|
|
|
| 38 |
A" |
898 |
859 |
3.67 |
|
|
|
| 39 |
A" |
390 |
373 |
0.60 |
|
|
|
| 40 |
A" |
311 |
297 |
0.17 |
|
|
|
| 41 |
A" |
276 |
264 |
0.01 |
|
|
|
| 42 |
A" |
225 |
215 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31603.2 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30234.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.525 |
|
|
-0.539 |
| 2 |
H |
0.170 |
|
|
0.136 |
| 3 |
H |
0.170 |
|
|
0.136 |
| 4 |
H |
0.171 |
|
|
0.134 |
| 5 |
C |
-0.524 |
|
|
-0.433 |
| 6 |
H |
0.179 |
|
|
0.118 |
| 7 |
H |
0.168 |
|
|
0.109 |
| 8 |
H |
0.168 |
|
|
0.109 |
| 9 |
C |
-0.007 |
|
|
0.310 |
| 10 |
C |
-0.042 |
|
|
0.075 |
| 11 |
C |
-0.042 |
|
|
0.075 |
| 12 |
H |
0.157 |
|
|
0.041 |
| 13 |
H |
0.157 |
|
|
0.041 |
| 14 |
H |
0.150 |
|
|
0.038 |
| 15 |
H |
0.150 |
|
|
0.038 |
| 16 |
O |
-0.500 |
|
|
-0.388 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.082 |
2.036 |
0.000 |
2.038 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.076 |
2.063 |
0.000 |
2.064 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.271 |
0.081 |
0.000 |
| y |
0.081 |
-43.016 |
0.000 |
| z |
0.000 |
0.000 |
-35.548 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.011 |
0.081 |
0.000 |
| y |
0.081 |
-6.606 |
0.000 |
| z |
0.000 |
0.000 |
4.595 |
|
| Polar |
|---|
| 3z2-r2 | 9.191 |
|---|
| x2-y2 | 5.745 |
|---|
| xy | 0.081 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
158.502 |
| (<r2>)1/2 |
12.590 |