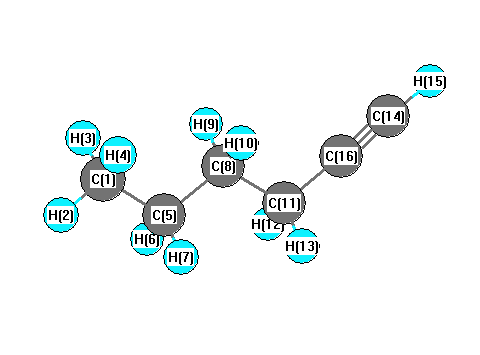
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3501 |
3350 |
57.34 |
|
|
|
| 2 |
A' |
3132 |
2996 |
36.70 |
|
|
|
| 3 |
A' |
3059 |
2927 |
52.52 |
|
|
|
| 4 |
A' |
3052 |
2920 |
20.26 |
|
|
|
| 5 |
A' |
3037 |
2905 |
30.82 |
|
|
|
| 6 |
A' |
3035 |
2904 |
1.81 |
|
|
|
| 7 |
A' |
2244 |
2147 |
5.70 |
|
|
|
| 8 |
A' |
1535 |
1469 |
5.66 |
|
|
|
| 9 |
A' |
1522 |
1456 |
1.13 |
|
|
|
| 10 |
A' |
1513 |
1448 |
0.60 |
|
|
|
| 11 |
A' |
1498 |
1433 |
1.21 |
|
|
|
| 12 |
A' |
1437 |
1375 |
2.74 |
|
|
|
| 13 |
A' |
1414 |
1353 |
1.42 |
|
|
|
| 14 |
A' |
1371 |
1312 |
2.74 |
|
|
|
| 15 |
A' |
1284 |
1228 |
4.60 |
|
|
|
| 16 |
A' |
1136 |
1087 |
3.81 |
|
|
|
| 17 |
A' |
1080 |
1034 |
0.27 |
|
|
|
| 18 |
A' |
1043 |
997 |
0.92 |
|
|
|
| 19 |
A' |
964 |
922 |
0.16 |
|
|
|
| 20 |
A' |
916 |
877 |
3.32 |
|
|
|
| 21 |
A' |
619 |
592 |
51.90 |
|
|
|
| 22 |
A' |
520 |
498 |
3.46 |
|
|
|
| 23 |
A' |
384 |
367 |
0.03 |
|
|
|
| 24 |
A' |
276 |
264 |
0.51 |
|
|
|
| 25 |
A' |
121 |
115 |
0.03 |
|
|
|
| 26 |
A" |
3127 |
2992 |
60.67 |
|
|
|
| 27 |
A" |
3107 |
2973 |
30.60 |
|
|
|
| 28 |
A" |
3072 |
2939 |
0.00 |
|
|
|
| 29 |
A" |
3067 |
2934 |
10.70 |
|
|
|
| 30 |
A" |
1527 |
1460 |
7.65 |
|
|
|
| 31 |
A" |
1341 |
1283 |
0.04 |
|
|
|
| 32 |
A" |
1327 |
1269 |
0.89 |
|
|
|
| 33 |
A" |
1252 |
1198 |
0.17 |
|
|
|
| 34 |
A" |
1144 |
1094 |
0.01 |
|
|
|
| 35 |
A" |
942 |
902 |
0.51 |
|
|
|
| 36 |
A" |
798 |
764 |
0.55 |
|
|
|
| 37 |
A" |
740 |
708 |
3.91 |
|
|
|
| 38 |
A" |
606 |
579 |
52.32 |
|
|
|
| 39 |
A" |
359 |
344 |
3.06 |
|
|
|
| 40 |
A" |
252 |
241 |
0.03 |
|
|
|
| 41 |
A" |
116 |
111 |
0.00 |
|
|
|
| 42 |
A" |
92 |
88 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31280.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29926.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.509 |
|
|
-0.316 |
| 2 |
H |
0.169 |
|
|
0.083 |
| 3 |
H |
0.168 |
|
|
0.075 |
| 4 |
H |
0.168 |
|
|
0.075 |
| 5 |
C |
-0.305 |
|
|
0.127 |
| 6 |
H |
0.160 |
|
|
-0.011 |
| 7 |
H |
0.160 |
|
|
-0.011 |
| 8 |
C |
-0.305 |
|
|
0.023 |
| 9 |
H |
0.173 |
|
|
0.031 |
| 10 |
H |
0.173 |
|
|
0.031 |
| 11 |
C |
-0.455 |
|
|
-0.249 |
| 12 |
H |
0.194 |
|
|
0.111 |
| 13 |
H |
0.194 |
|
|
0.111 |
| 14 |
C |
-0.546 |
|
|
-0.525 |
| 15 |
H |
0.221 |
|
|
0.319 |
| 16 |
C |
0.341 |
|
|
0.127 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.654 |
0.517 |
0.000 |
0.834 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.639 |
0.515 |
0.000 |
0.821 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.409 |
0.886 |
0.000 |
| y |
0.886 |
-38.967 |
0.000 |
| z |
0.000 |
0.000 |
-38.207 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.177 |
0.886 |
0.000 |
| y |
0.886 |
-2.158 |
0.000 |
| z |
0.000 |
0.000 |
-1.019 |
|
| Polar |
|---|
| 3z2-r2 | -2.038 |
|---|
| x2-y2 | 3.557 |
|---|
| xy | 0.886 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
266.079 |
| (<r2>)1/2 |
16.312 |