return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1-Hexyne)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-234.293309
Energy at 298.15K-234.302649
HF Energy-234.036281
Nuclear repulsion energy210.193067
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3517 3338 54.21      
2 A' 3153 2993 37.50      
3 A' 3080 2924 58.05      
4 A' 3076 2920 15.03      
5 A' 3062 2907 19.33      
6 A' 3060 2904 8.89      
7 A' 2208 2096 1.78      
8 A' 1561 1482 3.96      
9 A' 1548 1470 0.87      
10 A' 1539 1460 0.60      
11 A' 1527 1449 1.00      
12 A' 1462 1387 1.86      
13 A' 1436 1363 1.64      
14 A' 1392 1321 2.87      
15 A' 1304 1238 5.91      
16 A' 1149 1091 2.97      
17 A' 1086 1031 0.20      
18 A' 1046 992 0.73      
19 A' 963 914 0.35      
20 A' 923 876 2.97      
21 A' 585 555 51.45      
22 A' 505 479 0.49      
23 A' 388 368 0.05      
24 A' 272 258 0.27      
25 A' 120 113 0.02      
26 A" 3150 2990 60.17      
27 A" 3127 2968 36.27      
28 A" 3097 2939 0.17      
29 A" 3091 2934 9.11      
30 A" 1551 1473 6.33      
31 A" 1359 1290 0.06      
32 A" 1345 1277 0.81      
33 A" 1273 1209 0.07      
34 A" 1161 1102 0.03      
35 A" 957 908 0.25      
36 A" 811 770 0.52      
37 A" 750 711 2.62      
38 A" 571 542 50.38      
39 A" 333 316 1.05      
40 A" 255 242 0.03      
41 A" 117 111 0.00      
42 A" 92 87 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 31499.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 29899.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.49439 0.04390 0.04156

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.322 2.587 0.000
H2 2.321 3.030 0.000
H3 0.785 2.943 0.876
H4 0.785 2.943 -0.876
C5 1.398 1.054 0.000
H6 1.942 0.715 0.887
H7 1.942 0.715 -0.887
C8 0.000 0.414 0.000
H9 -0.548 0.748 0.886
H10 -0.548 0.748 -0.886
C11 0.070 -1.131 0.000
H12 0.624 -1.451 0.896
H13 0.624 -1.451 -0.896
C14 -2.334 -2.233 0.000
H15 -3.284 -2.648 0.000
C16 -1.229 -1.740 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 C16
C11.09301.08801.08801.53432.16212.16212.54382.76812.76813.92314.19444.19446.04976.97245.0226
H21.09301.77111.77112.18042.50752.50753.49753.77143.77144.73074.87424.87427.02637.97825.9458
H31.08801.77111.75292.17052.51073.06792.78972.56793.11444.22814.39694.74066.10636.96985.1721
H41.08801.77111.75292.17053.06792.51072.78973.11442.56794.22814.74064.39696.10636.96985.1721
C51.53432.18042.17052.17051.09421.09421.53782.15992.15992.55732.77092.77094.97355.96883.8352
H62.16212.50752.51073.06791.09421.77332.15602.49023.05662.77462.53533.09915.26906.27744.1071
H72.16212.50753.06792.51071.09421.77332.15603.05662.49022.77463.09912.53535.26906.27744.1071
C82.54383.49752.78972.78971.53782.15602.15601.09411.09411.54622.16062.16063.52884.48952.4793
H92.76813.77142.56793.11442.15992.49023.05661.09411.77182.16742.49193.06333.58644.44992.7271
H102.76813.77143.11442.56792.15993.05662.49021.09411.77182.16743.06332.49193.58644.44992.7271
C113.92314.73074.22814.22812.55732.77462.77461.54622.16742.16741.10081.10082.64463.68101.4347
H124.19444.87424.39694.74062.77092.53533.09912.16062.49193.06331.10081.79143.18844.18442.0788
H134.19444.87424.74064.39692.77093.09912.53532.16063.06332.49191.10081.79143.18844.18442.0788
C146.04977.02636.10636.10634.97355.26905.26903.52883.58643.58642.64463.18843.18841.03651.2100
H156.97247.97826.96986.96985.96886.27746.27744.48954.44994.44993.68104.18444.18441.03652.2465
C165.02265.94585.17215.17213.83524.10714.10712.47932.72712.72711.43472.07882.07881.21002.2465

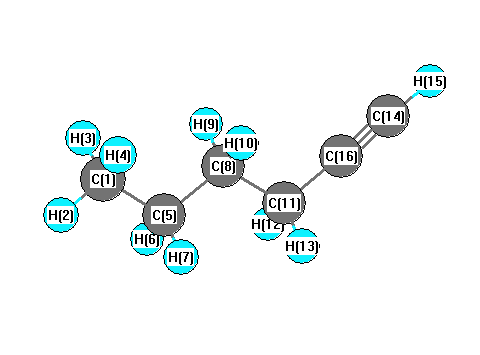
picture of 1-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.539 C1 C5 H7 109.539
C1 C5 C8 111.789 H2 C1 H3 108.595
H2 C1 H4 108.595 H2 C1 C5 111.057
H3 C1 H4 107.333 H3 C1 C5 110.575
H4 C1 C5 110.575 C5 C8 H9 109.141
C5 C8 H10 109.141 C5 C8 C11 112.037
H6 C5 H7 108.247 H6 C5 C8 108.826
H7 C5 C8 108.826 C8 C11 H12 108.243
C8 C11 H13 108.243 C8 C11 C16 112.500
H9 C8 H10 108.132 H9 C8 C11 109.155
H10 C8 C11 109.155 C11 C16 C14 178.979
H12 C11 H13 108.914 H12 C11 C16 109.435
H13 C11 C16 109.435 H15 C14 C16 179.499
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.460      
2 H 0.154      
3 H 0.152      
4 H 0.152      
5 C -0.275      
6 H 0.144      
7 H 0.144      
8 C -0.274      
9 H 0.157      
10 H 0.157      
11 C -0.384      
12 H 0.179      
13 H 0.179      
14 C -0.506      
15 H 0.227      
16 C 0.257      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 10.340 2.452 0.000
y 2.452 9.122 0.000
z 0.000 0.000 6.674


<r2> (average value of r2) Å2
<r2> 266.483
(<r2>)1/2 16.324