return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (Ethenylcyclopropane)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-194.632806
Energy at 298.15K-194.641002
HF Energy-193.938624
Nuclear repulsion energy162.574613
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3256 3101 33.47      
2 A' 3255 3100 7.84      
3 A' 3178 3026 7.05      
4 A' 3175 3024 4.54      
5 A' 3170 3019 15.97      
6 A' 3149 2999 10.14      
7 A' 1745 1662 15.23      
8 A' 1564 1489 1.27      
9 A' 1507 1436 0.22      
10 A' 1396 1329 0.85      
11 A' 1346 1282 3.24      
12 A' 1268 1207 2.39      
13 A' 1255 1195 1.50      
14 A' 1109 1056 1.99      
15 A' 1034 984 4.00      
16 A' 993 945 30.08      
17 A' 847 807 1.01      
18 A' 792 754 0.30      
19 A' 450 429 0.89      
20 A' 273 260 1.03      
21 A" 3242 3088 0.13      
22 A" 3164 3013 19.69      
23 A" 1517 1444 1.15      
24 A" 1233 1174 0.60      
25 A" 1165 1110 1.42      
26 A" 1112 1059 4.13      
27 A" 1019 971 12.02      
28 A" 961 915 7.77      
29 A" 913 869 40.74      
30 A" 858 817 6.64      
31 A" 679 646 3.98      
32 A" 318 303 1.09      
33 A" 114 108 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 25527.0 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 24309.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.50390 0.10128 0.09741

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.387 0.291 0.000
C2 0.258 -1.045 0.000
C3 -0.404 -2.210 0.000
C4 0.258 1.436 0.751
C5 0.258 1.436 -0.751
H6 -1.478 0.272 0.000
H7 1.352 -1.048 0.000
H8 0.119 -3.164 0.000
H9 -1.492 -2.250 0.000
H10 -0.393 2.135 1.271
H11 1.196 1.235 1.265
H12 -0.393 2.135 -1.271
H13 1.196 1.235 -1.265

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.48332.50051.51371.51371.09042.19503.49132.77122.24022.23592.24022.2359
C21.48331.33962.59172.59172.17841.09382.12342.12543.48602.77103.48602.7710
C32.50051.33963.78033.78032.70392.10481.08771.08944.52704.00324.52704.0032
C41.51372.59173.78031.50252.22052.81644.66234.14921.08781.08842.23702.2330
C51.51372.59173.78031.50252.22052.81644.66234.14922.23702.23301.08781.0884
H61.09042.17842.70392.22052.22053.12223.78832.52252.50313.11082.50313.1108
H72.19501.09382.10482.81642.81643.12222.44833.08773.84652.61523.84652.6152
H83.49132.12341.08774.66234.66233.78832.44831.85185.47324.70215.47324.7021
H92.77122.12541.08944.14924.14922.52253.08771.85184.69694.58024.69694.5802
H102.24023.48604.52701.08782.23702.50313.84655.47324.69691.82622.54293.1256
H112.23592.77104.00321.08842.23303.11082.61524.70214.58021.82623.12562.5304
H122.24023.48604.52702.23701.08782.50313.84655.47324.69692.54293.12561.8262
H132.23592.77104.00322.23301.08843.11082.61524.70214.58023.12562.53041.8262

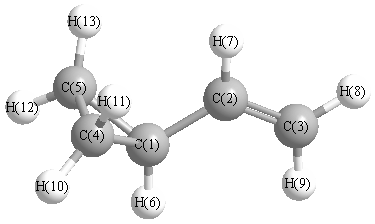
picture of Ethenylcyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.617 C1 C2 H7 115.977
C1 C4 C5 60.246 C1 C4 H10 117.953
C1 C4 H11 117.536 C1 C5 C4 60.246
C1 C5 H12 117.953 C1 C5 H13 117.536
C2 C1 C4 119.711 C2 C1 C5 119.711
C2 C1 H6 114.789 C2 C3 H8 121.695
C2 C3 H9 121.753 C3 C2 H7 119.407
C4 C1 H6 116.059 C4 C5 H12 118.571
C4 C5 H13 118.176 C5 C1 H6 116.059
C5 C4 H10 118.571 C5 C4 H11 118.176
H8 C3 H9 116.552 H10 C4 H11 114.100
H12 C5 H13 114.100
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability